Readable HTML

English — complete official images and native search text

CDSCO Medical Devices FAQ Portal Compilation — Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation

Currentness under review. Whether this instrument remains applicable is under review. Consult the later instruments and source context below before relying on it.

Original page images are authoritative. Searchable text is the PDF text layer and may omit image text or contain extraction artifacts. Quote and verify against the page images.

Link directly to a passage with its anchor — for example #pdf-page-1 — to open it highlighted. This reference version is generated from the authority’s publication and does not replace it. Language: English.

Contents
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 1
Searchable PDF text layer

Central Drugs Standard Control Organization (Medical Devices Division) Medical Devices Frequently Asked Questions Doc No.: CDSCO/FAQ/MD/01/2024 Notice: The replies to the FAQs are aimed only for creating public awareness about Medical Devices Regulation by CDSCO and are not meant to be used for legal or professional purposes. The readers are advised to refer to the statutory provisions of Drugs and Cosmetics Act & Rules and respective Guidelines / Clarifications issued by CDSCO from time to time for all their professional needs. Page 1 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 2
Searchable PDF text layer

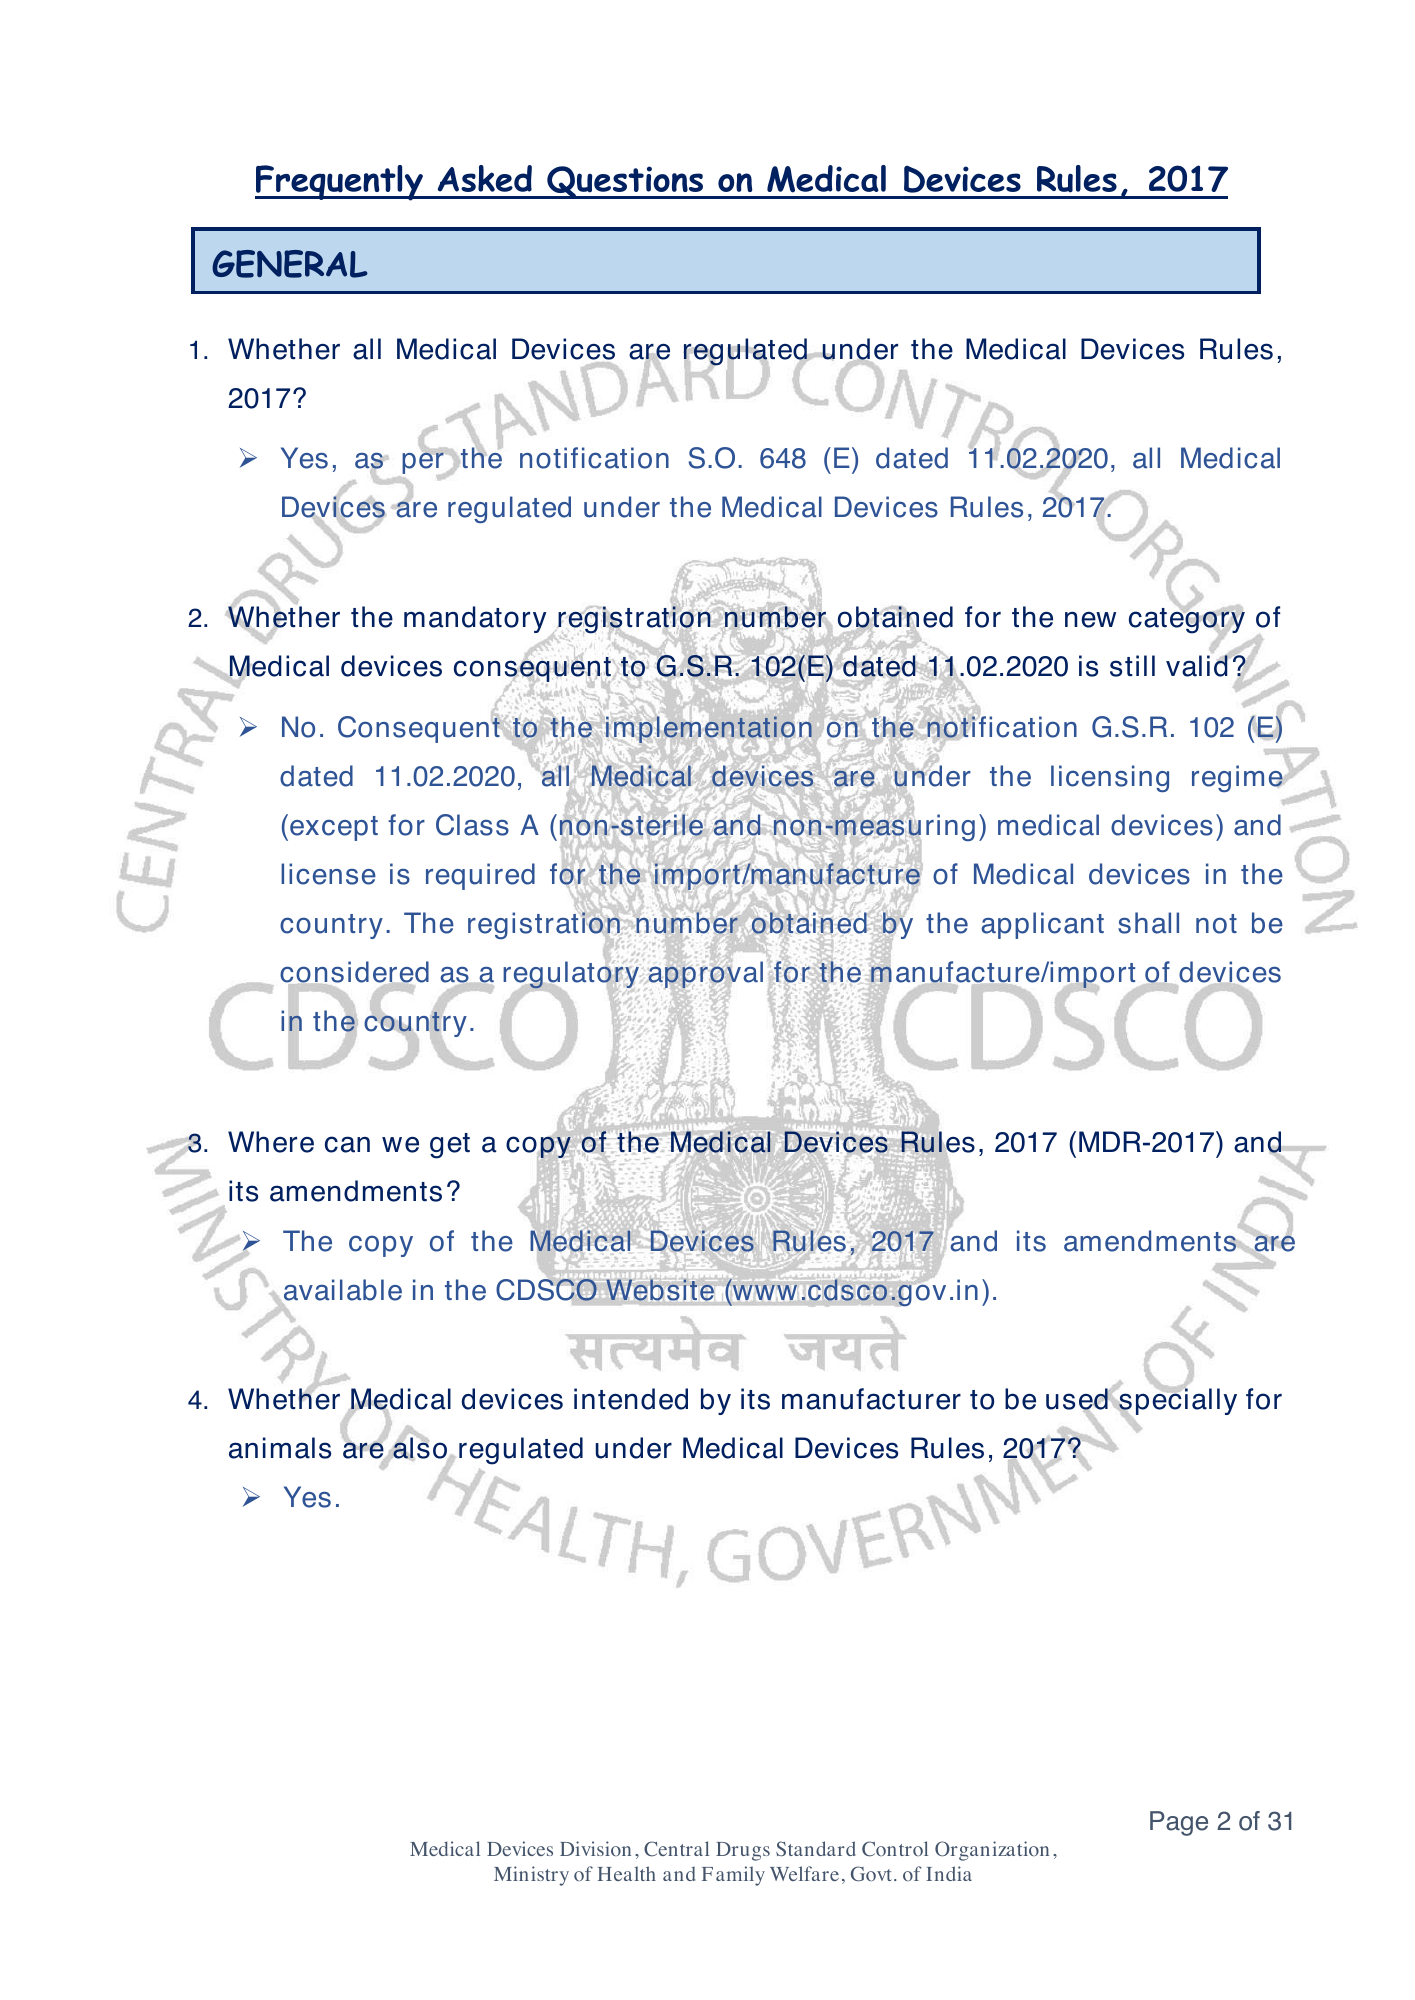
Frequently Asked Questions on Medical Devices Rules, 2017 GENERAL 1. Whether all Medical Devices are regulated under the Medical Devices Rules, 2017?  Yes, as per the notification S.O. 648 (E) dated 11.02.2020, all Medical Devices are regulated under the Medical Devices Rules, 2017. 2. Whether the mandatory registration number obtained for the new category of Medical devices consequent to G.S.R. 102(E) dated 11.02.2020 is still valid?  No. Consequent to the implementation on the notification G.S.R. 102 (E) dated 11.02.2020, all Medical devices are under the licensing regime (except for Class A (non-sterile and non-measuring) medical devices) and license is required for the import/manufacture of Medical devices in the country. The registration number obtained by the applicant shall not be considered as a regulatory approval for the manufacture/import of devices in the country. 3. Where can we get a copy of the Medical Devices Rules, 2017 (MDR-2017) and its amendments?  The copy of the Medical Devices Rules, 2017 and its amendments are available in the CDSCO Website (www.cdsco.gov.in). 4. Whether Medical devices intended by its manufacturer to be used specially for animals are also regulated under Medical Devices Rules, 2017?  Yes. Page 2 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 3
Searchable PDF text layer

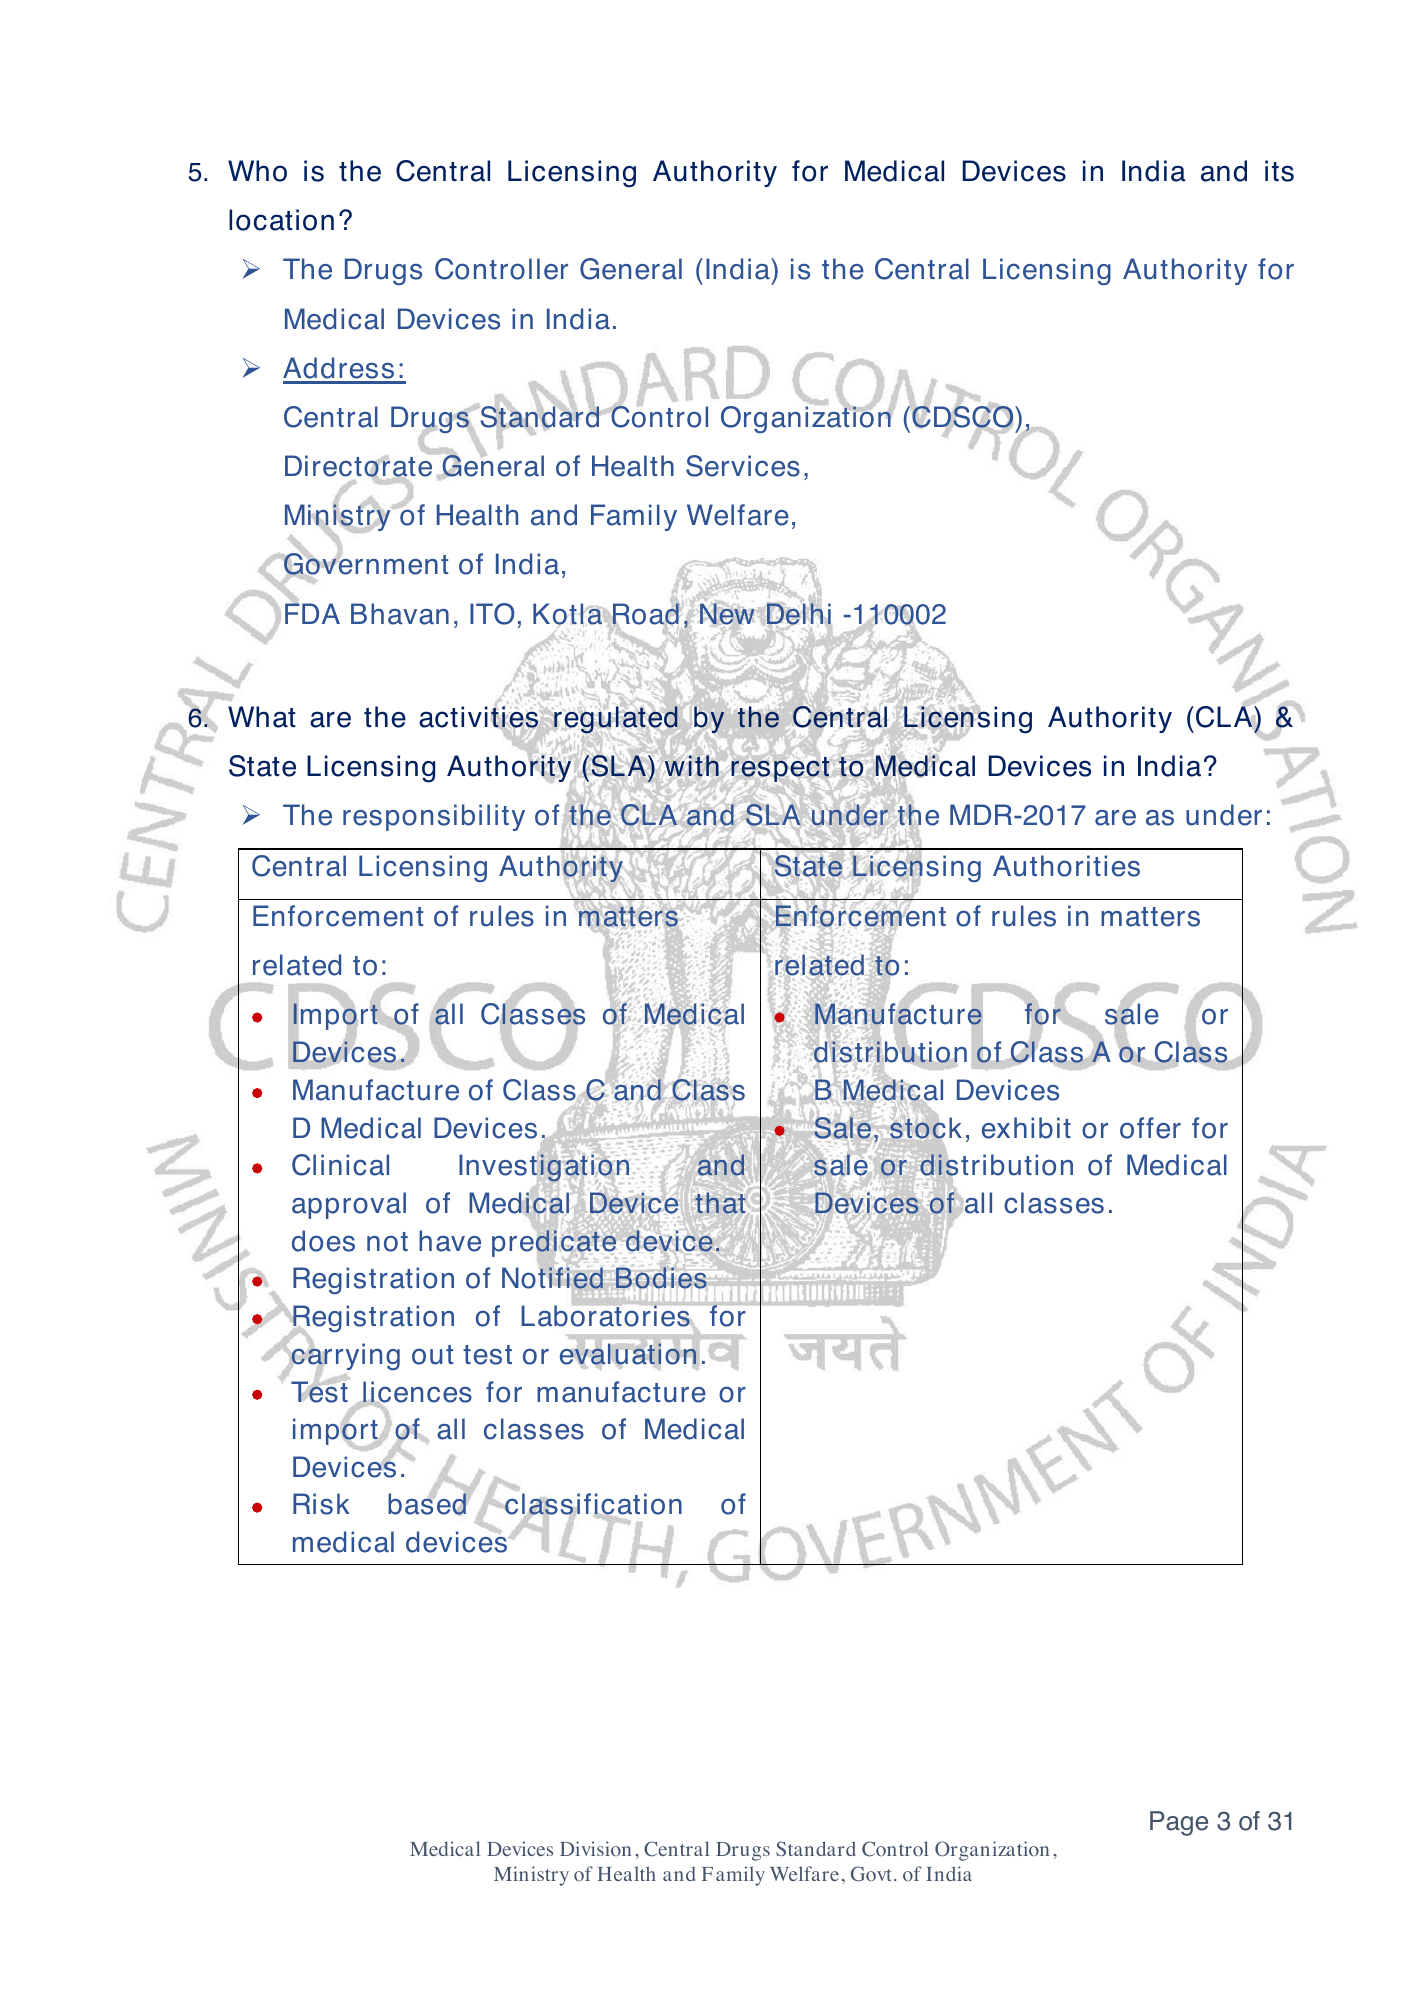
5. Who is the Central Licensing Authority for Medical Devices in India and its location?  The Drugs Controller General (India) is the Central Licensing Authority for Medical Devices in India.  Address: Central Drugs Standard Control Organization (CDSCO), Directorate General of Health Services, Ministry of Health and Family Welfare, Government of India, FDA Bhavan, ITO, Kotla Road, New Delhi -110002 6. What are the activities regulated by the Central Licensing Authority (CLA) & State Licensing Authority (SLA) with respect to Medical Devices in India?  The responsibility of the CLA and SLA under the MDR-2017 are as under: Central Licensing Authority State Licensing Authorities Enforcement of rules in matters Enforcement of rules in matters related to: related to:  Import of all Classes of Medical  Manufacture for sale or Devices. distribution of Class A or Class  Manufacture of Class C and Class B Medical Devices D Medical Devices.  Sale, stock, exhibit or offer for  Clinical Investigation and sale or distribution of Medical approval of Medical Device that Devices of all classes. does not have predicate device.  Registration of Notified Bodies  Registration of Laboratories for carrying out test or evaluation.  Test licences for manufacture or import of all classes of Medical Devices.  Risk based classification of medical devices Page 3 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 4
Searchable PDF text layer

7. Whether CDSCO provides any consultation system for Start-ups/ Innovators/ Importers/ Manufacturers?  Yes. The Public Relation Office (PRO) cell is established in CDSCO Head Quarter to address the issue of startups/ innovators/ importers/ manufacturers in the field of Medical devices pertaining to regulatory pathway.  Link: https://cdsco.gov.in/opencms/opencms/en/PRO/;  Email ID: startupinnov@cdsco.nic.in 8. What is the mode of submission of various applications in MDR-2017 to the Licensing Authority?  The applicant is required to submit their application through online mode for which specific provision for submission of application is available as under: Website link: www.cdscomdonline.gov.in , www.nsws.gov.in 9. What are the details of the Plant Master File (PMF) / Site Master File (SMF)?  The contents of PMF/SMF have been prescribed in Appendix - I of Fourth Schedule of MDR-2017. 10. Whether Essential principles for safety and performance of medical devices shall be applicable for all risk class of medical devices?  Yes 11. Can Third party / Authorized Consultant ask the status of the application to the Licensing Authority?  Only authorized person of the company can ask the status of their application. 12. What are the essential documents required for submission to the Licensing Authority under various application forms prescribed under Medical Devices Rules, 2017?  The essential documents shall be submitted as per the checklist available in the Medical Devices Online portal under Medical Devices Rules, 2017. Page 4 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 5
Searchable PDF text layer

13. What is the Fee required to be submitted for various applications under Medical Devices Rules, 2017?  The Fee required to be submitted for various applications is prescribed in the Second Schedule of Medical Devices Rules, 2017. 14. What is the method for getting refund of challan amount if any manufacturer/importer does not want to register the product or withdraw their application?  As per Medical Devices Rules, 2017, there is no provision/ clause for the refund of the fee paid by the applicant. 15. Whether a separate fee is required for multiple Brands of a device applied for grant of manufacturing or import licence?  Yes, a separate fee is required to be submitted for each applied Brands of a device as per Second Schedule of MDR-2017. 16. Whether the manufacturer may send their devices to private testing laboratory for test or evaluation?  Yes, the manufacturers may send their devices for test or evaluation to the Medical Device Testing Laboratory (MDTL) registered with CDSCO under MDR-2017. A list of the registered MDTL is published on CDSCO website which is dynamic in nature. 17. What is the approval procedure for addition of medical devices as an additional product in the existing licence?  An endorsement application along with requisite fees and requisite documents as per Fourth Schedule of MDR-2017 & checklist available in the Medical Devices Online portal. Page 5 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 6
Searchable PDF text layer

18. What is a License granted for the purpose of Clinical Investigations or Test or Evaluation or Demonstration or Training under MDR-2017?  Under MDR-2017, the License for Manufacture is issued in Form MD-13 and License for Import is issued in Form MD-17 for the purposes of Clinical Investigations or Test or Evaluation or Demonstration or Training. 19. Whether the License granted in Form MD-13/ Form MD-17 for the purpose of Clinical Investigations or Test or Evaluation or Demonstration or Training under MDR-2017 is issued in perpetuity?  No, the licence is issued with the validity of three years from the date of issue of the license under MDR-2017. 20. What is Central Medical Device Testing Laboratory (CMDTL)?  Central medical devices testing laboratory (CMDTL) means a medical devices testing laboratory established or designated by the Central Government under rule 19 of MDR-2017. 21. What is considered as “Change of constitution” of the company under the MDR- 2017?  The Change of constitution is defined under Rule 3(j) of MDR-2017 as under: o a firm means change from proprietorship to partnership including Limited Liability Partnership or vice versa; o a company means- (A) its conversion from a private to a public company, or from a public to a private company; or (B) any change in the ownership of shares of more than fifty per cent. Of the voting capital in the body corporate or in case of a body corporate not having a share capital, any change in its membership; and where the managing agent, being a body corporate is a subsidiary of another body corporate, includes a change in the constitution of that other body corporate within the meaning of this clause; Page 6 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 7
Searchable PDF text layer

22. In case of Change of constitution of the company whether fresh license is required?  Yes, the applicant shall inform to the Licensing Authority about such change within 45 days and submit an application under MDR-2017 within a period of 180 days from the date of such change in constitution. 23. Who are the competent body to audit the manufacturing facility of Class A & Class B medical devices in the country?  Only those Notified body who are registered with the CDSCO are eligible for audit the manufacturing facility of Class A & Class B medical devices in the country. 24. Where can we get a list of registered Notified bodies?  The list of Notified bodies registered with CDSCO is available on the CDSCO website. The list is dynamic in nature. 25. If a device is complying with ISO/IEC standards, would it still need to follow BIS standards?  If the BIS standard is available for the particular medical device, then in such cases, the fulfilment of BIS standard is mandatory. In case, there is no BIS standard available, then other international standards (ISO/IEC) would be applicable. 26. In case if there is no BIS/ISO/IEC standard for medical devices then which standard to be followed?  In such cases, the medical device shall conform to any other pharmacopoeial standards or validated manufacturers standard. Page 7 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 8
Searchable PDF text layer

27. What is the timeline for carrying out the inspection for Class C & Class D medical devices for grant of manufacturing licence?  For Class C and Class D medical device, the inspection will be carried out by the Medical Device Officers within a period of 60 days from the date of application, provided the data submitted by the applicant is found satisfactory.  28. Is it sufficient to provide a single IFU (Instructions for use) for Multi pack of a medical device used by Health Care Professionals?  The medical device when offered for sale as single or multi pack, shall be accompanied by either its IFU (Instructions for use), prescribing information, relevant literature, etc. 29. Will e-IFU (electronic Instructions for use) be allowed under the MDR-2017?  Yes. 30. Who is responsible for the submission of Post Marketing Surveillance (PMS) data?  It is the responsibility of the licensed holder for the submission of Post Marketing Surveillance (PMS) data to the Licensing Authority. 31. What are the Labeling requirements for Medical Devices in India?  Product labels shall comply with the Labelling requirements as prescribed in Chapter VI of Medical Devices Rules, 2017. 32. Can the date of manufacture/sterilization/expiry be mentioned as DD/MM/YY or MM/YY?  The date of expiry shall be in terms of the month and the year. 33. What is the residual shelf life applicable for Medical Devices?  The Residual shelf life for the medical devices is specified in Rule 47 of MDR-2017. Page 8 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 9
Searchable PDF text layer

34. What are the documents to be provided to support the Claimed Shelf Life of a Medical Device?  A real-time aging data shall be submitted to support the claimed shelf life. However, if real-time data is not available, accelerated stability data shall be submitted to support the claimed shelf life. Such a provisional claimed shelf life may be approved provided that the manufacturer immediately initiates real-time stability testing to validate the proposed shelf life and the real-time stability data shall be submitted after completion of the study by the manufacturer. 35. What are the documents to be provided to get approval for Shelf Life of a medical device beyond 5 years under MDR-2017?  A real-time aging data of the medical device generated by the manufacturer along with the accelerated stability data and other relevant supporting data shall be submitted to support the claimed shelf life beyond 5 years. 36. Whether Shelf life is applicable for Medical Equipments/Instruments/Apparatus?  In case of Medical Equipments/Instruments/Apparatus, the date of expiry may not be necessary, however, the applicant may mention use life of such device in the Technical dossier. 37. In case the real-time stability study data is not available for the devices at the time of submission of application, whether accelerated stability data can be considered?  The accelerated stability study data can be considered provided the applicant submits the on-going real-time stability study data initiated at the time of accelerated stability study and the detailed real time stability study data shall be submitted after completion of the study to the Licensing authority. Page 9 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 10
Searchable PDF text layer

38. Whether any change in labeling need to be notified to Licensing Authority?  Yes. The changes in respect of label excluding change in font size, font type, color, label design shall be considered as major change as per Sixth Schedule of MDR-2017. 39. Whether GMP compliance and GMP certification is applicable to medical devices as per Medical Devices Rules, 2017?  Under the Medical Devices Rules, 2017, the applicant needs to comply with the requirements of the Quality Management System (QMS) in respect of the manufacturing of Medical devices. 40. Whether WHO GMP certificate is issued for medical devices?  No. The WHO GMP certificate is generally issued for drug formulations based on the WHO guidelines. However, for any certificate required for export purpose, the Licensing authority may consider for issuance of necessary certificate for this purpose under MDR-2017. 41. Who will issue the certificates like Non-Conviction Certificate, Free Sale Certificate, Market Standing certificate etc. which are required on request of procurement / tendering agencies or for export purpose?  The Licensing Authority who has issued license may issue such certificates. 42. What is the validity of Market Standing Certificate issued by the Licensing Authority?  The Market Standing Certificate issued by the Licensing Authority is valid for a period of one year from the date of issuance of the certificate. 43. What is the validity of Free Sale Certificate issued by the Licensing Authority?  Free Sale Certificate issued by the Licensing Authority is valid upto the validity of the manufacturing licence. Page 10 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 11
Searchable PDF text layer

44. Is there any provision for issue of GLP certificate in MDR-2017 for testing laboratory?  There is no provision in MDR-2017 for issuance of GLP certificate. However, the testing lab shall conform to the requirements of Laboratory Quality Management System (LQMS). 45. What is the regulatory expectation to ensure quality of components (raw materials which are to be used for further manufacturing of finished medical devices) under the valid licence for manufacturing?  As regard to ensuring the quality of components/ raw materials to be used for further manufacturing of finished medical devices, it is required that manufacturer of these components need to qualify product standards and Quality Management System. 46. What is the Environmental requirements for manufacturing of medical devices?  The Environmental requirements for certain category of medical devices are prescribed in Annexure A of the Fifth Schedule of MDR-2017. For other medical devices which are not specified in the Annexure A, the manufacturer shall adhere the applicable standards/norms for such manufacturing activities. 47. Whether single Test licence can have multiple testing site?  Yes. If the applicant intends to send the medical device to different testing centers, they may submit single test license application by mentioning each testing site along with the quantity to be tested at each site.  48. What is distinct medical device and Medical Device Grouping?  For this purpose, the Grouping Guidelines for Medical Devices Applications is published by the Ministry of Health & Family Welfare, Government of India which is available in CDSCO website. Page 11 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 12
Searchable PDF text layer

49. If an Investigational medical device is approved by the Central Licensing Authority for export purpose only, whether such devices can be considered as a predicate device?  No. 50. What information is required to be submitted for the substantial equivalence with respect to the predicate device?  The applicant shall submit a comparative statement/document between applied product and predicate device in respect to the name and address of the manufacturer, intended use, risk class, Material of construction, applicable standards, design characteristics, energy used or delivered, manufacturing and testing process, biocompatibility, performance, safety, effectiveness other characteristics etc (as applicable) as per Rule 51 of MDR-2017. 51. Whether the massagers intended for soothing or wellness purpose are regulated under Medical Devices Rules, 2017?  If the massager is intended for soothing or general wellness purpose and not for any therapeutic purpose, then it does not come under the regulation. However, if it is intended for the purpose like therapeutic, alleviation of disease or disorder, etc. are regulated under the provisions of Medical Devices Rules, 2017.  52. What is the mode of submission of application for Free Sale Certificate/ Market Standing Certificate/ Non-Conviction Certificate etc. to the Licensing Authority?  The applicant may submit all such applications through Online mode in Medical Devices online portal (www.cdscomdonline.gov.in) to concerned Licensing Authority. Page 12 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 13
Searchable PDF text layer

53. What is the payment mode for submission of application for the grant of manufacturing license for Class A (other than Class A (Non-sterile and Non- measuring) devices)) and Class B medical devices ?  The fee as specified in Second Schedule of MDR-2017 is payable to the State Licensing Authority, through a challan or by electronic mode as may be specified by the State Government concerned.  54. Whether any exemption on the labeling requirement for Software as a Medical Device (SaMD)/ Standalone software?  No.   55. What is the validity of Import License, Manufacturing License or Registration certificate issued under MDR-2017?  Import License, Manufacturing License or Registration certificate issued under MDR-2017 shall continue to be perpetually valid till suspension or cancellation, provided that the licencee/registration holder shall pay a Licence/ Registration retention fee in every five years under the provisions of MDR- 2017. 56. What is the validity of an Endorsement license?  The endorsement license issued by the Licensing Authority is valid till the date of validity of its base license. 57. Whether all disinfectants are regulated under Medical Devices Rules, 2017?  Only those disinfectants which are intended for the disinfection of Medical devices are regulated under the Medical Devices Rules, 2017. 58. Whether license is required for medical devices that are meant for Research use purposes or quality control testing under Medical Devices Rules, 2017?  The devices which attracts the definition as per S.O. 648(E) dated 11.02.2020 are regulated under the Medical Devices Rules, 2017 and a license is required under the said rule. Page 13 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 14
Searchable PDF text layer

59. Do we need to get test license for prototype medical devices, that are manufactured /Imported for the purpose of techno-commercial feasibility testing, verification & validation testing?  In such cases, the applicant may obtain Test licence in Form MD-13 for manufacturing of test batches or obtain Test licence in Form MD-17 for the import of small quantity of devices under the provisions of Medical Devices Rules, 2017. 60. What is an ‘Accessory’ to a Medical Device?  An ‘Accessory’ means a device which attracts the definition of device as per S.O. 648 (E) dated 11.02.2020.  Such accessory shall be intended specifically by its manufacturer to be used together (in combination) with a particular medical device (parent) to enable or assist that medical device (parent) to be used in accordance with its intended use. 61. What is meant by a component of a Medical device?  Component means any raw material, substance, piece, part, software, firmware, labeling, or assembly which is intended to be included as part of the finished, packaged, and labeled device. 62. What is meant by ‘Spare part’ of a Medical Device?  Spare parts refer to component or part of a medical device, specifically intended to replace an identical or similar integral part or component of a medical device that is defective or worn in order to maintain or restore the function of the medical device without changing its performance or safety characteristics or its intended use and which does not adversely affect the safety and performance of the device.  If such component or part of a medical device, changes the performance or safety characteristics or the intended purpose of the medical device, then it shall be considered to be a medical device and shall meet the requirements laid down the Medical Devices Rules, 2017. Page 14 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 15
Searchable PDF text layer

63. What are the documents required to be submitted for obtaining license for ‘Accessory’ to a Medical Device which is to be marketed individually?  At the time of submission of application, the applicant may follow the Medical Devices Grouping Guideline for requisite fee and submit requisite documents of the main device(s). 64. Which accessories or components needs to be listed under the application for manufacturing/import license?  Accessories or components which are clearly defined in the Device master file as a part of the main device(s) and also in its Instruction for use, are only to be listed under the application for manufacturing/import license. 65. Is there any exemptions for requirement of provisions for Medical devices under MDR-2017?  The Eighth Schedule prescribes the exemptions for certain provisions for Medical devices under MDR-2017.  The Central Government may, by notification, from time to time, amend or modify the entries in the Eighth Schedule. 66. What are the conditions of the License/ permission issued by the Licensing Authority?  The conditions of the License/ permission issued by the Licensing Authority is mentioned in the respective rules under MDR-2017 and also mentioned in the covering letter issued along with the License/ permission. 67. Does Standalone Software require registration/ license?  Yes. Standalone software which attracts the definition of device as per S.O. 648 (E) dated 11.02.2020 are regulated under the Medical Devices Rules, 2017. 68. Whether the software meant for monitoring of fitness, or wellbeing or wellness purpose are regulated?  Software which attracts the definition of device as per S.O. 648 (E) dated 11.02.2020 are regulated under the Medical Devices Rules, 2017. Page 15 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 16
Searchable PDF text layer

CLASS A (NON-STERILE AND NON-MEASURING) MEDICAL DEVICES 69. Whether Class A (non-sterile and non-measuring) medical devices require license to import or manufacturing under Medical Devices Rules, 2017 for marketing in the country?  No. Class A (non-sterile and non-measuring) medical devices are exempted from the requirements of Chapter IV, V, VII, VIII and XI of Medical Devices Rules, 2017.  However, the applicant has to do mandatory registration through Medical Devices Online portal and comply with the labeling requirements prescribed in Chapter VI of MDR-2017 & applicable standards. 70. Whether any Registration certificate is issued by the licensing authority to the applicant after registration of Class A (non-sterile and non-measuring) medical devices ?  A system generated registration number for a Class A (non-sterile and non- measuring) medical device is obtained after furnishing of the requisite information on the Medical Devices online portal. 71. Whether any fees is required for the obtaining registration number for Class A (non-sterile and non-measuring) medical device?  No 72. How to identify medical device having a measuring function?  Any medical device which is intended by the manufacturer to measure quantitatively a physiological or anatomical parameter or measure a quantity or a quantifiable characteristic of energy or of substances (including medicinal products) delivered to or removed from the human body and the result of the measurement is displayed in legal units or other acceptable units of measurement, by the device is considered as a measurable medical device. Page 16 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 17
Searchable PDF text layer

73. Who will issue the Free Sale Certificate/ Non Conviction Certificate/ Market Standing Certificate in respect of Class A (non-sterile and non-measuring) medical device?  As the control of Class A medical devices is under the State Licensing Authority, therefore the applicant may approach to the concerned State Licensing Authority for obtaining such certificates. CLASSIFICATION OF MEDICAL DEVICES 74. What are the criteria for classification of Medical Devices under Medical Devices Rules, 2017?  The parameters for risk based classification for Medical devices are prescribed in Part I of the First Schedule of MDR-2017, as under :— a. low risk - Class A; b. low moderate risk- Class B; c. moderate high risk- Class C; d. high risk- Class D 75. What is the process for risk classification of medical devices prior to submission of application to the licensing authority?  As per Rule 3 of MDR-2017, the Central Licensing Authority classifies medical devices based on their intended use and other parameters specified in the First Schedule of Medical Devices Rules, 2017.  Risk-based classification list of medical devices are published on the website of the Central Drugs Standard Control Organization (CDSCO). The Central Licensing Authority may, from time to time, make additions or deletions in such list of medical devices or modify the class of any medical device.  CDSCO has already displayed the list of medical devices with risk- classification, which is dynamic in nature. Page 17 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 18
Searchable PDF text layer

76. Which Medical Device fall under the category of Class A, Class B, Class C or Class D?  Please refer to the Risk-based classification list issued by Central Licensing Authority available in CDSCO website. 77. Will a list of products classified into Class A, B, C and D be released by CDSCO or the manufacturer has to classify their devices as per risk factors?  Only the Central Licensing Authority under MDR, 2017 is the competent authority to classify the medical devices as per the risk-based parameters specified in the First Schedule of MDR-2017. 78. In case, if the classification of medical device is not included in risk classification published by Central Licensing Authority, then which class will be followed for such medical device?  The applicant may submit separate request to Central Licensing Authority along with the product technical documents (label and IFU), regulatory status of the product / similar product in the other countries to classify the devices and same will be conveyed to the applicant and the existing risk- classification list will be updated accordingly. 79. Is the risk-based classification be harmonized with the already existing and established global classification systems?  Yes. MANUFACTURING OF MEDICAL DEVICES  80. Who is the authority for issuance of manufacturing license for medical devices in the country?  The applicant may submit the application to the Licensing authority through online portal as under: o For Class A (other than non-sterile and non-measuring) medical device & Class B Medical Devices - approach the State Licensing Authority under whose jurisdiction, the manufacturing premises is located. o For Class C & Class D Medical Devices - Central Licensing Authority. Page 18 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 19
Searchable PDF text layer

81. Whether manufacturing site of Medical Device will be inspected before grant of Manufacturing License?  For Indigenous manufacturers of Medical Devices: (i) For Class A (other than non-sterile and non-measuring) medical device, no audit of the manufacturing site shall be necessary prior to grant of licence or loan licence to manufacture for sale or for distribution; and (ii) For Class B, Class C and Class D medical devices, before grant of the manufacturing licence the audit/inspection of the manufacturing site shall be carried out. 82. What is Loan Licence?  Loan licence means a licence issued by the State Licensing Authority or the Central Licensing Authority, for manufacturing a medical device, as the case may be, to a person who intends to utilize the manufacturing site of other licencee for manufacturing the same medical device as manufactured by the licencee at that site. 83. Will the manufacturer have an option to choose Notified body?  No. The Notified body which is registered with the CDSCO for the purpose will be assigned by the concerned State Licensing Authority after review of the online applications. 84. If Notified body is not having competency to evaluate specific class(es) of devices, what would be the process?  In such cases, the notified body shall inform the scope of their audit to the concerned State Licensing Authority inorder to designate the alternate Notified body for the QMS verification. Page 19 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 20
Searchable PDF text layer

85. In case of submission of application of grant of manufacturing license, whether it is mandatory to submit an undertaking by the manufacturer that their manufacturing facility complies with the provisions of the Fifth schedule of MDR-2017?  Yes, the manufacturer shall submit an undertaking signed stating that their manufacturing site is in compliance with the provisions of the Fifth Schedule of MDR-2017. 86. Who approves the Neutral code/ Special code number for export purpose?  The Central Licensing Authority approves the Neutral code/ Special code number for export purpose in respect of all the classes of medical devices. 87. If a manufacturer intends to manufacture medical device only for “Export purpose”, whether they need to specify the same in the application form?  Yes, the applicant shall mention the term “For Export only” under the Generic name of the medical device in the application form. IMPORT OF MEDICAL DEVICES  88. Whether a license is required for import of Medical device for marketing in India?  Yes, for the import of Medical Devices, the applicant has to obtain import license under MDR-2017 from the Central Licensing Authority. However, for Class A (non-sterile and non-measuring) medical devices, the importer has to fulfil the registration requirements to obtain the registration number. 89. What is the procedure to obtain Import license?  The importer shall submit an application in Form MD-14 along with the relevant documents as prescribed in the Fourth schedule (Part I, Part II and Part III (Appendix I & II only)) and requisite fee as per Second schedule of Medical Devices Rules, 2017. Page 20 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 21
Searchable PDF text layer

90. Who can Import a Medical device in the country?  Any firm or organization who is holding valid license to manufacture the devices or holding wholesale licence or holding a registration certificate in Form MD-42 for sale or distribution under Medical Devices Rules, 2017. 91. Who is an “Authorized agent” for import of medical devices in the country?  An “Authorized agent” means a person including any firm or organisation who has been appointed by an overseas manufacturer through a power of attorney to undertake import of medical device in India. 92. What are the changes that require an applicant to obtain a fresh import license?  Fresh import license application shall be made - in case of change in the constitution of a licensee, overseas manufacturer who executed that power of attorney and also, in case of, the change in the location of the actual manufacturer site. 93. What if the classification of a product being imported is different in IMDRF (GHTF) countries from the classification in India?  In such cases, the higher risk class of Medical device may be considered by the Licensing Authority for approval of the devices in the country. 94. Could there be multiple authorized Indian agent for import of the same product(s) for marketing in the country (i.e. same legal & actual manufacturer)?  Yes. 95. In case of multiple importers for same product, do all the documents related to site and product need to be submitted?  The subsequent applicant (new agent) has to submit the documents viz. Form MD-14, Power of Attorney, Free sale certificate, requisite fees, wholesale license/ Form MD-42/ manufacturing license, Label, IFU and copy of import licence issued to earlier authorized agent along with the undertaking from the manufacturer stating that there is no change in the Device master file, Plant master file, under MDR-2017. Page 21 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 22
Searchable PDF text layer

96. Would there be a provision to list multiple sites for a specific product on an existing license?  If the legal manufacturer, having multiple manufacturing sites for manufacturing of a specific product, that may be endorsed in the existing Import license provided a requisite documents and fee are submitted.  97. What is the timeline for grant of Import license in Form MD-15?  As per Medical Devices Rules, 2017, the timeline for grant of Import license is nine months, provided the documents are found satisfactory.  98. Whether it is mandatory that the overseas manufacturing site of Medical device will be inspected before grant of import License?  The Central Licensing Authority may carry out an inspection of the overseas manufacturing site before or after grant of the import licence as and when it is required. 99. Can Importer affix the India specific details as a sticker on retail pack in India or would the manufacturer be required to do so prior to shipping to India?  Importer can provide the India specific label, by way of stickering, when such details are not already printed on the label of the importer devices, which may include import licence number, name and address of the importer, address of the actual manufacturing premises, etc.  100. What fee would be applicable for the manufacturing site, if the importers wish to register devices belonging to multiple risk classes?  As per the second schedule the applicant needs to submit the applicable fee based on the risk class of the devices.   101. Is the Quality Management System as prescribed in Fifth Schedule of MDR-2017 applicable for importers?  The importers need to submit Quality certificate conforming to the Quality Management System as per Fourth Schedule of MDR-2017. Page 22 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 23
Searchable PDF text layer

102. What shall be the content of a Power of Attorney that needs to be submitted in the application for grant of Import license?  An applicant shall submit Power of Attorney along with the undertaking from authorized agent as specified in the Part I of Fourth Schedule of MDR- 2017, duly authenticated in India either by a Magistrate of First Class or by Indian Embassy in the country of origin or by an equivalent authority through apostille, for the grant of Import license. 103. In which form permission to import small quantities of medical devices for personal use can be obtained?  An applicant may apply in Form MD-20 with all requisite documents and permission can be issued by CLA in Form MD-21. 104. Will change in authorized agent require Fresh Import License?  Yes. Change in Indian agent will require Fresh Import License. 105. Are there any requirements for exporting medical devices already imported into the country under Import license in Form MD-15?  If the facility is located in the Special Economic Zone (SEZ) and the firm is export oriented unit, they may export the devices after obtaining necessary permission from the competent authority. 106. Whether Medical Device, having valid Import License, can be imported from any Ports of India?  No. The devices can only be imported through the port notified under the Drug Rules, 1945 from time to time. 107. Is it mandatory for the importers to stock the imported medical devices only at the warehouse address that is mentioned on the import licence?  No, the importer can import the Medical Devices from any notified ports of India and stock for sale and distribution of the devices from any of their registered warehouse in the country. Page 23 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 24
Searchable PDF text layer

108. Whether both legal (If any) and actual manufactures name and address should be stated in the Free Sale Certificate issued by the National Regulatory agency of the exporting country for the purpose of Import of medical devices in India?  Yes. 109. Any changes in name and/or address of Indian agent/ Importer or change in constitution after issue of import licence are required to be communicated to the Licensing Authority?  Yes, Indian authorized agent shall inform such change to CLA in writing within a period of forty-five days in the event of any change in the constitution of the overseas manufacturer or authorized agent. 110. Whether fees required for change in address of authorized agent, without change in constitution as Post Approval Change under MDR-2017 ?  Please refer to the public notice issued vide F. no. 29/Misc/03/2020-DC (124) dated 31.08.2020 issued in this regard. 111. Whether Certificate of Exportability (which reflects that the proposed products may not be freely sold in the country of origin but can be exported), is acceptable as Free Sale Certificate?  No.  INVESTIGATIONAL MEDICAL DEVICES  112. What is an Investigational medical device?  Investigation medical device is a medical device: i. which does not have its predicate device; or ii. which is licensed and claims for new intended use or new population or new material or major design change, and is being assessed for safety or performance or effectiveness in a clinical investigation. Page 24 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 25
Searchable PDF text layer

113. Can a person or sponsor conduct clinical investigation in respect of Investigational medical device in human participants without obtaining any prior permission from the Central Licensing Authority?  No, the person or sponsor shall obtain prior permission from the Central Licensing Authority in Form MD-23 for conduct any clinical investigation in respect of Investigational medical device. 114. What shall be the content of a Clinical Investigation Plan?  The applicant shall submit a Clinical Investigation Plan as prescribed in Table 5 of Seventh Schedule of MDR-2017. 115. What shall be the content of a Clinical Investigation Report?  The applicant shall submit a Clinical Investigation Report as prescribed in Table 10 of Seventh Schedule of MDR-2017. 116. In case of an Investigational medical device, does subsequent applicant need to obtain Investigational medical device approval?  Once an Investigational medical device is approved by the Central Licensing Authority and the product is already marketed in the country, then same can be considered as a predicate device & subsequent applicant may obtain license after complying with the requirements of MDR-2017. 117. To import a medical device which does not have a predicate device, would the clinical investigation be waived off ?  No. However, if such medical device is approved by the regulatory authorities of either the UK or USA or Australia or Canada or Japan and the said device has been marketed for at least two years in that country and if the Central Licensing Authority is satisfied with the data of safety, performance and pharmacovigilance of the device, in such cases the requirement of the Clinical Investigation may be waived off which is decided on case to case basis in consultation with the Subject Expert Committee. Page 25 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 26
Searchable PDF text layer

118. Which Form is required to obtain the Permission for Import or manufacture of Investigational medical device?  The applicant shall submit an application in Form MD-26 to obtain the permission in Form MD-27 under MDR-2017. 119. What documents are required for obtaining permission in Form MD-27 for Medical devices which does not have predicate device?  The applicant is required to submit documents as per Part IV of Fourth Schedule of Medical Devices Rules, 2017. 120. In which Form the License to Import Investigational medical devices by a Government hospital or statutory medical institution for the treatment of patients can be obtained?  An application may be submitted in Form MD-18 with all requisite documents and fee as specified in Second Schedule for grant of Import license in Form MD-19 from Central Licensing Authority. 121. Whether permission in Form MD-23 is required for conduct of academic Clinical study on licensed medical device?  No. However, the Ethics Committee approval needs to be obtained before initiation of the study and the data generated during the study shall not be used to furnish to the Central Licensing Authority to manufacture or to import for marketing any investigational medical device in the country. 122. Whether permission in Form MD-23 is required for conduct of academic Clinical study on Investigational medical device?  Yes. 123. Whether a Permission in Form MD-23 is required for conduct Clinical Investigation on the Medical devices having predicate device?  No, the permission for Clinical Investigation study in Form MD-23 is required for medical devices which falls under the definition of Investigational medical device. Page 26 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 27
Searchable PDF text layer

124. Whether permission fee is required for any institute, organisation, hospital run or funded by the Central Government or the State Government, as the case may be, for conduct of clinical investigation?  No. 125. Whether Clinical Investigation can be initiated after a period of one year from the date of grant of permission in Form MD-23?  If the study is not initiated by the sponsor within a period of one year from the date of grant of permission, they need to take prior permission in Form MD-23 from the Central Licensing Authority. 126. What is the frequency for submission of Periodic Safety Update Report (PSUR) to the Central Licensing Authority?  The permission holder in Form MD-27 shall submit the Periodic Safety Update Report to the Central Licensing Authority from the date of launch of device in the market and such report shall be submitted every six months for first two years followed by submission of the said report annually for the two more successive years. 127. Whether the Pilot Clinical Investigation is mandatory for Investigational medical device developed in India?  Yes, the Pilot Clinical Investigation data generated may be required to be submitted to the Central Licensing Authority and will be useful for designing of Pivotal study. 128. Will clinical investigation be required for any Class A Investigational medical device?  The Clinical investigation for Class A Investigational medical device may not be required, however, where depending on the nature of the medical device, the Central Licensing Authority, may considers such requirement.   Page 27 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 28
Searchable PDF text layer

129. Whether the fee for the grant of permission to import or manufacture a medical device which does not have its predicate device is for distinct product?  Yes. POST APPROVAL CHANGES 130. What are considered to be the major changes and minor changes in Post approval change application for Medical devices?  Certain Major changes and Minor changes are prescribed in the Sixth Schedule of Medical Devices Rules, 2017. 131. Will post-approval change notification approval require submission of fee?  For major changes there may be requirement of fees as per Second Schedule of MDR-2017 however for minor changes no fee is required.  132. What is the provision for change of Model Number or Accessories in already approved medical device?  In such case, the applicant may submit an application under Post approval changes to the Licensing Authority. 133. Is any documentation required including approval certificate from Country of Origin or any other IMDRF countries to include additional Accessory/component under Post approval category?  The applicant may submit the documents as per the checklist for Post approval changes provided in the CDSCO Medical Devices online portal. 134. If there is any change in the location of the manufacturing site approved under the license, then what would be the procedure?  In such cases, the applicant has to inform to the Licensing Authority through Post approval changes (notification), thereafter, necessary approval is required to be obtained from the Licensing Authority as per the requirement. Page 28 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 29
Searchable PDF text layer

135. As software will require updates at certain time over its lifetime, does the importer/manufacturer need to register different versions individually?  For any change in the version of the approved software the importer/manufacturer shall submit an application through Post Approval Change in Medical Devices Online portal. 136. What will be time-period for approval by CLA for implementation of a Major change?  In case of manufacturing the timeline is 45 days and for import it is 60 days. 137. What will be time-period for approval by CLA for implementation of a Minor change?  Implementation of minor change do not need prior approval provided licensee inform CLA within a period of 30 days after the change takes place or becomes effective. RETENTION OF LICENSE/REGISTRATION CERTIFICATE 138. When the applicant is required to submit retention of their endorsement licence?  Once the application for the retention of the base license is submitted by the applicant, they may submit retention application to Licensing Authority for retention of their endorsement license issued under the base license   139. What is the proof of issuance of retention of license in CDSCO Medical Devices online portal?  An acknowledgment email is received from the official email ID of the Licensing Authority. The validity of this retention will be effective from the date of expiry of the original manufacturing/import license. Page 29 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 30
Searchable PDF text layer

140. What is the mode for inclusion of medical device(s) that were deleted/removed by the applicant at the time of submission of their retention application and obtained the retention acknowledgment email copy?  In such cases, the applicant may submit subsequent endorsement application to the Licensing authority.  141. If the application for the retention is submitted by the applicant in due timeline, however, the application is still under process and the applicant did not receive the acknowledgment of their retention application, then can they continue to market their product in the country?  Yes. If the applicant has submitted their Retention application along with the requisite fee, then their license is deemed to be valid and they can market their product in the country.  142. What are the documents required for the submission of Retention application to the Licensing Authority?  The applicant may submit their retention application to the concerned Licensing Authority along with the requisite fee and the documents as per the checklist for retention available on the Medical devices online portal.  SALE  OF MEDICAL DEVICES  143. Whether the wholesale license issued under the Drugs and Cosmetics Rules, 1945 will be applicable for stock and distribution of medical devices under Medical Devices Rules, 2017?  Yes. 144. If the trader/distributor involved only for stock, sale and distribution of medical device only, whether the wholesale license (Form 20B & Form 21B or Form 21C) issued under the Drugs and Cosmetics Rules, 1945 is mandatory?  No. If the trader/distributor is involved exclusively for stock, sale and distribution of medical device only, then they are required to obtain Registration certificate in Form MD-42. Page 30 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 31
Searchable PDF text layer

145. Which authority grants the registration certificate in Form MD-42 to sell, stock, exhibit or offer for sale or distribute a medical device ?  The concerned State Licensing Authority. 146. What is the mode of submission of application in Form MD-41 ?  The applicant may submit an application in Form MD-41 along with requisite documents and fees as per Second Schedule of MDR-2017 through online mode in the Medical Devices online portal to concerned State Licensing Authority. 147. What is the timeline for grant of registration certificate in Form MD-42 ?  The State Licensing Authority may issue Form MD-42 within ten days from the date the application, provided the data found satisfactory. 148. What is the validity period of Registration certificate granted in Form MD-42 ?  Registration certificate issued by the State Licensing Authority in Form MD- 42 is in perpetuity, however, the registration certificate holder shall pay a registration certificate retention fee in every five years under the provisions of MDR- 2017. ***** Page 31 of 31 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 32
Searchable PDF text layer

Central Drugs Standard Control Organization (Medical Devices Division) Medical Devices Frequently Asked Questions (FAQ) Doc No.:CDSCO/FAQ/MD/01/2024 Addendum No.: 01 Date: 03.04.2025 Notice: The replies to the FAQs are aimed only for creating public awareness about Medical Devices Regulation by CDSCO and are not meant to be used for legal or professional purposes. The readers are advised to refer to the statutory provisions of Drugs and Cosmetics Act & Rules and respective Guidelines / Clarifications issued by CDSCO from time to time for all their professional needs. Page 1 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 33
Searchable PDF text layer

Addendum to FAQ on Medical Devices Rules, 2017 1. Whether the manufacturer or importer need to conduct the Biocompatibility test on the all applied products? ➢ The biocompatibility is required for the devices which comes in direct or indirect contact of human body. The manufacturer shall generate the Biocompatibility data on the applied device as per applicable standard. These tests need to be conducted by the manufacturer as a part of the Material validation initially and shall revalidate if there are any changes to the manufacturer of the material or change in the process or supplier. The Biocompatibility testing of medical devices need to be viewed rationally and shall be tested as per IS/ISO 10993 series of standards. ➢ However, if the manufacturer is already carried out the Biocompatibility test on the particular category of the devices and subsequently applied for approval of the similar devices which is being manufactured by using the same manufacturing process, same supplier / vendor of raw materials of the particular devices, in such cases the manufacturer may submit the initial biocompatibility data generated on the particular category of the devices along with the justification & biological risk assessment reports of the applied devices for consideration. ➢ Further, if the material is used for a product with different invasiveness, the additional tests which need to be conducted as per the requirements listed in IS/ISO10993-part 1 standard need to be conducted and validated. ➢ However, the manufacturer shall perform a fore cause testing based on adverse event reported to the them or based on trending result of the complaints received by them. 2. Whether the manufacturer can release the sterile medical device to the market based on parametric release? ➢ As per Clause 7.5.2.2 of Fifth Schedule of Medical Devices Rules, 2017 which prescribes the requirements of Quality Management System for manufacture, storage, installation and distribution of Medical Devices, the manufacturer shall establish documented procedures for the Page 2 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 34
Searchable PDF text layer

validation of the sterilization process. The sterilization process shall be validated prior to initial use. As per Rule 7 of the Medical Devices Rules, 2017, the manufacturers need to comply with the Bureau of Indian Standards (BIS) and in the absence of BIS standards shall comply with ISO / IEC/ Pharmacopoeial standards. If no standards are available, then validated manufacturers standard shall be used. ➢ The BIS has standards for various sterilization validation process viz Ethylene Oxide (IS/ISO 11135), Gamma (IS/ISO 11137-1) and Steam sterilization (IS 17812 (Part 1)/ ISO 17665) ➢ All these standards as a part of routine control and release of sterilized material, allow parametric release. The requirements of the parametric release for each method of sterilization is mentioned in the said standard under routine control and release. ➢ If the manufacturer has validated the sterilization process as per the respective BIS/ISO standards and has plans/records for routine revalidation of sterilization process, then the manufacturer may release the sterilized medical device based on the criteria mentioned in the routine process control requirements of the respective sterilization standards. Page 3 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 35
Searchable PDF text layer

3. Whether for a Medical device which is supplied in non-sterile state, the expiry/shelf life is mandatory on its label? ➢ For the Medical Device which is supplied in non-sterile state, the date of expiry/shelf life on the label of such device may not be necessary. 4. Whether the validation and QC data of medical device generated by the manufacturer for the medical device which is marketed in the country prior to implementation of the mandatory licensing regime under MDR, 2017 can be considered for grant of manufacturing license in the country? ➢ Yes, if the data found satisfactory by the Licensing Authority and for such cases test license in Form MD -13 is not mandatory. 5. In case of Change of constitution of the company/firm holding the manufacturing license under the MDR-2017, whether fresh license is required? ➢ Yes. The licensee shall inform to the Licensing Authority about such change within 45 days and submit an application under MDR-2017 within a period of 180 days from the date of such change in constitution. ➢ In such cases, the existing licence shall be deemed to be valid till such time, a fresh licence is issued or application is rejected by the Licensing Authority. 6. In case of Change of constitution of the company/firm holding the import license under the MDR-2017, whether fresh license is required? ➢ Yes. The licensee shall inform the Central Licensing Authority in writing within a period of 30 days in the event of any change in the constitution of the overseas manufacturer or the authorized agent and submit an application under MDR-2017 within a period of 180 days from the date of such change in constitution. ➢ In such cases, the existing licence shall be deemed to be valid till such time, the fresh licence is issued or application is rejected by the Central Licensing Authority. Page 4 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 36
Searchable PDF text layer

7. In case of Change of constitution of the company/firm holding the registration certificate under the MDR-2017, whether fresh registration is required? ➢ Yes. The registration holder shall inform to the Central Licensing Authority in writing in the event of any change in its constitution and where such change in the constitution takes place, the current registration shall be deemed to be valid for a maximum period of 90 days from the date on which the change took place unless, in the meantime, a fresh approval has been taken from the Central Licensing Authority with the changed constitution. 8. Whether the applicant can submit a single application for multiple actual manufacturing site for obtaining import license? ➢ No. A separate application along with requisite fees and documents need to be submitted. 9. In case of change in the location of manufacturing site of the manufacturer whether a fresh license is required? ➢ Yes. The manufacturer shall obtain fresh manufacturing license under MDR-2017. 10. What is a ‘non-sterile medical device’ ? ➢ A medical device that is intended by the manufacturer to be supplied in a non-sterile state. 11. What are the packaging and labelling requirements for a ‘non-sterile medical device’ if it is intended to be sterilized before use? ➢ If such device is intended to be sterilized before it is used, then the device must be packed in a way that: (a) The label of such device must clearly indicate that the device is in a ‘non-sterile’ state. (b) ensures that the risk of microbial contamination is minimized. (c) is suitable, having regard to the method of sterilization that the Page 5 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 37
Searchable PDF text layer

manufacturer indicates is to be used for the device. (d) the finished product package shall include Instructions for use which clearly mentions the method of sterilization to be used for the device prior to its end-use/use on the patients. 12. What is a ‘sterile medical device’ ? ➢ A medical device that is intended by its manufacturer to be supplied in a sterile state. ➢ The label of such device must clearly indicate that the device is in a ‘sterile’ state along with method of sterilization. ➢ Such device must be designed, produced and sterilized using an appropriate validated method and packed in a way that ensures that the device is sterile when it is supplied, and will remain sterile, until the protective packaging is opened or damaged. 13. Whether wholesale licence in Form 20B/21B under Drug Rules, 1945 or Registration certificate in Form MD-42 under Medical Devices Rules,2017 is mandatory for Class A (non-sterile and non-measuring) medical devices? ➢ No. Class A (non-sterile and non-measuring) medical devices are exempted from the requirements of Chapter XI (Sale of Medical Devices). 14. What is the regulatory pathway to be followed for obtaining manufacturing/ import license for medical device under MDR-2017? ➢ The information on the Regulatory pathway to be followed for the Medical Device from its development to commercialization under Medical Devices Rules, 2017 is available in CDSCO website. (https://cdsco.gov.in/opencms/export/sites/CDSCO_WEB/Pdf- documents/medical-device/RegulatoryMDR-2017.pdf ) 15. Whether the manufacturer of medical device require to obtain ISO 13485:2016 certificate for obtaining manufacturing license under MDR-2017? ➢ No Page 6 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 38
Searchable PDF text layer

16. In case of Import of medical devices, whether Brand name of a medical device owned by an Indian authorized agent can be mentioned on the application for import in Form MD-14? ➢ Brand names (which is /registered under the Trade Marks Act, 1999) for a medical device in Form MD-14 need to be present in the Free Sale Certificate (FSC) issued to the legal manufacturer for indicating it is freely sold in the countries viz. Australia, Canada, Japan, European Union Countries, United Kingdom or the United States of America. In case if the Brand name is not mentioned in the FSC, then the Brand name should be mentioned as ‘Not Applicable’ in the Form MD-14. ***** Page 7 of 7 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 39
Searchable PDF text layer

Central Drugs Standard Control Organization (Medical Devices Division) Medical Devices Frequently Asked Questions (FAQ) Doc No.:CDSCO/FAQ/MD/01/2024 Addendum No.: 02 Date: 09.07.2025 Notice: The replies to the FAQs are aimed only for creating public awareness about Medical Devices Regulation by CDSCO and are not meant to be used for legal or professional purposes. The readers are advised to refer to the statutory provisions of Drugs and Cosmetics Act & Rules and respective Guidelines / Clarifications issued by CDSCO from time to time for all their professional needs. Page 1 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 40
Searchable PDF text layer

Addendum to FAQ on Medical Devices Rules, 2017 1. What are the requirements for re-import of medical device in the country? ➢ The requirement for re-import of the medical device is prescribed in the Guidance document for functions and responsibilities of Zonal, Sub-zonal & Port offices of CDSCO, 2024. ➢ The decision on the requirement for testing of such consignment at the lab before re-import into the country and its release may be taken by the Port office in consultation with concerned Zonal office of CDSCO/ State Drugs Control office, as appropriate. 2. What is the requirement for carrying out outsourced activity (e.g. Coating, Laser Engraving, batch testing, etc) by the manufacturer at an overseas site? ➢ The manufacturer needs to obtain a No objection certificate (NOC) from the concerned Licensing Authority for carrying out such activity at the overseas premises, by submitting relevant documents viz. copy of manufacturing license, the name and quantity of the medical device to be sent for such activity, copy of the agreement with the overseas firm to carry out such activity, etc. 3. Whether change from Partnership firm to LLP firm, or vice versa, is also considered as change in constitution? ➢ Yes. In such cases the applicant may obtain fresh license under the MDR- 2017. 4. Whether change from firm to company, as defined in the MDR-2017, or vice versa, is also considered as change in constitution? ➢ Yes. In such cases the applicant may obtain fresh license under the MDR- 2017. 5. Whether approval is required from Licensing authority for endorsement/change of competent Technical staff for manufacturing or testing of medical devices under MDR-2017? ➢ No. The manufacturer shall appoint the competent Technical staff for manufacturing or testing of medical devices as per Rule 22 of MDR-2017. Page 2 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 41
Searchable PDF text layer

6. Whether the product manufactured in the premises located at Special Economic Zone (SEZ) can be diverted to Domestic Tariff Area (DTA) for sale? ➢ For import of medical devices in India, an Import license is required to be obtained under Chapter V of Medical Devices Rules, 2017. ➢ For the medical devices manufactured in the premises located at Special Economic Zone (SEZ) and which need to be diverted to Domestic Tariff Area (DTA) for sale, by the manufacturer, the following procedure should be followed for transfer of the medical device to DTA to ensure that the devices shall meet the quality, safety and performance requirement as per the Drug & Cosmetics Act, 1940 and rules made thereunder: • In case, the device manufactured in the SEZ, the manufacturer shall comply with the requirement specified under Chapter IV of MDR-2017 for obtaining manufacturing license. • If the critical component of the medical device which attract medical device definition under the Rule, is imported for manufacturing of Finished Device which is manufactured in SEZ to be diverted to DTA for sale and distribution, the applicant shall comply with the requirement specified under Chapter V of MDR-2017 for obtaining import license for such component. • Further, the applicant may also refer to the Circular issued by CDSCO vide IMP/141/024-eoffice dated 08.04.2025 in this regard. 7. For regulation of medical devices whether the provisions of Drug Rules, 1945 is applicable? ➢ As per Rule 96 Overriding effect, the provisions of Medical Devices Rules, 2017 shall have effect, notwithstanding anything inconsistent therewith contained in the Drug Rules, 1945. ➢ However, if any specific provisions under the Drug Rules, 1945 for which there is no provision under the MDR-2017, the same may be applicable for Medical devices also. 8. What is the provision for inclusion of additional brand name in the existing license? ➢ The applicant shall apply separately as per exclusive provision viz. ‘Brand approval’ made under the Medical Devices Online portal. Page 3 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 42
Searchable PDF text layer

9. Whether a single use medical device claimed by the manufacturer can be reused? ➢ No, as such medical device is assessed by the manufacturer for its safety and performance for single use only. 10. Can multiple models comprising of different intended use, material of construction, design characteristics, etc may be considered under a same medical device grouping category? ➢ No, products/models having different intended use, material of construction, design characteristics, etc which are beyond the permissible limits cannot be applied under a single medical device category. The manufacturer has to apply separately for the same. 11. Whether the stickering activity is allowed to the importer at the licensed/registered warehouse? ➢ Yes, in case of import, the license holder may carry out the stickering of India specific information as provided in Rule 44(n) of MDR-2017 on label of medical device. 12. Is it mandatory to perform a design qualification test for every batch released for a particular medical device having same material composition, manufacturing process, etc. ➢ It is not mandatory to perform a full Design Qualification test for every batch, provided, the material of composition of the device and its manufacturing process remain same and the validation of the manufacturing process (such as Process Validation and periodic requalification) is carried out to ensure the product's safety and effectiveness. ➢ However, if there are significant changes to the design, material composition, or manufacturing process, a requalification or additional testing might be required. Page 4 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 43
Searchable PDF text layer

13. Whether the Standard prescribed under Rule 7 of MDR-2017 is mandatory for medical devices manufactured exclusively for export purpose? ➢ The manufacturer of the medical device in India shall obtain manufacturing license under the provisions of Chapter IV of MDR-2017 to manufacture medical devices that meets the standards as prescribed under said rules. ➢ Further, in case of export, the manufacturer may follow the applicable standard as per the requirement of the importing country. 14. What information needs to be mentioned in the specimen/draft label in case of the import of a medical device for marketing in the country? ➢ The specimen label of the imported device shall conform to the requirement as prescribed under Rule 44 of MDR-2017. The label should also provide the specific space for stickering of India specific label in case if it is to be stickered in India. 15. What information needs to be mentioned in the specimen/draft label in case of the manufacturing of a medical device for marketing in the country ? ➢ The specimen label of the device to be manufactured shall conform to the requirement as prescribed under Rule 44 of MDR-2017. 16. In case of multiple Bharatkosh fee receipts submitted in an application, do all Bharatkosh fee receipts need to be linked with the application? ➢ Yes. Each Bharatkosh fee receipts needs to be linked with the application if there are multiple challans so that total fee details shall be freezed with the application on the portal. 17. What are the requirements of the checklist for obtaining various types of approvals under MDR-2017? ➢ The checklist of the various types of application forms for obtaining approvals is available on the website of CDSCO (https://cdsco.gov.in/) / MD online portal (https://cdscomdonline.gov.in/) /NSWS (https://www.nsws.gov.in/). Page 5 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 44
Searchable PDF text layer

18. How to fill in the application form (legal form) while submitting applications for obtaining approval under MDR-2017? ➢ For submitting specific information in the application form (legal form) a requisite tooltip has been provided at the applicant end on the portal so that the applicant may be aware on specific information to be filled in the legal form. 19. Whether Free Sale Certificate from any of the following countries viz. USA, UK, EU, Canada, Japan or Australia is required for obtaining Import license for Class A & Class B ? ➢ Yes. However, incase if it is not approved by any of the said authority the applicant may submit published safety and performance data or clinical investigation report generated in the country of origin and a free sale certificate from the country of origin or any other country where the device is being marketed. 20. Whether Free Sale Certificate from any of the following countries viz. USA, UK, EU, Canada, Japan or Australia is required for obtaining Import license for Class C & Class D ? ➢ Yes. However, incase if it is not approved by any of the said authority the applicant may submit clinical investigation report generated in India and a free sale certificate from the country of origin or any other country where the device is being marketed. 21. Is there any requirement for obtaining test license for testing of a custom-made medical device? ➢ No 22. Whether the devices intended for human beings for its intended purpose, can also be used for veterinary purposes? ➢ No. The device intended for veterinary use shall be licensed under the MDR- 2017 by the Licensing Authority separately for veterinary purpose. Page 6 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
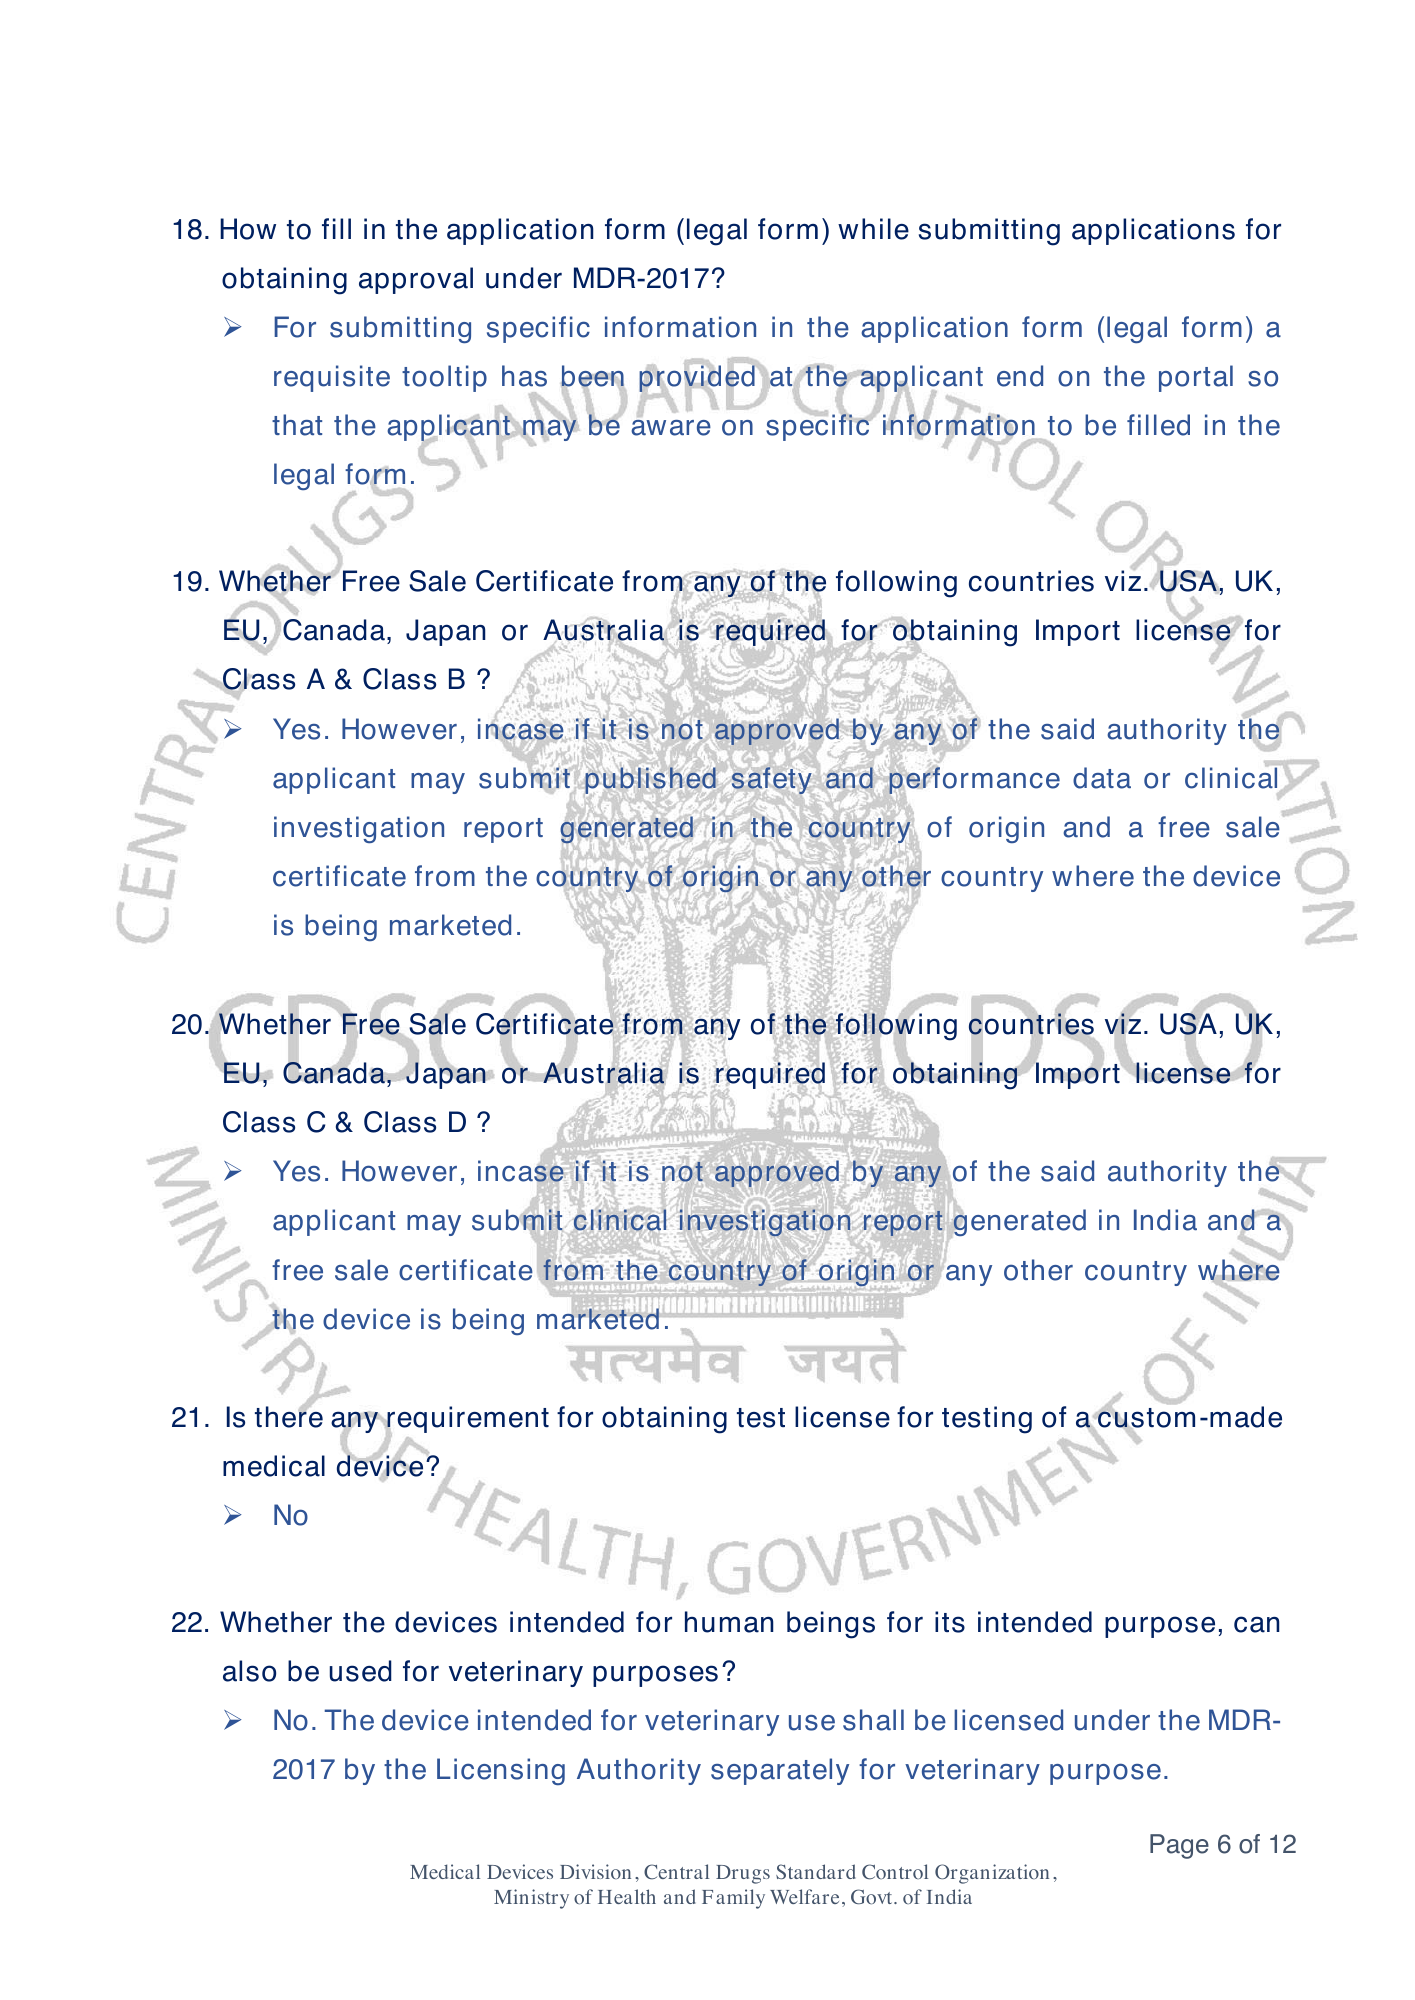
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 45
Searchable PDF text layer

23. What is the residual shelf life required for a medical device for its import into the country for marketing? ➢ The residual shelf-life of a medical device is prescribed under Rule 47 of Medical Devices Rules, 2017 as under: Total shelf life claim of the medical Residual shelf life of the device as per its manufacturer medical device on the date of import (a) Less than ninety days Not Less than 40% (b) Between ninety days to one year Not Less than 50% (c) more than one year Not Less than 60% 24. What is a custom-made medical device? ➢ As per the definition prescribed under Rule 3(r) of MDR-2017, "custom made medical device" means a medical device made specifically in accordance with a written prescription of a registered medical practitioner, specialised in the relevant area, under his responsibility for the sole use of a particular patient, but does not include a mass production of such device. 25. Whether license is required under MDR-2017 for manufacture/import of Custom-made medical Devices for marketing in the country? ➢ The custom-made medical device is exempted for the provisions of Chapter IV and Chapter V of MDR-2017. However, the facility of such devices shall conform to the requirement of the Quality management system and the labelling provisions under the said rule. 26. Whether Custom-made medical device manufactured/imported in bulk also exempted from the provisions of Chapter IV and Chapter V of MDR-2017? ➢ No. Mass produced devices, which only need adoption to meet the specific requirement of a medical practitioner or any other professional user, shall not be considered as custom made device and license is required for such devices for marketing in the country. Page 7 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
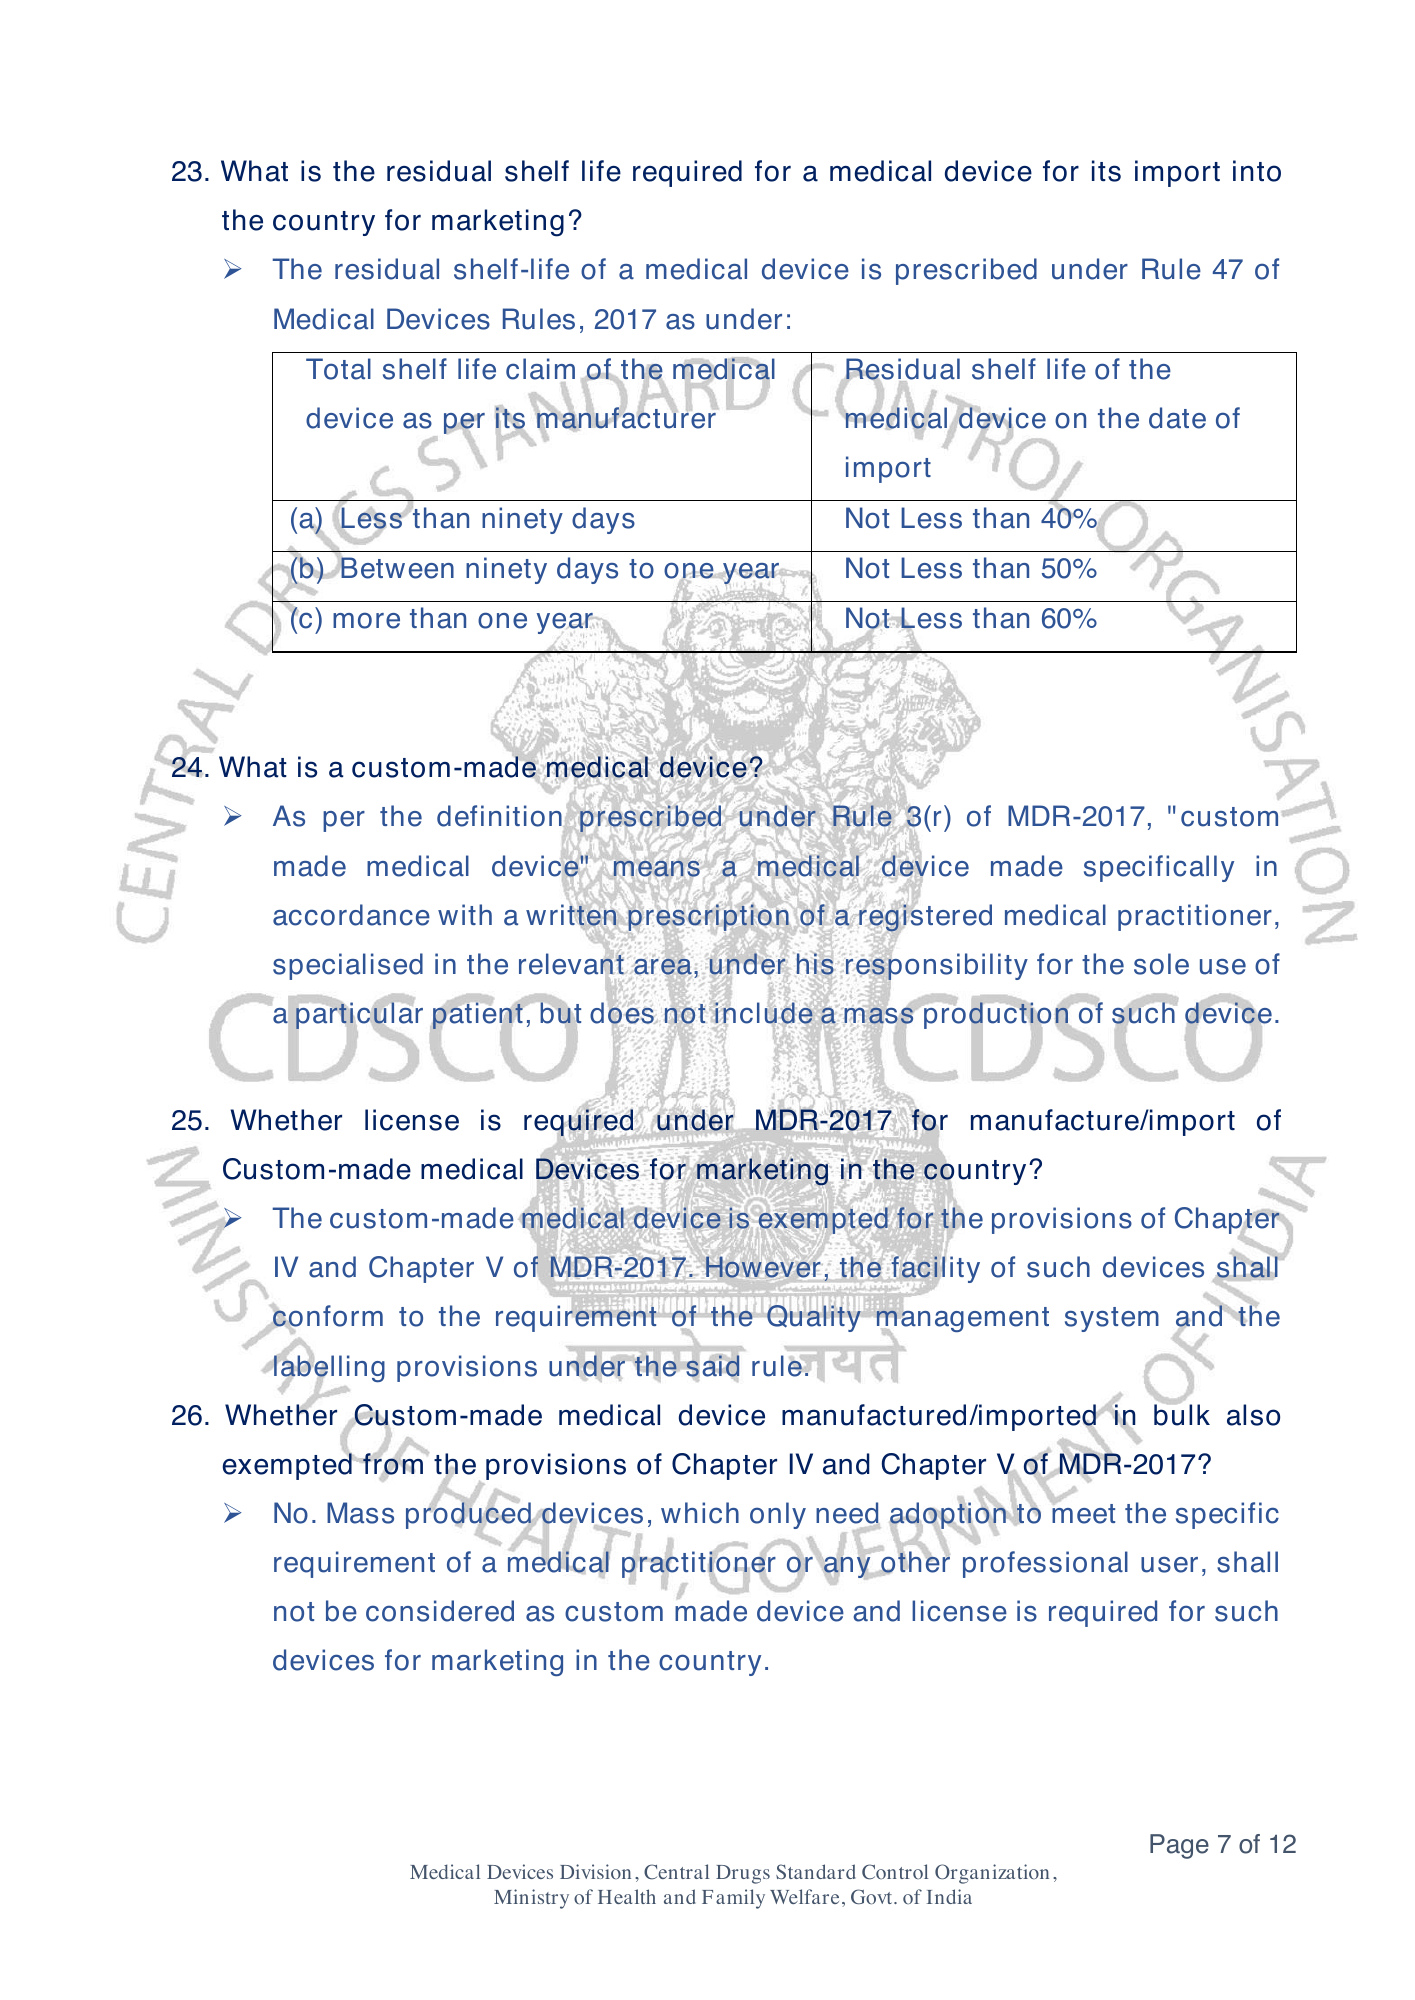
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 46
Searchable PDF text layer

27. Whether the veterinary medical devices can be approved if it is not approved by National regulatory authority of countries viz. USA, UK, EU, Canada, Japan or Australia? ➢ Incase if the veterinary medical device is not approved by above authorities, the applicant may submit approval obtained from other competent authority from the said countries for its marketing. 28. Whether NOC from Department of Animal Husbandry and Dairying (DAHD) is required to be obtained for all medical devices intended for veterinary purpose prior to grant of license? ➢ The NOC of DAHD is required only for the veterinary medical devices which does not have predicate device. 29. What are the documents required to be submitted for obtaining NOC from Central Licensing Authority for import of small quantity of medical devices for donation to a charitable hospital for treatment of patients free of cost by that hospital? ➢ The applicant may submit following documents for obtaining NOC from Central Licensing Authority: • A copy of registration document of Charitable hospital/trust • Bill of Entry or Bill of Landing/ Commercial Invoice • Undertaking from charitable hospital stating that the devices will be donated free of cost to the patients. • Undertaking stating that the Charitable hospital will take the responsibility of any safety issues related to the device to be used on the patient. 30. What is the requirement for becoming an Indian-authorized agent for the import of medical devices in the country? ➢ The Indian-authorized agent shall possess a valid wholesale license obtained in Form 20B and Form 21B under the Drug Rules, 1945 or a Registration Certificate obtained in Form MD-42 under the Medical Devices Rules, 2017 or manufacturing license obtained under Medical Devices Rules, 2017. Page 8 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
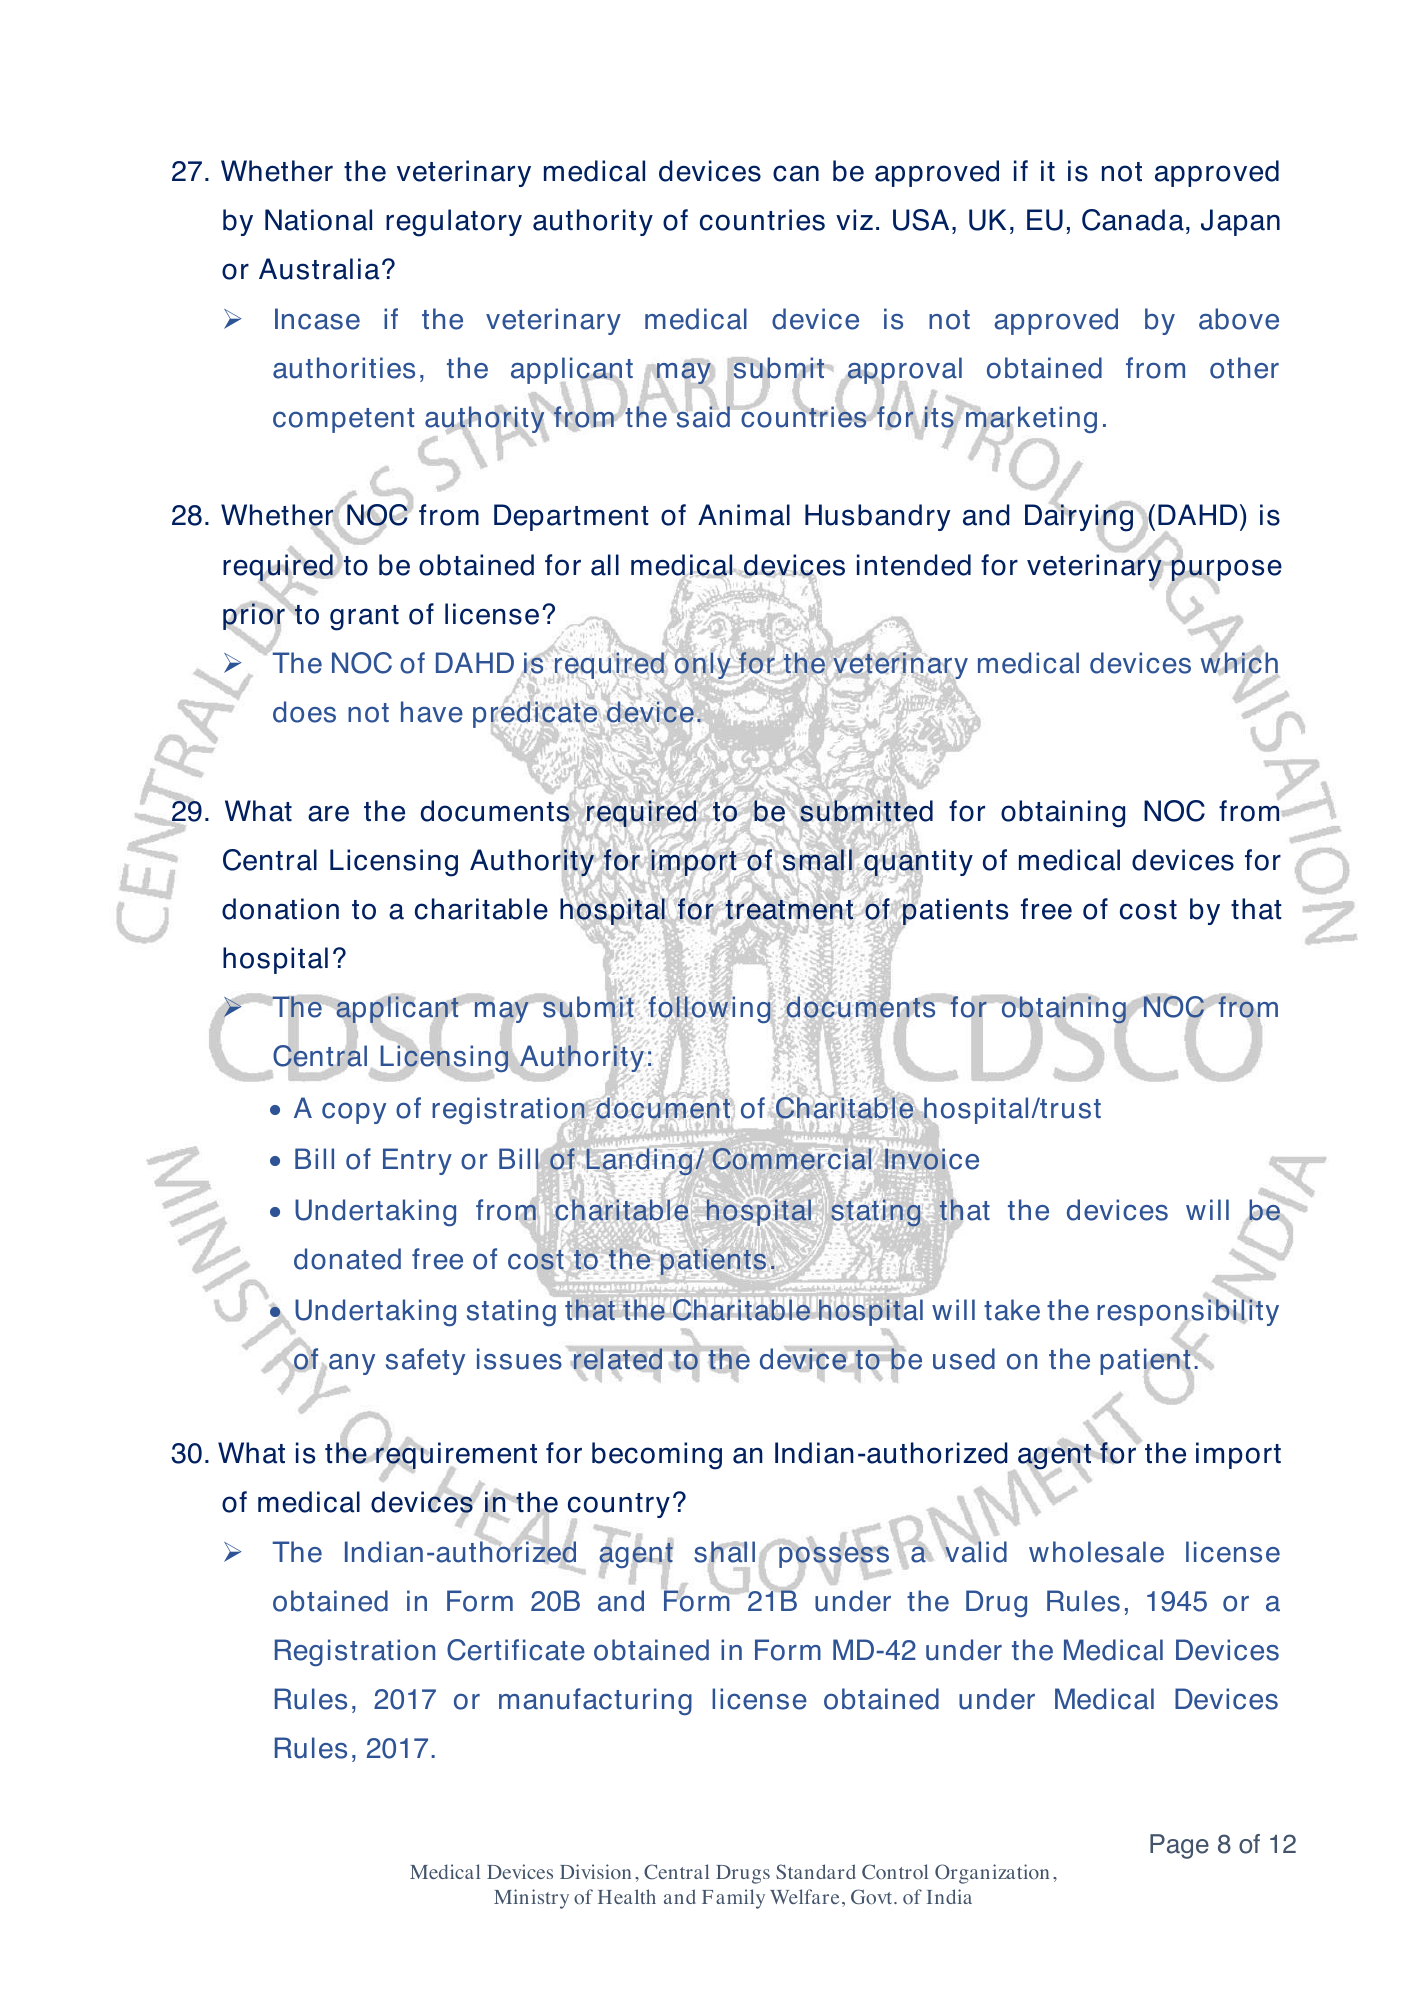
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 47
Searchable PDF text layer

31. Whether the requirement of Quality Management system is applicable in case of Class A (non-sterile and non-measuring) medical devices? ➢ For manufacturing of such medical devices, the manufacturing facility shall conform to the requirements of Quality Management system for medical devices. 32. Whether all gloves are covered under the Medical Devices Rules, 2017? ➢ No. Only the Surgical Gloves, Medical Examination gloves and other gloves which are intended for medical purposes, are regulated under the Medical Devices Rules 2017. 33. Whether import license is required under the Medical Devices Rules, 2017 for import of small quantity of medical devices for donation to a charitable hospital for treatment of patients free of cost by that hospital? ➢ No. ➢ In such case, the provisions of Chapter V of MDR-2017 are exempted, the applicant shall obtain No Objection Certificate for such purpose from the Central Licensing Authority. 34. What is the regulatory requirements for outsourcing sterilization activity of medical devices by the manufacturer under MDR-2017? ➢ Please refer to CDSCO Letter no. MED/48/2025-eoffice dated 24.06.2025 published in CDSCO website. 35. Whether the components/accessories of the Medical device require license under MDR-2017? ➢ The ‘Component’ or ‘Accessory’ of a parent medical device which attracts the definition of ‘medical device’ as per S.O. 648 (E) dated 11.02.2020 and which is packed or labelled for the commercial distribution, require license under MDR-2017. Page 9 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
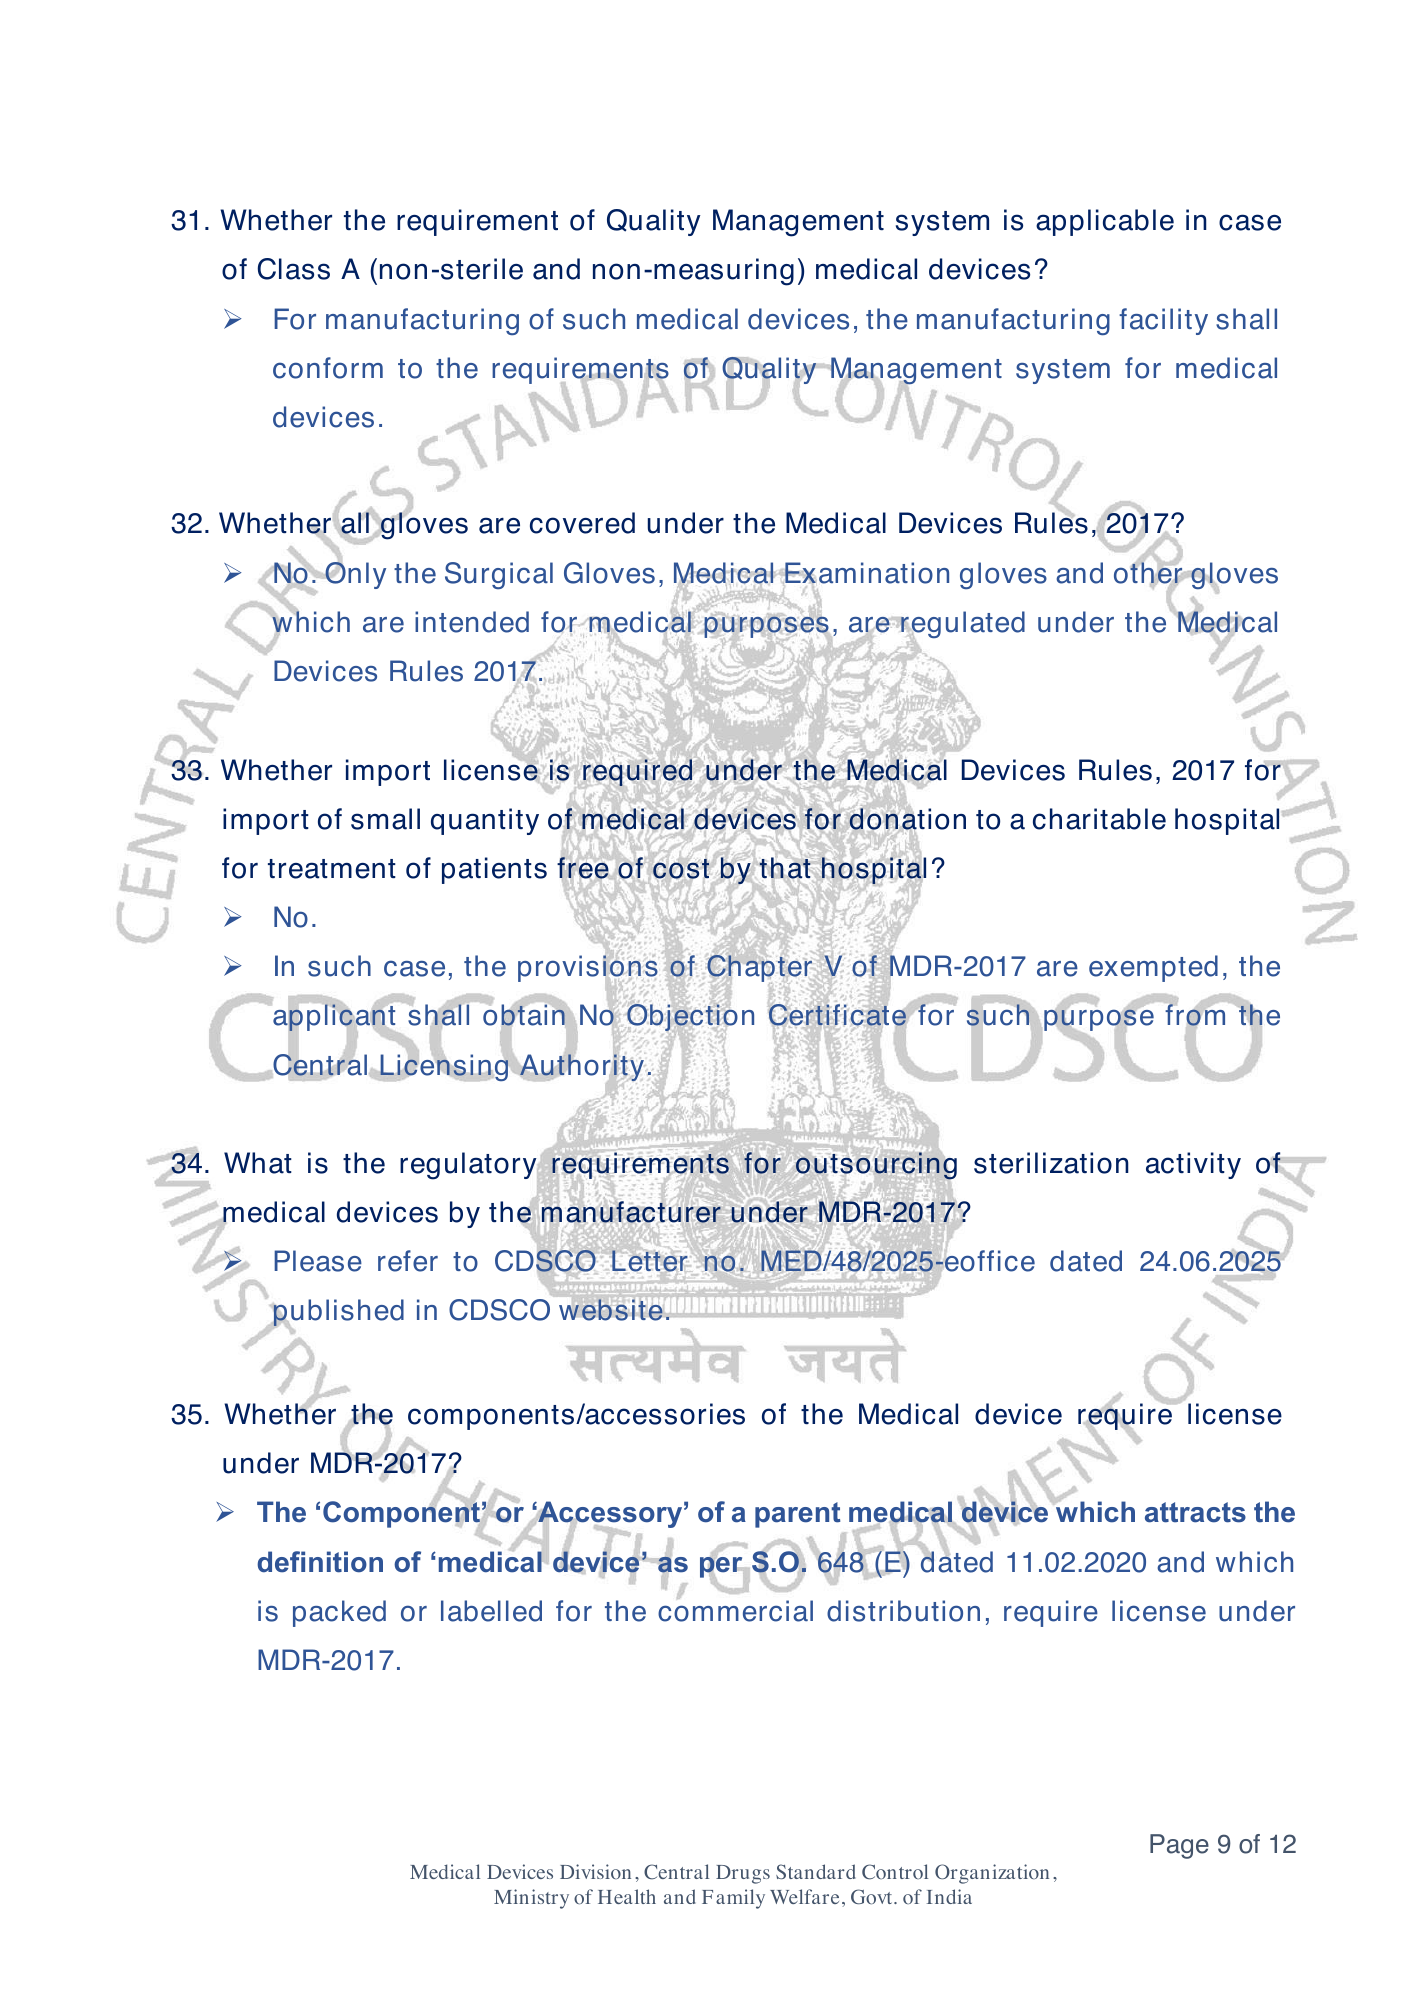
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 48
Searchable PDF text layer

36. Whether change in the method of sterilization of medical device (eg. from Gamma to ETO sterilization, etc) is considered as a separate device ? ➢ Yes. In such cases the applicant need to obtain an endorsement to their license obtained under the MDR-2017. 37. Whether the intended use of a medical device as mentioned in the CDSCO classification list is to be adhered by the applicant while submitting application for obtaining manufacturing/import license for marketing in the country? ➢ No, the intended use of the medical device in the application for manufacturing/import may not be exactly the same as that of the intended use of the device mentioned in the CDSCO classification list. Further the essence of the intended use of the particular device should not deviate from that mentioned in the CDSCO classification list. Otherwise, the risk class of the device may change based on the intended use. 38. Whether the medical devices manufactured or imported under the Test license in Form MD-13 or Form MD-17 can be used for commercialization in the country? ➢ No. The medical devices manufactured or imported under Test license shall shall be used exclusively for purposes of clinical investigations/ test/ evaluation/ demonstration/ training and it shall not be commercialized. The unused quantity may be either exported (incase of import) or destroyed under the intimation to the Central Licensing Authority. Commercialization of devices manufactured/imported under Test license may attract penalty as per the Drugs & Cosmetics Act, 1940. Page 10 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 49
Searchable PDF text layer

39. Whether the risk-class of a medical device, as mentioned in the test license (MD-13/MD-17), can be considered for market authorization of the medical device in the country? ➢ No, the risk-class mentioned in the Test license is the proposed risk classification of the applied medical device as applied by the applicant at the stage of development of the product. ➢ The applicant(s) may ensure whether the medical device, for which application is to be submitted for obtaining market authorization, is enlisted in the risk classification lists published by the CLA. In case if the said medical device is not enlisted in the published risk classification lists, they shall seek clarification from the CLA regarding its risk classification 40. What is the process for obtaining Neural code for medical device under Medical Devices Rules, 2017? ➢ The manufacturer who obtained license for medical device for marketing in the country may obtain system auto-generated Neutral code by applying through the Medical devices online portal (https://cdscomdonline.gov.in/) from the Central Licensing Authority. 41. What is the process of obtaining Market Standing Certificate/ Non-Conviction Certificate for Class C & D medical devices? ➢ The applicant may submit an application through Medical devices online portal (https://cdscomdonline.gov.in/) to obtain system auto-generated Market Standing Certificate/ Non-Conviction Certificate. 42. What are the requirements for registering a testing laboratory under the Medical Devices Rules, 2017 for testing of medical devices on behalf of the manufacturer? ➢ The Chapter X of the Medical Devices Rules, 2017 prescribes the procedure for obtaining registration certificate for such Medical Device Testing laboratory. Further the testing facility shall be NABL accredited and conform to the Laboratory Quality Management System as per ISO/IEC 17025. Page 11 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 50
Searchable PDF text layer

43. What is the mode of payment of fees for obtaining license under MDR-2017? ➢ The applicant shall pay the fees to concerned Licensing Authority through a challan or by electronic mode as specified under MDR-2017. Further it is advisable that the applicant may upload the receipt of the payment as received after making the payment through challan. 44. Whether all Beauty care equipments for human use are regulated under Medical Devices Rules, 2017? ➢ No, provided, the Beauty care equipment is not intended for any therapeutic purpose, etc or modification of anatomy/physiology of the human body. 45. Whether Batch release certificate/ Certificate of Analysis of consecutive three batches is required for Software as a Medical Device (SaMD) and Medical equipments? ➢ For such devices the applicant may submit final software release certificate/factory release certificate indicating the product meets the applicable specifications, wherever required, along with proper justification with respect to the requirement. 46. Whether a fresh application for obtaining Registration certificate for Notified body responsible for audit of class A & B devices can be obtained through MD online portal? ➢ No. The applicant shall submit an application in Form MD-1 through NSWS only (https://www.nsws.gov.in/). 47. What is the procedure for waiver of the additional condition mentioned in the permission/license granted for Class C & D medical devices under MDR-2017? ➢ The applicant may submit an application through the provision made in the MD online portal. ***** Page 12 of 12 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 51
Searchable PDF text layer

Central Drugs Standard Control Organization (Medical Devices Division) Medical Devices Frequently Asked Questions (FAQ) Doc No.:CDSCO/FAQ/MD/01/2024 Addendum No.: 03 Date: 03.11.2025 Notice: The replies to the FAQs are aimed only for creating public awareness about Medical Devices Regulation by CDSCO and are not meant to be used for legal or professional purposes. The readers are advised to refer to the statutory provisions of Drugs and Cosmetics Act & Rules and respective Guidelines / Clarifications issued by CDSCO from time to time for all their professional needs. Page 1 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 52
Searchable PDF text layer

Addendum to FAQ on Medical Devices Rules, 2017 1. What are the risk classification criteria for medical devices intended to be used on animals?  Risk parameters for classification devices used in animal are the same as per prescribed under First schedule of MDR-2017 and same risk class as mentioned in the risk classification notice is applicable. 2. In case if a Medical Device intended for animal use, is not regulated in the country of origin, not approved by any other stringent regulatory authority and facility not audited for Quality Management System (QMS) as per ISO 13485; what are the documents to be submitted with the import application?  In such case, the applicant can submit the any other relevant certificates issued by the competent authority, confirming the Quality Management System in the manufacturer premises. 3. Whether the Power of Attorney (PoA) signed by the manufacturer and undertaking signed by the authorised agent and thereafter apostille or authenticate together either in country of origin or in India; need to be a single binded/punched document?  Yes. 4. In case of change in Actual manufacturing site name, without change in constitution, whether fresh license is required?  No, in such case, the firm need to obtain approval from the licensing authority through Post Approval Change. 5. For an authorised agent having multiple retail or wholesale license under Drug Rules, 1945 or registration certificate (From MD-42) under MDR-2017, whether address of all such premises licensed/registered for sell or stock for sale or distribute need to be mentioned as authorised agent address in the import license obtained under MDR 2017?  No. Page 2 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 53
Searchable PDF text layer

6. Who will be considered as Subsequent importer for import of medical device in the country?  An authorised agent of the foreign manufacturer who intend to obtain the import license for import of medical device(s) which is already licensed to another agent/importer under MDR-2017, provided the legal and actual manufacturing site of the device is the same. 7. Whether the AERB type approval or compliance certificate, is a mandatory requirement prior to obtain license for import or manufacture of ionization radiation medical devices for marketing in the country?  No. However the applicant shall submit AERB approvals to the licensing authority, prior to use of such device on Indian population. 8. Whether an import license is required to be obtained for importing raw materials/components to be used for further manufacturing of finished Single- Use medical devices under a valid manufacturing license issued under the provisions of D&C Act and Rules made thereunder?  No. However, such raw materials/components need to qualify applicable standards and Quality Management System. 9. Whether an import license is required to be obtained for importing components/parts to be used for further manufacturing/assembling of finished Medical Equipment/instrument under a valid manufacturing license issued under the provisions of D&C Act and Rules made thereunder?  As per the existing practice, in case if such component or part which is to be imported is critical in nature which attracts the definition of medical device and affects the performance or safety characteristics or the intended purpose of the medical device, then import license is required. Page 3 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 54
Searchable PDF text layer

10. Whether import license required to be obtained for importing unfinished medical devices in bulk for carrying out further manufacturing activity viz. sterilization, labelling and packaging to make finished Medical device under a valid manufacturing license issued under the provisions of D&C Act and Rules made thereunder?  Yes. 11. Whether an Investigational medical device can be imported by any hospital for treatment of patient?  Only a Government hospital or a statutory medical institution allowed to import small quantity of investigational medical device which is approved in the country of origin, for treatment of a patient suffering from a life threatening disease or disease causing serious permanent disability or disease requiring therapy for unmet medical need. Application in Form MD-18 need to be submitted through Online System for Medical Devices. www.cdscomdonline.gov.in/NewMedDev/ViewDevicePersonal 12. Whether the hospital can also import medical devices other than investigational medical devices in Form MD-21 for treatment of patient?  No. 13. Whether the products (eg: clinical mannequin etc) intended to be used for the purpose of education (not for planning or practicing clinical procedures with actual patient data) only by healthcare professionals, academician etc. comes under the purview of MDR-2017?  No. 14. Whether Radiopharmaceutical products are covered under the MDR-2017?  No. For import/manufacture of radiopharmaceutical products, the applicant may obtain license under Drug Rules,1945 of Drug & Cosmetics Act, 1940. Page 4 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 55
Searchable PDF text layer

15. In case of only “change in constitution” of the firm, does the manufacturing premises require QMS re-inspection/audit?  Not required, however the authority may verify if there is any change in the QMS requirement. 16. In case of only “Change in constitution” of the Medical Devices Testing Laboratory registered with Central Licensing Authority, does the laboratory need to be re-inspected as per MDR-2017 requirement?  Not required, however the authority may verify if there is any change in the laboratory QMS requirement. 17. In case of change in location of a manufacturing site in India, whether QMS re- inspection/audit is mandatory under MDR-2017?  Yes. 18. Whether products such as laminar flow safety cabinet, industrial sterilizer, microscope, protective coveralls etc. which is intended for non-medical purposes in production unit or testing or R&D of various sectors/industry like food technology, pharmaceutical or biotechnology or chemical or electronic, etc. comes under the purview of MDR-2017?  No. 19. What is the regulatory pathway to be followed for obtaining license for marketing an Investigational medical device (i.e. Device for which predicate device is not available in the country)?  In such cases the applicant may follow the pathways as under: The applicant may initially obtain Test license in Form MD-13/MD-17 (for purpose of testing or clinical investigation, as the case may be) to manufacture/import test batches for generation of validation data, etc. Thereafter, apply in Form MD-26 along with the requisite fees and documents including clinical evidence data generated on the device to obtain Form MD-27 prior to apply for license to manufacture or import in relevant forms from the concerned licensing authority. In case no clinical data or inadequate Page 5 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 56
Searchable PDF text layer

data on safety and performance is generated on the device, the applicant may conduct the study after obtaining permission in Form MD-23 and submit the same for consideration of their application for marketing in the country. 20. What are the extent and conditions of exemption for Custom-made medical devices?  Mass produced devices, which only need adaption to meet the specific requirement of a medical practitioner or any other professional user, shall not be considered as Custom-made device. All provisions of Chapter IV and Chapter V of MDR-2017 are exempted, subject to the condition that the device is being specifically made in accordance with a duly qualified medical practitioner's written prescription under his responsibility, in accordance with specific design, characteristics and the same is intended for the sole use of a particular patient and the label contains the words 'Custom made device'. Only change in dimension of devices cannot be considered under custom medical devices. 21. Whether medical devices with different dimensions only are considered as Custom-made device as prescribed by the registered medical practitioner's?  No. 22. Whether clinical investigation permission in Form MD-23 is required for Class A (non-sterile and non-measuring) medical device?  No. Page 6 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 57
Searchable PDF text layer

23. How to obtain permission to import small quantity of medical devices for personal use by the patient?  The patient or his nominee, may apply in Form MD-20 along with documents and medical prescriptions to the Central licensing authority, through Online System for Medical Devices for obtaining the permission in Form MD-21. www.cdscomdonline.gov.in/NewMedDev/ViewDevicePersonal 24. Whether the import or manufacturing license holder need to submit endorsement application to add model name or model number of an approved device?  In cases where the model name or number added is in consonance risk class, intended use and technical characteristics as mentioned in the current device master file, the applicant need to submit Post approval change application for its addition. 25. What procedure need to follow for approval, if the medical device is licensed as a ‘System’ as per grouping guideline and license holder intended to include additional model of the said system?  In such cases, the applicant may obtain post approval changes in respect of grouping from ‘System’ to ‘Family’ and addition of model name/number 26. How to delete or modify an existing model name or model number of device from the license?  The applicant needs to submit an application under Post approval changes for change in model name or model number of the licensed device. Page 7 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 58
Searchable PDF text layer

27. What is the procedure to be followed by license holder to inform licensing authority on any change or anticipated change in approved medical device or manufacturing site?  Any changes on the device or site licensed by the licensing authority require submission of post approval change application to concerned licensing authority under the respective category through Online systems for medical devices. 28. Whether applicant need to submit fee for change in address of authorized agent without change in constitution as post approval changes under MDR 2017?  No. The applicant may refer CDSCO notice vide F.No. 29/Misc/03/2020- DC(124) dated 31.08.2020. 29. What is the procedure to change registered office address/correspondence address of a licensee in Online System for Medical Devices?  Applicant may write to IT cell of CDSCO-HQ through CRU (cru.division@cdsco.nic.in) for making changes in registered office address/correspondence address in Online System for Medical Devices. 30. Incase of change in constitution of the firm , whether the license holder need to create separate login credential in CDSCO Medical devices online portal to obtain the fresh license?  Not mandatory, in such cases the license holder may submit a separate request to IT cell, CDSCO for necessary changes in the existing login credential along with relevant documents and updated credentials may be used for submitting fresh application. However, if they wish to create a separate credential for obtaining the fresh license, the existing credentials may be considered deactivated and the same need to be informed to IT cell-registration desk for necessary action to deactivate the credential. Page 8 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India
Frequently Asked Questions on Medical Devices Rules 2017 — official portal compilation — official PDF physical page 59
Searchable PDF text layer

31. Whether registered Class A (non-sterile and non-measuring) medical device under MDR-2017, can be imported from any Ports of India?  No. The devices can only be imported through the port notified under the Drug Rules, 1945 from time to time. 32. Can the State/UT Licensing Authority who initially referred the application to Notified body for QMS audit and want to transfer the application to other Notified body due to certain reasons?  Yes. The State/UT Licensing Authority can transfer the application file to another Notified body through their concerned Nodal officer to another Notified body for the conduct of QMS audit. 33. What is the mode of communication between the State/UT Licensing Authority and the Notified body in the Online System for Medical Devices with respect to QMS audit?  The communication between State/UT Licensing Authority and the Notified body can be made through e-vartalap (communication box) as provided in Online System for Medical Devices. ***** Page 9 of 9 Medical Devices Division, Central Drugs Standard Control Organization, Ministry of Health and Family Welfare, Govt. of India