Searchable PDF text layer
MDA/GD/0020 21 November 2022 Fourth Edition MEDICAL DEVICE GUIDANCE DOCUMENT CHANGE NOTIFICATION FOR REGISTERED MEDICAL DEVICE Medical Device Authority MINISTRY OF HEALTH MALAYSIA
Currentness under review. Whether this instrument remains applicable is under review. Consult the later instruments and source context below before relying on it.
Original page images are authoritative. Searchable text is the PDF text layer and may omit image text or contain extraction artifacts. Quote and verify against the page images.
Link directly to a passage with its anchor — for example #pdf-page-1 — to open it highlighted. This reference version is generated from the authority’s publication and does not replace it. Language: English.
MDA/GD/0020 21 November 2022 Fourth Edition MEDICAL DEVICE GUIDANCE DOCUMENT CHANGE NOTIFICATION FOR REGISTERED MEDICAL DEVICE Medical Device Authority MINISTRY OF HEALTH MALAYSIA
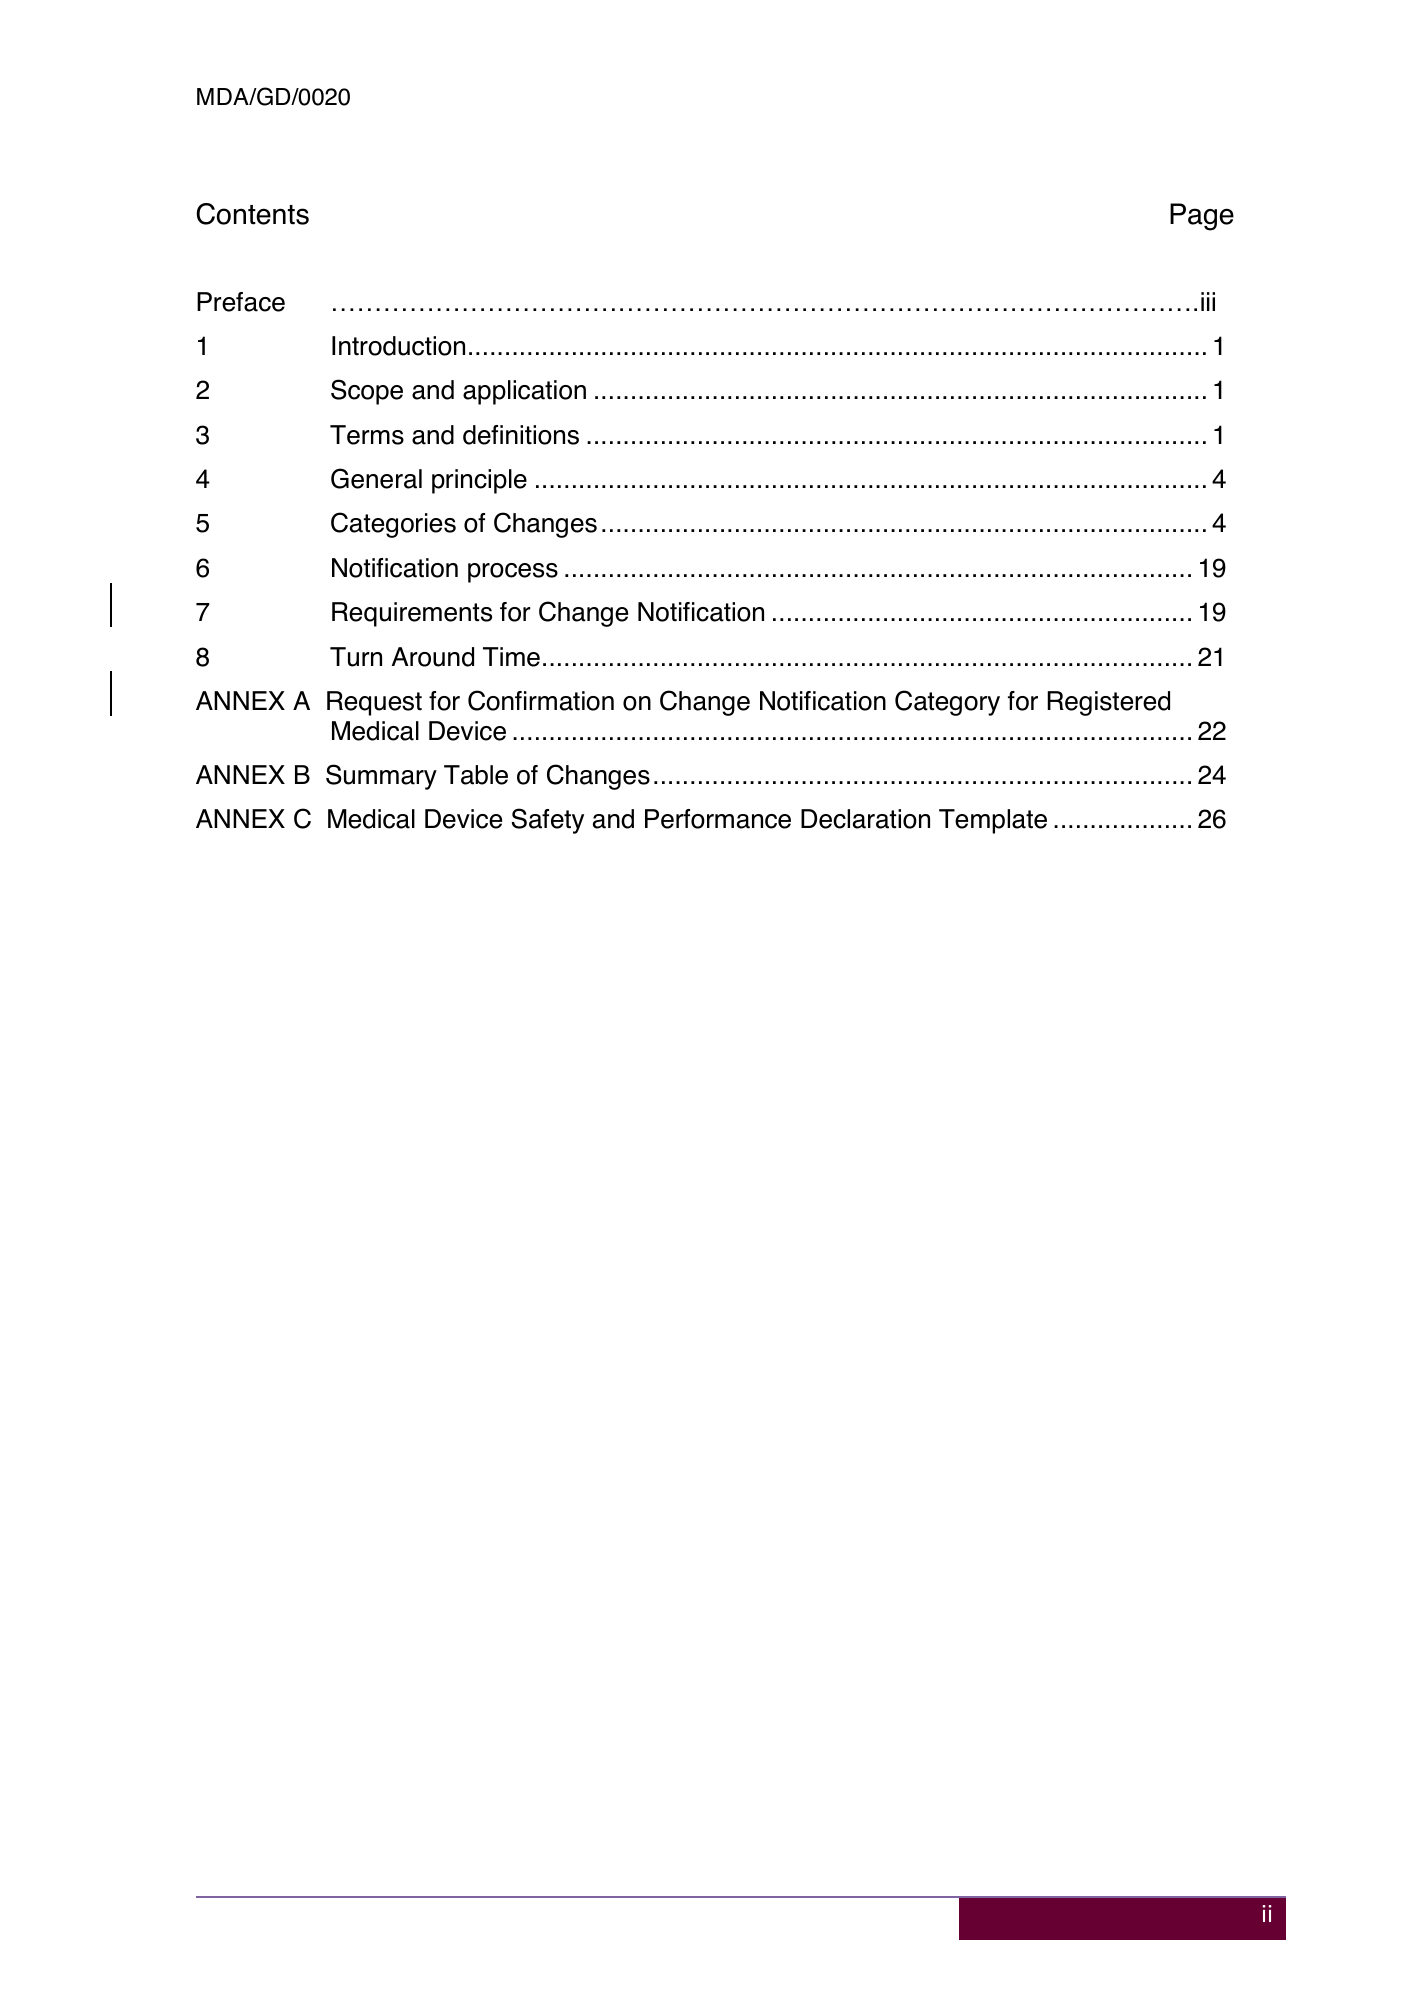
MDA/GD/0020 Contents Page Preface ……………………………………………………………………………………….iii 1 Introduction.................................................................................................... 1 2 Scope and application ................................................................................... 1 3 Terms and definitions .................................................................................... 1 4 General principle ........................................................................................... 4 5 Categories of Changes .................................................................................. 4 6 Notification process ..................................................................................... 19 7 Requirements for Change Notification ......................................................... 19 8 Turn Around Time........................................................................................ 21 ANNEX A Request for Confirmation on Change Notification Category for Registered Medical Device ............................................................................................ 22 ANNEX B Summary Table of Changes ......................................................................... 24 ANNEX C Medical Device Safety and Performance Declaration Template ................... 26 ii
MDA/GD/0020 Preface This Guidance Document was prepared by the Medical Device Authority (MDA) to help the industry and healthcare professionals in their quest to comply with the Medical Device Act (Act 737) and the regulations under it. This Guidance Document shall be read in conjunction with the current laws and regulations used in Malaysia, which include but not limited to the following- a) Medical Device Act 2012 (Act 737); and b) Medical Device Regulations 2012; and c) Medical Device (Duties and Obligations of Establishments) Regulations 2019. In this Guidance Document, the following verbal forms are used: — “shall” indicates a requirement; — “should” indicates a recommendation; — “may” indicates a permission; and — “can” indicates a possibility or a capability. Irrespective of the requirements of this Guidance Document, MDA has the right to request for information or material, or define conditions not specifically described in this document that is deemed necessary for the purpose of regulatory control. MDA has put much effort to ensure the accuracy and completeness of this guidance document. In the incident of any contradiction between the contents of this document and any written law, the latter should take precedence. MDA reserves the right to amend any part of the guidance document from time to time. CONTACT INFORMATION For further information, please contact: MEDICAL DEVICE AUTHORITY Ministry of Health Malaysia Level 6, Prima 9, Prima Avenue II, Block 3547, Persiaran APEC, 63000 Cyberjaya, Selangor, MALAYSIA Fax: (03) 8230 0200 Email: mdb@mda.gov.my Website: http://www.mda.gov.my iii
MDA/GD/0020 CHANGE NOTIFICATION FOR REGISTERED MEDICAL DEVICE 1 Introduction Changes in medical devices may take place from time to time as part of their life-cycle. Any change to a registered medical device is linked to the principles of safety and performance and the ability of the regulatory framework to manage the risk of the medical devices. Before making any decision whether a changed medical device can continue to be placed in the market, the Authority will determine whether evidence of safety and performance have been appropriately collected and reviewed based on the notification made by the registration holder. For any anticipated change to a medical device, a manufacturer must consider the impact of the change on the patient, practitioner and/or user of the medical device, and the impact of the change on the specifications of the medical device, and decide whether the change is expected to impact the safety and performance of the medical device. This document provides guidance on the categories of changes, the principles of change categorisation, and what should be done by the registration holder in relation to each category of change to its registered medical device. 2 Scope and application This guidance document specifies the categories of changes in relation to registered medical devices and the requirements to be met to continue the importation, exportation or placement of the medical devices in the market. This document applies to all registered medical devices under the Act. It sets out points for consideration by the registration holder when a registered medical device is in the process of change or modification. Owing to the various possible scenarios for changes made to a medical device, it is not the intention of this document to describe every permutation and type of change that can occur. This document is also applicable to situations when a registered medical device undergoes any changes or proposed changes as a result of a mandatory reportable incident or field corrective action under Section 40 or Section 41 of Act 737 respectively. 3 Terms and definitions For the purpose of this document, the terms and definitions in Act 737, the regulations under it and the following terms and definitions apply. 1
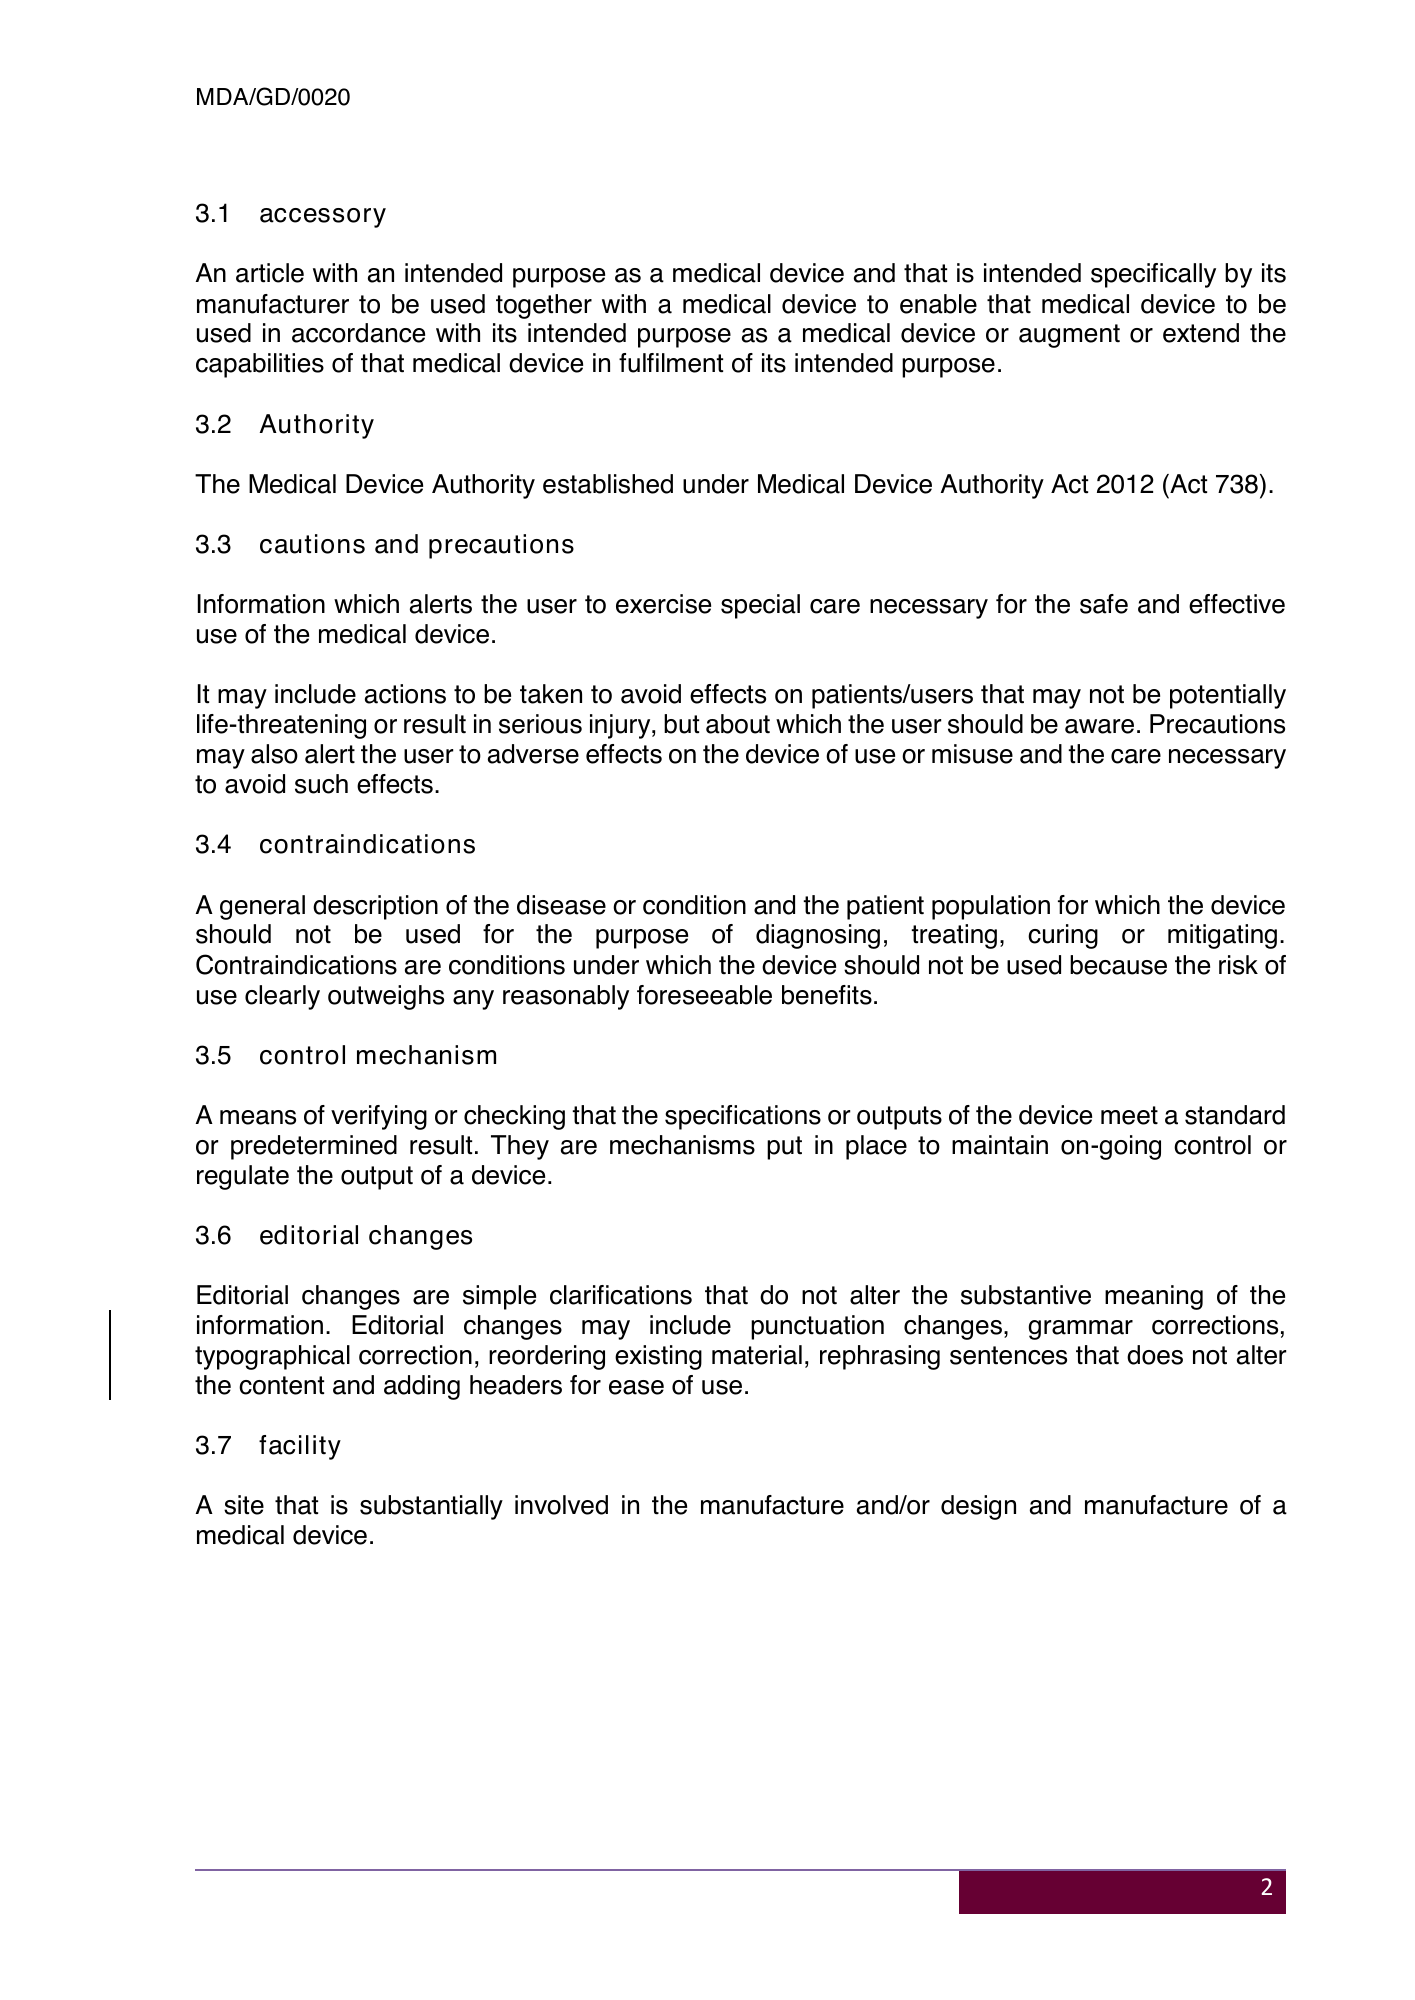
MDA/GD/0020 3.1 accessory An article with an intended purpose as a medical device and that is intended specifically by its manufacturer to be used together with a medical device to enable that medical device to be used in accordance with its intended purpose as a medical device or augment or extend the capabilities of that medical device in fulfilment of its intended purpose. 3.2 Authority The Medical Device Authority established under Medical Device Authority Act 2012 (Act 738). 3.3 cautions and precautions Information which alerts the user to exercise special care necessary for the safe and effective use of the medical device. It may include actions to be taken to avoid effects on patients/users that may not be potentially life-threatening or result in serious injury, but about which the user should be aware. Precautions may also alert the user to adverse effects on the device of use or misuse and the care necessary to avoid such effects. 3.4 contraindications A general description of the disease or condition and the patient population for which the device should not be used for the purpose of diagnosing, treating, curing or mitigating. Contraindications are conditions under which the device should not be used because the risk of use clearly outweighs any reasonably foreseeable benefits. 3.5 control mechanism A means of verifying or checking that the specifications or outputs of the device meet a standard or predetermined result. They are mechanisms put in place to maintain on-going control or regulate the output of a device. 3.6 editorial changes Editorial changes are simple clarifications that do not alter the substantive meaning of the information. Editorial changes may include punctuation changes, grammar corrections, typographical correction, reordering existing material, rephrasing sentences that does not alter the content and adding headers for ease of use. 3.7 facility A site that is substantially involved in the manufacture and/or design and manufacture of a medical device. 2
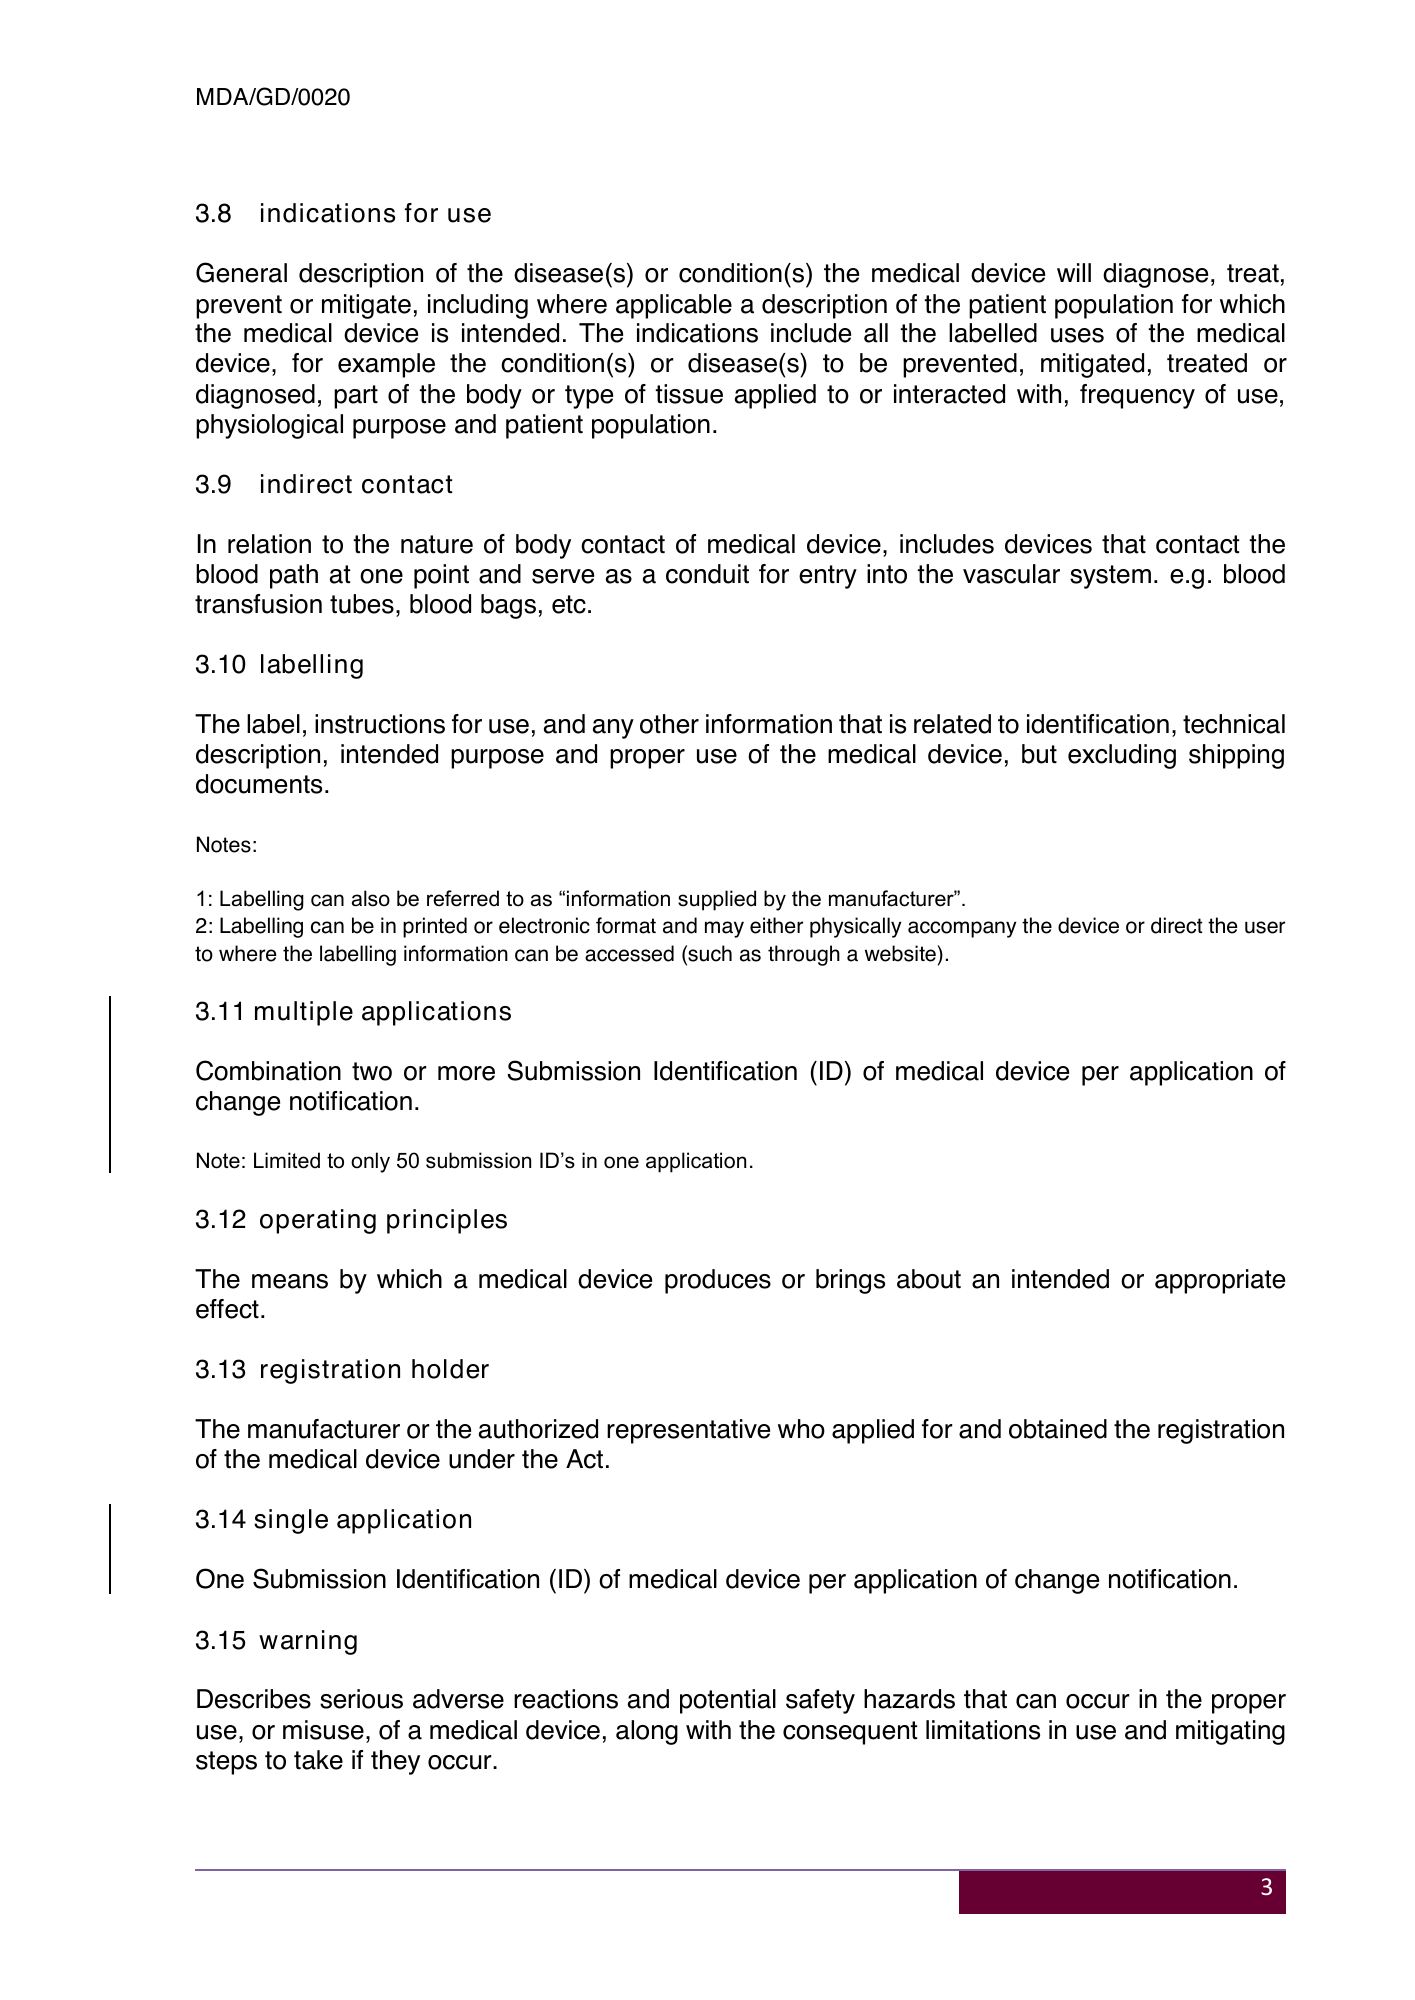
MDA/GD/0020 3.8 indications for use General description of the disease(s) or condition(s) the medical device will diagnose, treat, prevent or mitigate, including where applicable a description of the patient population for which the medical device is intended. The indications include all the labelled uses of the medical device, for example the condition(s) or disease(s) to be prevented, mitigated, treated or diagnosed, part of the body or type of tissue applied to or interacted with, frequency of use, physiological purpose and patient population. 3.9 indirect contact In relation to the nature of body contact of medical device, includes devices that contact the blood path at one point and serve as a conduit for entry into the vascular system. e.g. blood transfusion tubes, blood bags, etc. 3.10 labelling The label, instructions for use, and any other information that is related to identification, technical description, intended purpose and proper use of the medical device, but excluding shipping documents. Notes: 1: Labelling can also be referred to as “information supplied by the manufacturer”. 2: Labelling can be in printed or electronic format and may either physically accompany the device or direct the user to where the labelling information can be accessed (such as through a website). 3.11 multiple applications Combination two or more Submission Identification (ID) of medical device per application of change notification. Note: Limited to only 50 submission ID’s in one application. 3.12 operating principles The means by which a medical device produces or brings about an intended or appropriate effect. 3.13 registration holder The manufacturer or the authorized representative who applied for and obtained the registration of the medical device under the Act. 3.14 single application One Submission Identification (ID) of medical device per application of change notification. 3.15 warning Describes serious adverse reactions and potential safety hazards that can occur in the proper use, or misuse, of a medical device, along with the consequent limitations in use and mitigating steps to take if they occur. 3
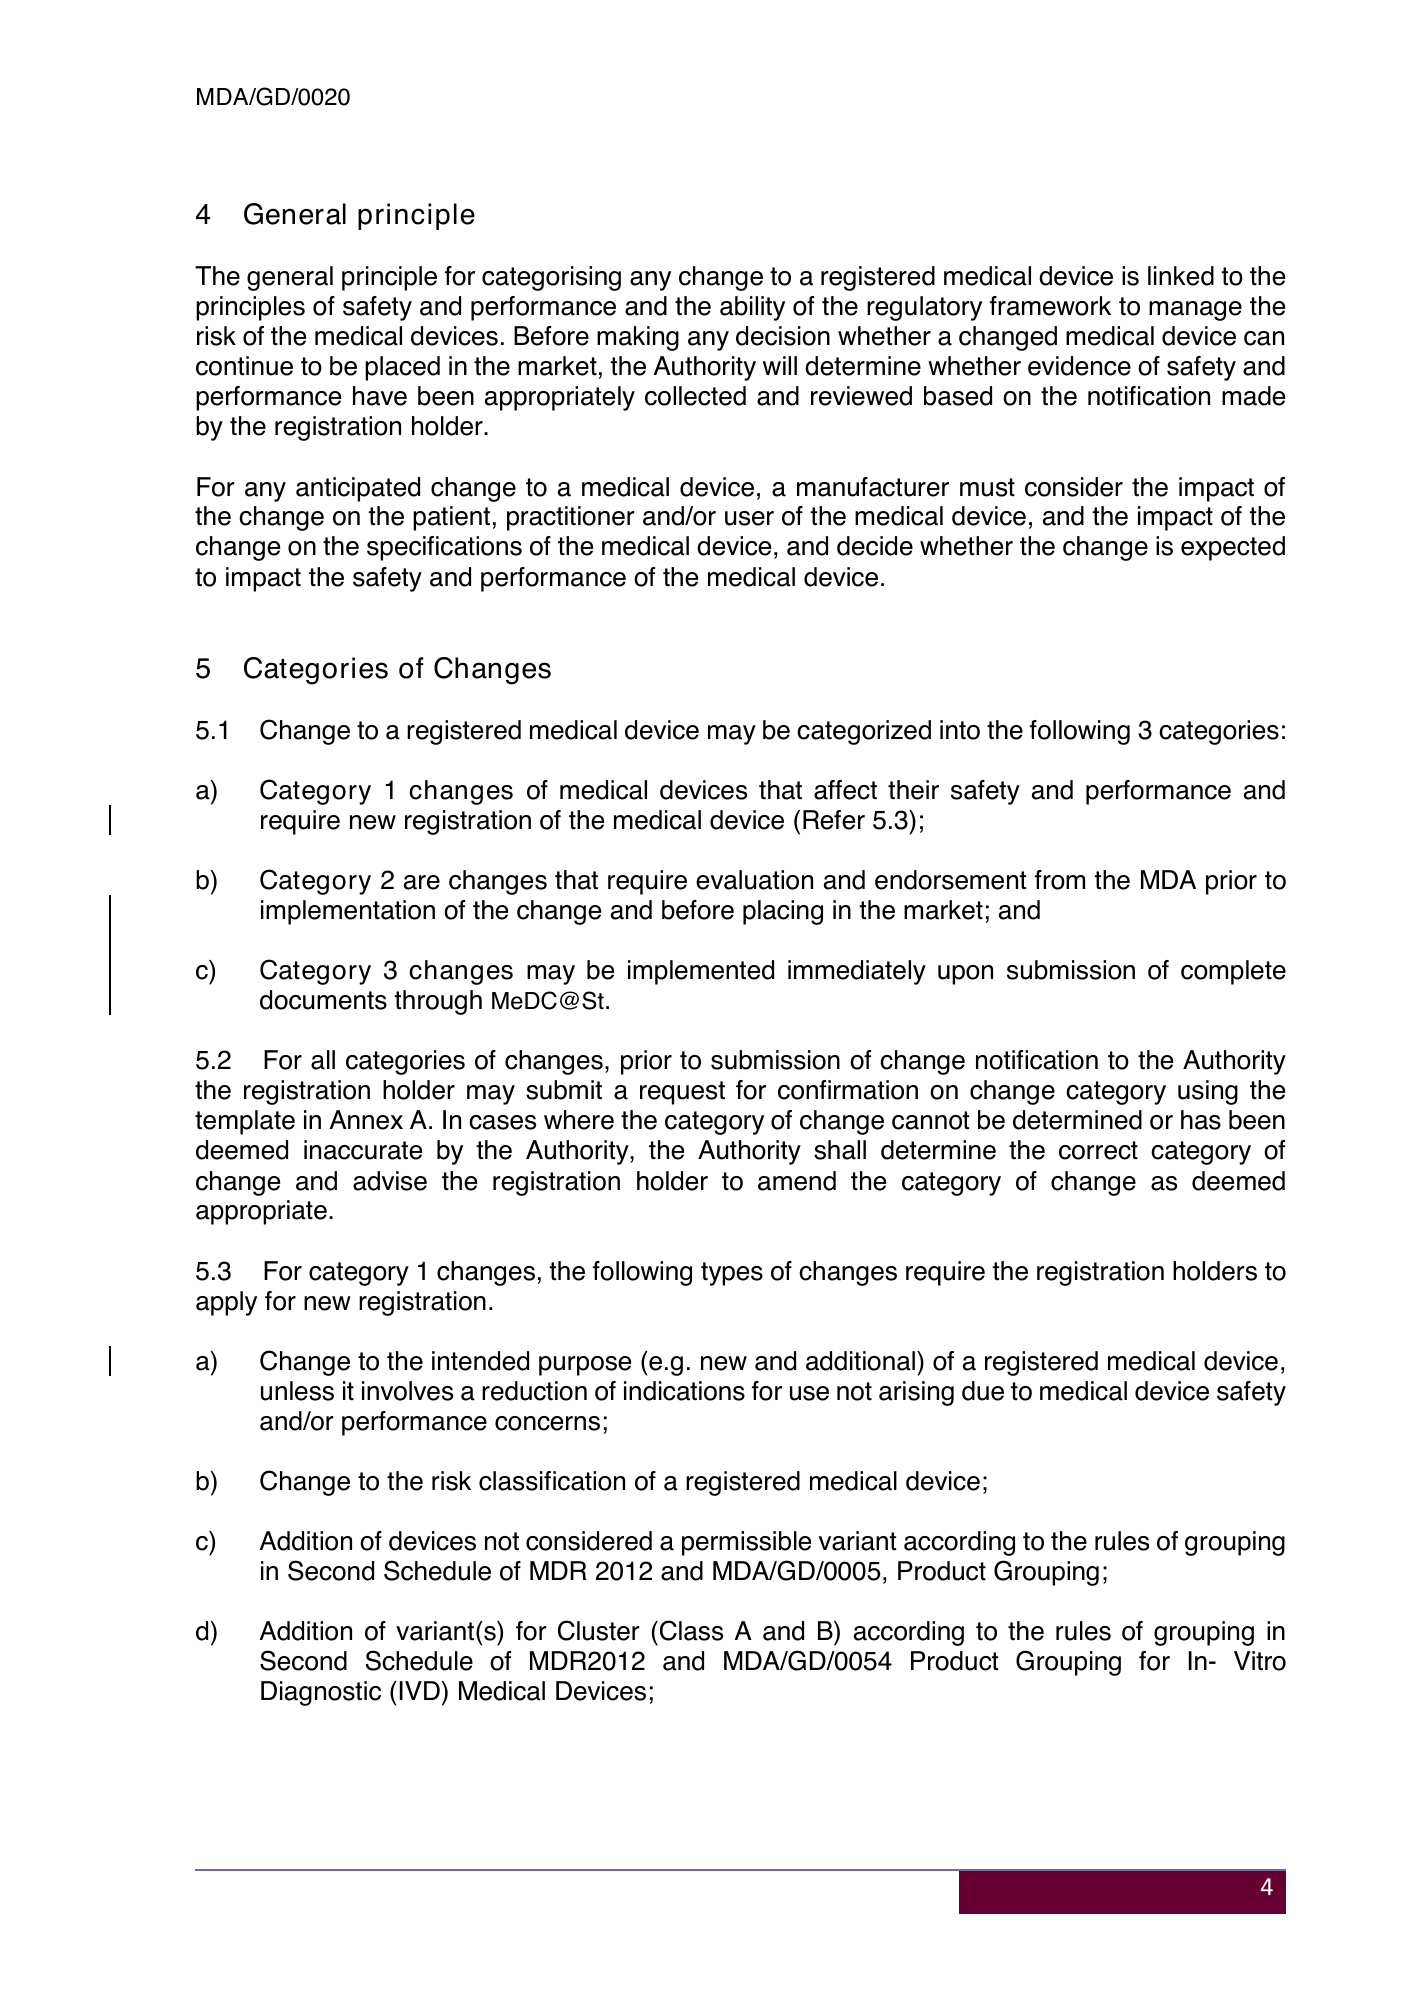
MDA/GD/0020 4 General principle The general principle for categorising any change to a registered medical device is linked to the principles of safety and performance and the ability of the regulatory framework to manage the risk of the medical devices. Before making any decision whether a changed medical device can continue to be placed in the market, the Authority will determine whether evidence of safety and performance have been appropriately collected and reviewed based on the notification made by the registration holder. For any anticipated change to a medical device, a manufacturer must consider the impact of the change on the patient, practitioner and/or user of the medical device, and the impact of the change on the specifications of the medical device, and decide whether the change is expected to impact the safety and performance of the medical device. 5 Categories of Changes 5.1 Change to a registered medical device may be categorized into the following 3 categories: a) Category 1 changes of medical devices that affect their safety and performance and require new registration of the medical device (Refer 5.3); b) Category 2 are changes that require evaluation and endorsement from the MDA prior to implementation of the change and before placing in the market; and c) Category 3 changes may be implemented immediately upon submission of complete documents through MeDC@St. 5.2 For all categories of changes, prior to submission of change notification to the Authority the registration holder may submit a request for confirmation on change category using the template in Annex A. In cases where the category of change cannot be determined or has been deemed inaccurate by the Authority, the Authority shall determine the correct category of change and advise the registration holder to amend the category of change as deemed appropriate. 5.3 For category 1 changes, the following types of changes require the registration holders to apply for new registration. a) Change to the intended purpose (e.g. new and additional) of a registered medical device, unless it involves a reduction of indications for use not arising due to medical device safety and/or performance concerns; b) Change to the risk classification of a registered medical device; c) Addition of devices not considered a permissible variant according to the rules of grouping in Second Schedule of MDR 2012 and MDA/GD/0005, Product Grouping; d) Addition of variant(s) for Cluster (Class A and B) according to the rules of grouping in Second Schedule of MDR2012 and MDA/GD/0054 Product Grouping for In- Vitro Diagnostic (IVD) Medical Devices; 4
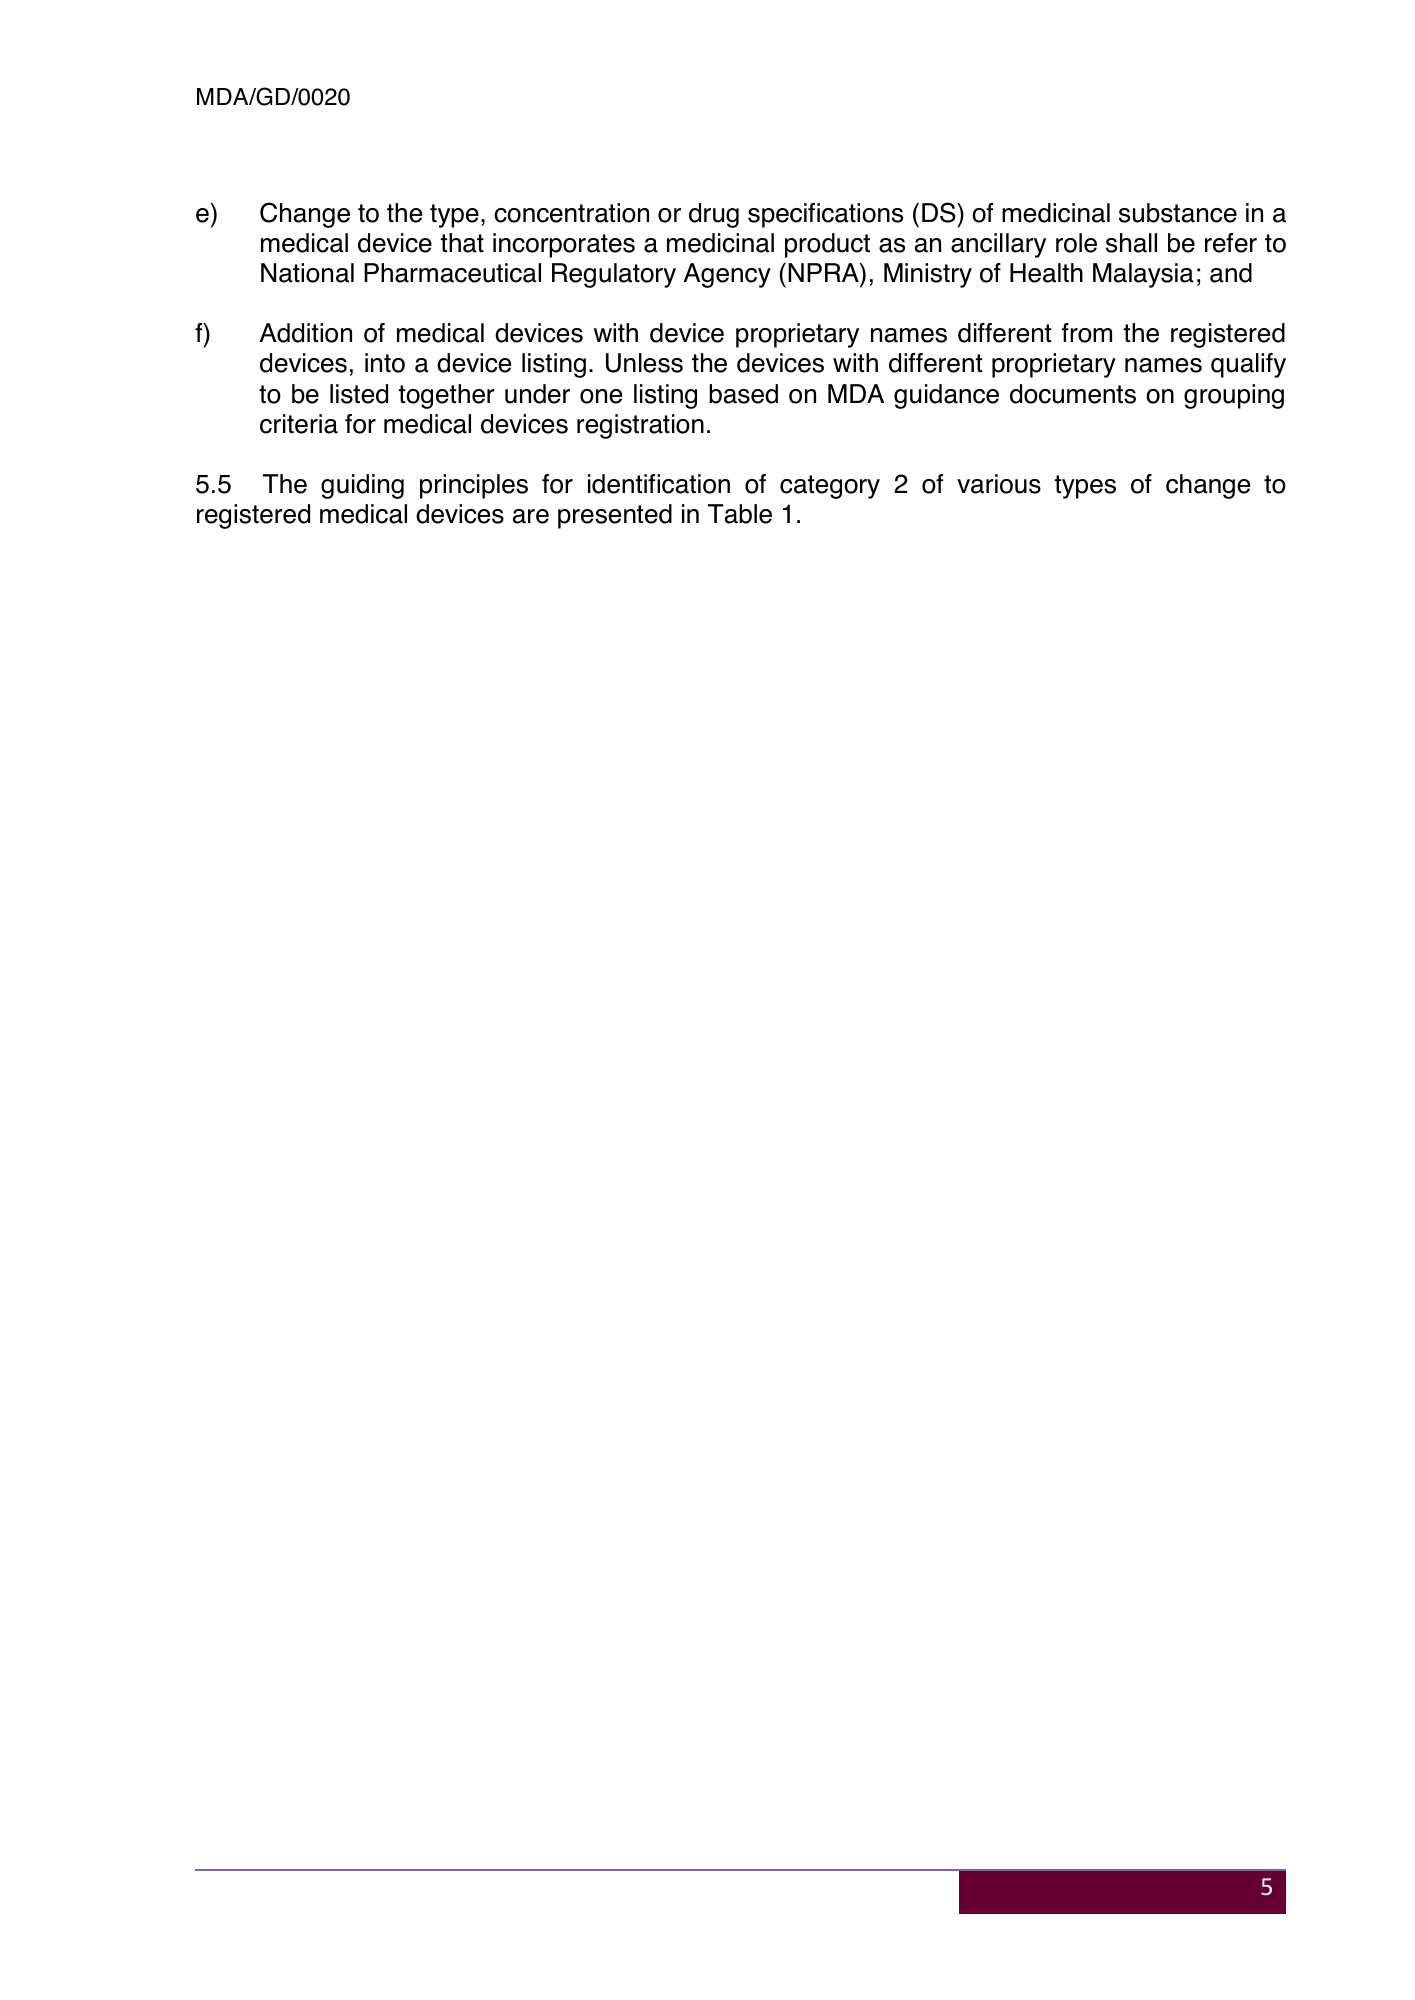
MDA/GD/0020 e) Change to the type, concentration or drug specifications (DS) of medicinal substance in a medical device that incorporates a medicinal product as an ancillary role shall be refer to National Pharmaceutical Regulatory Agency (NPRA), Ministry of Health Malaysia; and f) Addition of medical devices with device proprietary names different from the registered devices, into a device listing. Unless the devices with different proprietary names qualify to be listed together under one listing based on MDA guidance documents on grouping criteria for medical devices registration. 5.5 The guiding principles for identification of category 2 of various types of change to registered medical devices are presented in Table 1. 5
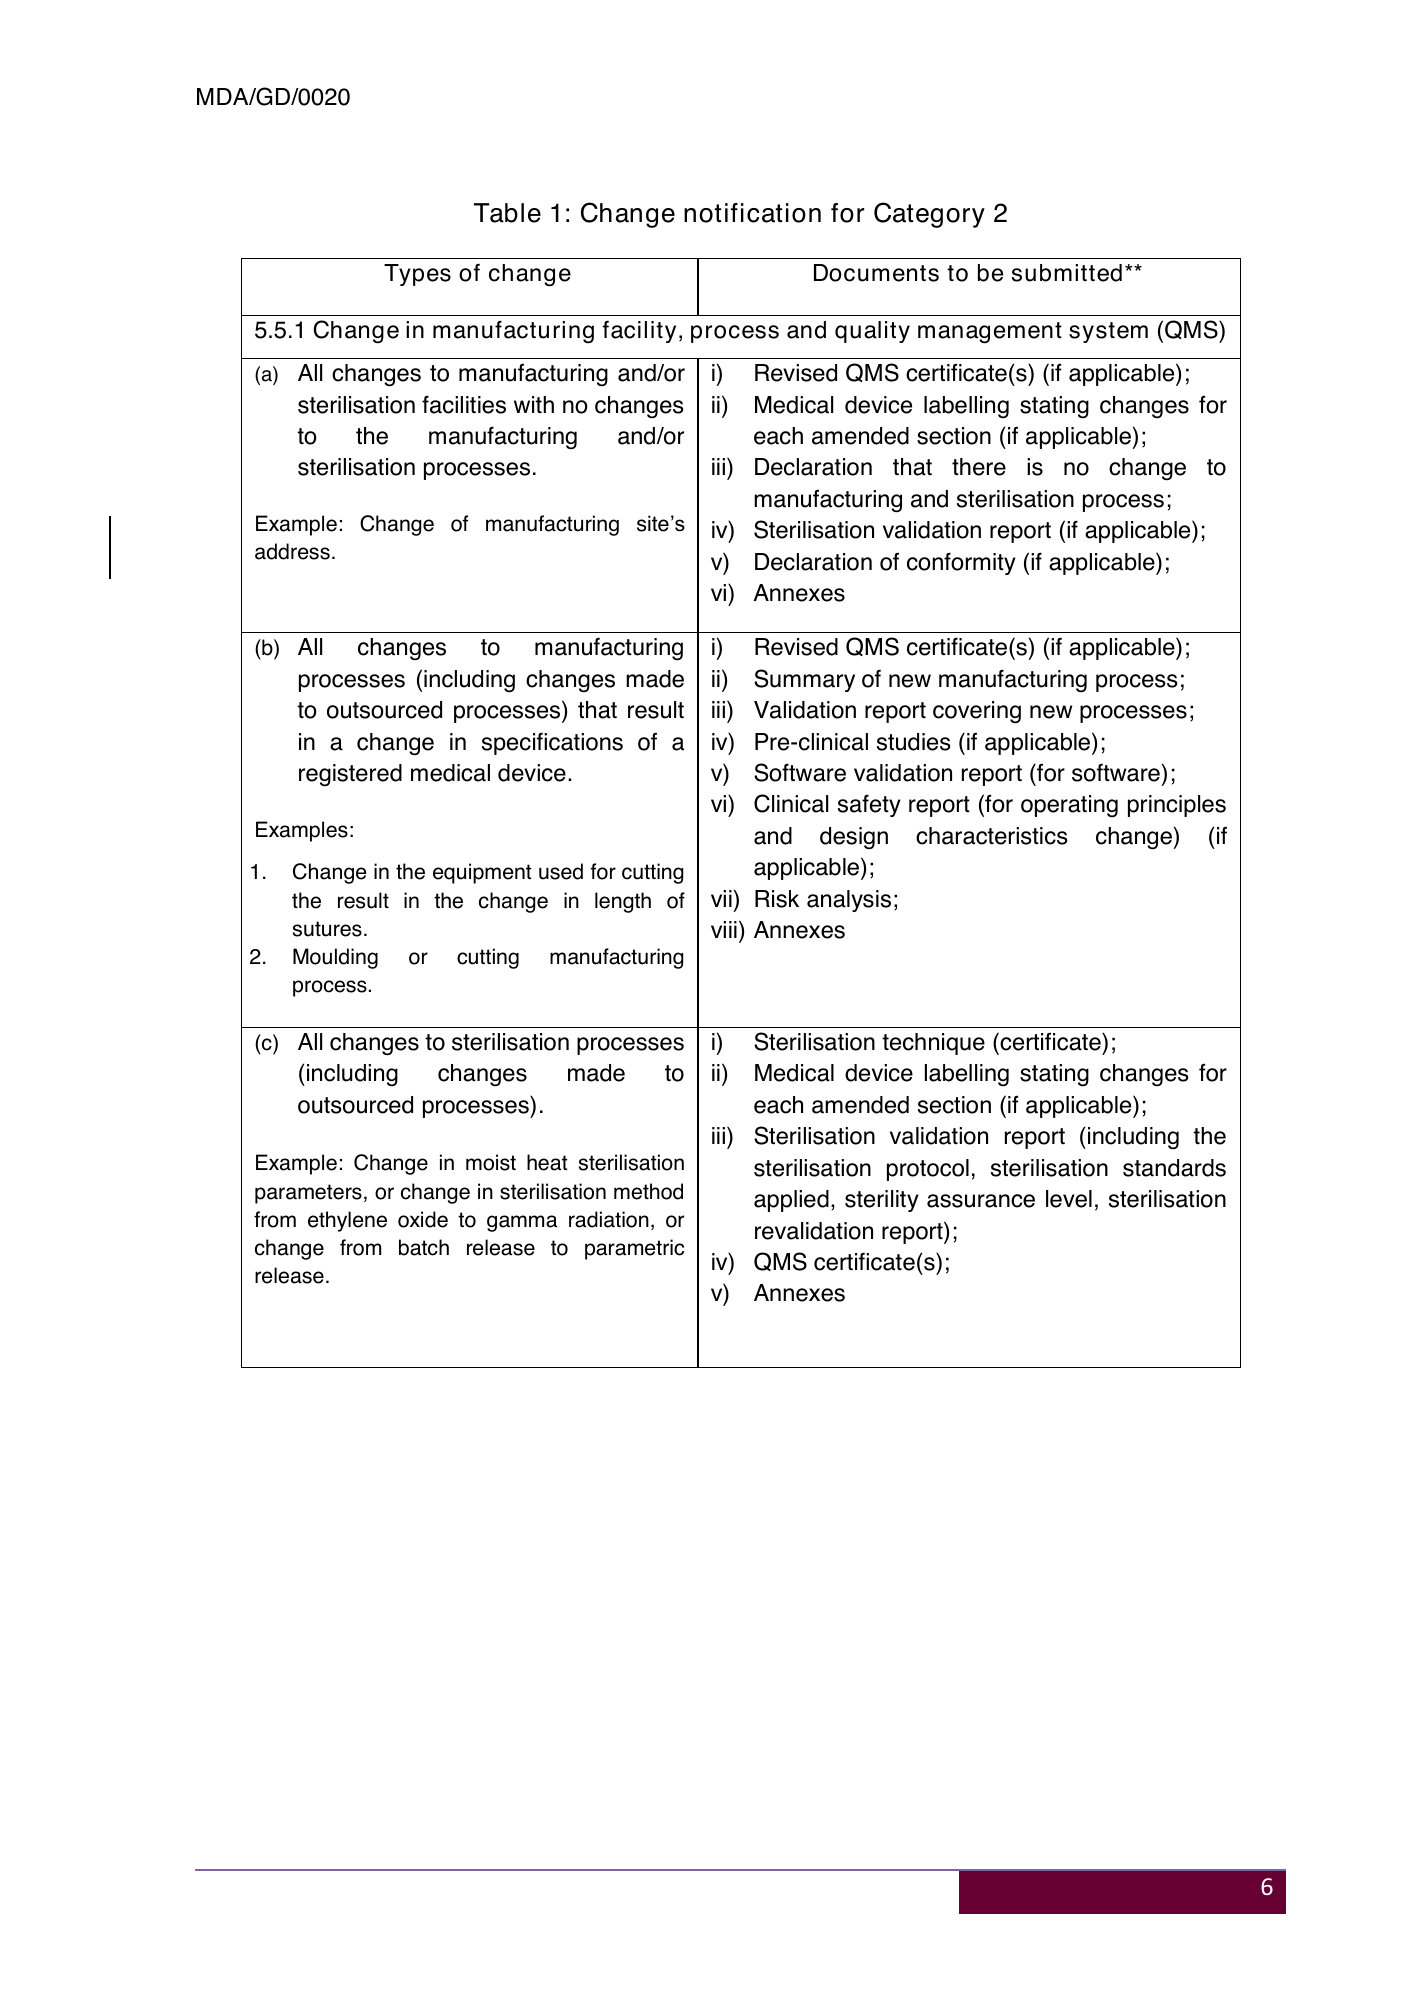
MDA/GD/0020 Table 1: Change notification for Category 2 Types of change Documents to be submitted** 5.5.1 Change in manufacturing facility, process and quality management system (QMS) (a) All changes to manufacturing and/or i) Revised QMS certificate(s) (if applicable); sterilisation facilities with no changes ii) Medical device labelling stating changes for to the manufacturing and/or each amended section (if applicable); sterilisation processes. iii) Declaration that there is no change to manufacturing and sterilisation process; Example: Change of manufacturing site’s iv) Sterilisation validation report (if applicable); address. v) Declaration of conformity (if applicable); vi) Annexes (b) All changes to manufacturing i) Revised QMS certificate(s) (if applicable); processes (including changes made ii) Summary of new manufacturing process; to outsourced processes) that result iii) Validation report covering new processes; in a change in specifications of a iv) Pre-clinical studies (if applicable); registered medical device. v) Software validation report (for software); vi) Clinical safety report (for operating principles Examples: and design characteristics change) (if 1. Change in the equipment used for cutting applicable); the result in the change in length of vii) Risk analysis; sutures. viii) Annexes 2. Moulding or cutting manufacturing process. (c) All changes to sterilisation processes i) Sterilisation technique (certificate); (including changes made to ii) Medical device labelling stating changes for outsourced processes). each amended section (if applicable); iii) Sterilisation validation report (including the Example: Change in moist heat sterilisation sterilisation protocol, sterilisation standards parameters, or change in sterilisation method applied, sterility assurance level, sterilisation from ethylene oxide to gamma radiation, or revalidation report); change from batch release to parametric iv) QMS certificate(s); release. v) Annexes 6
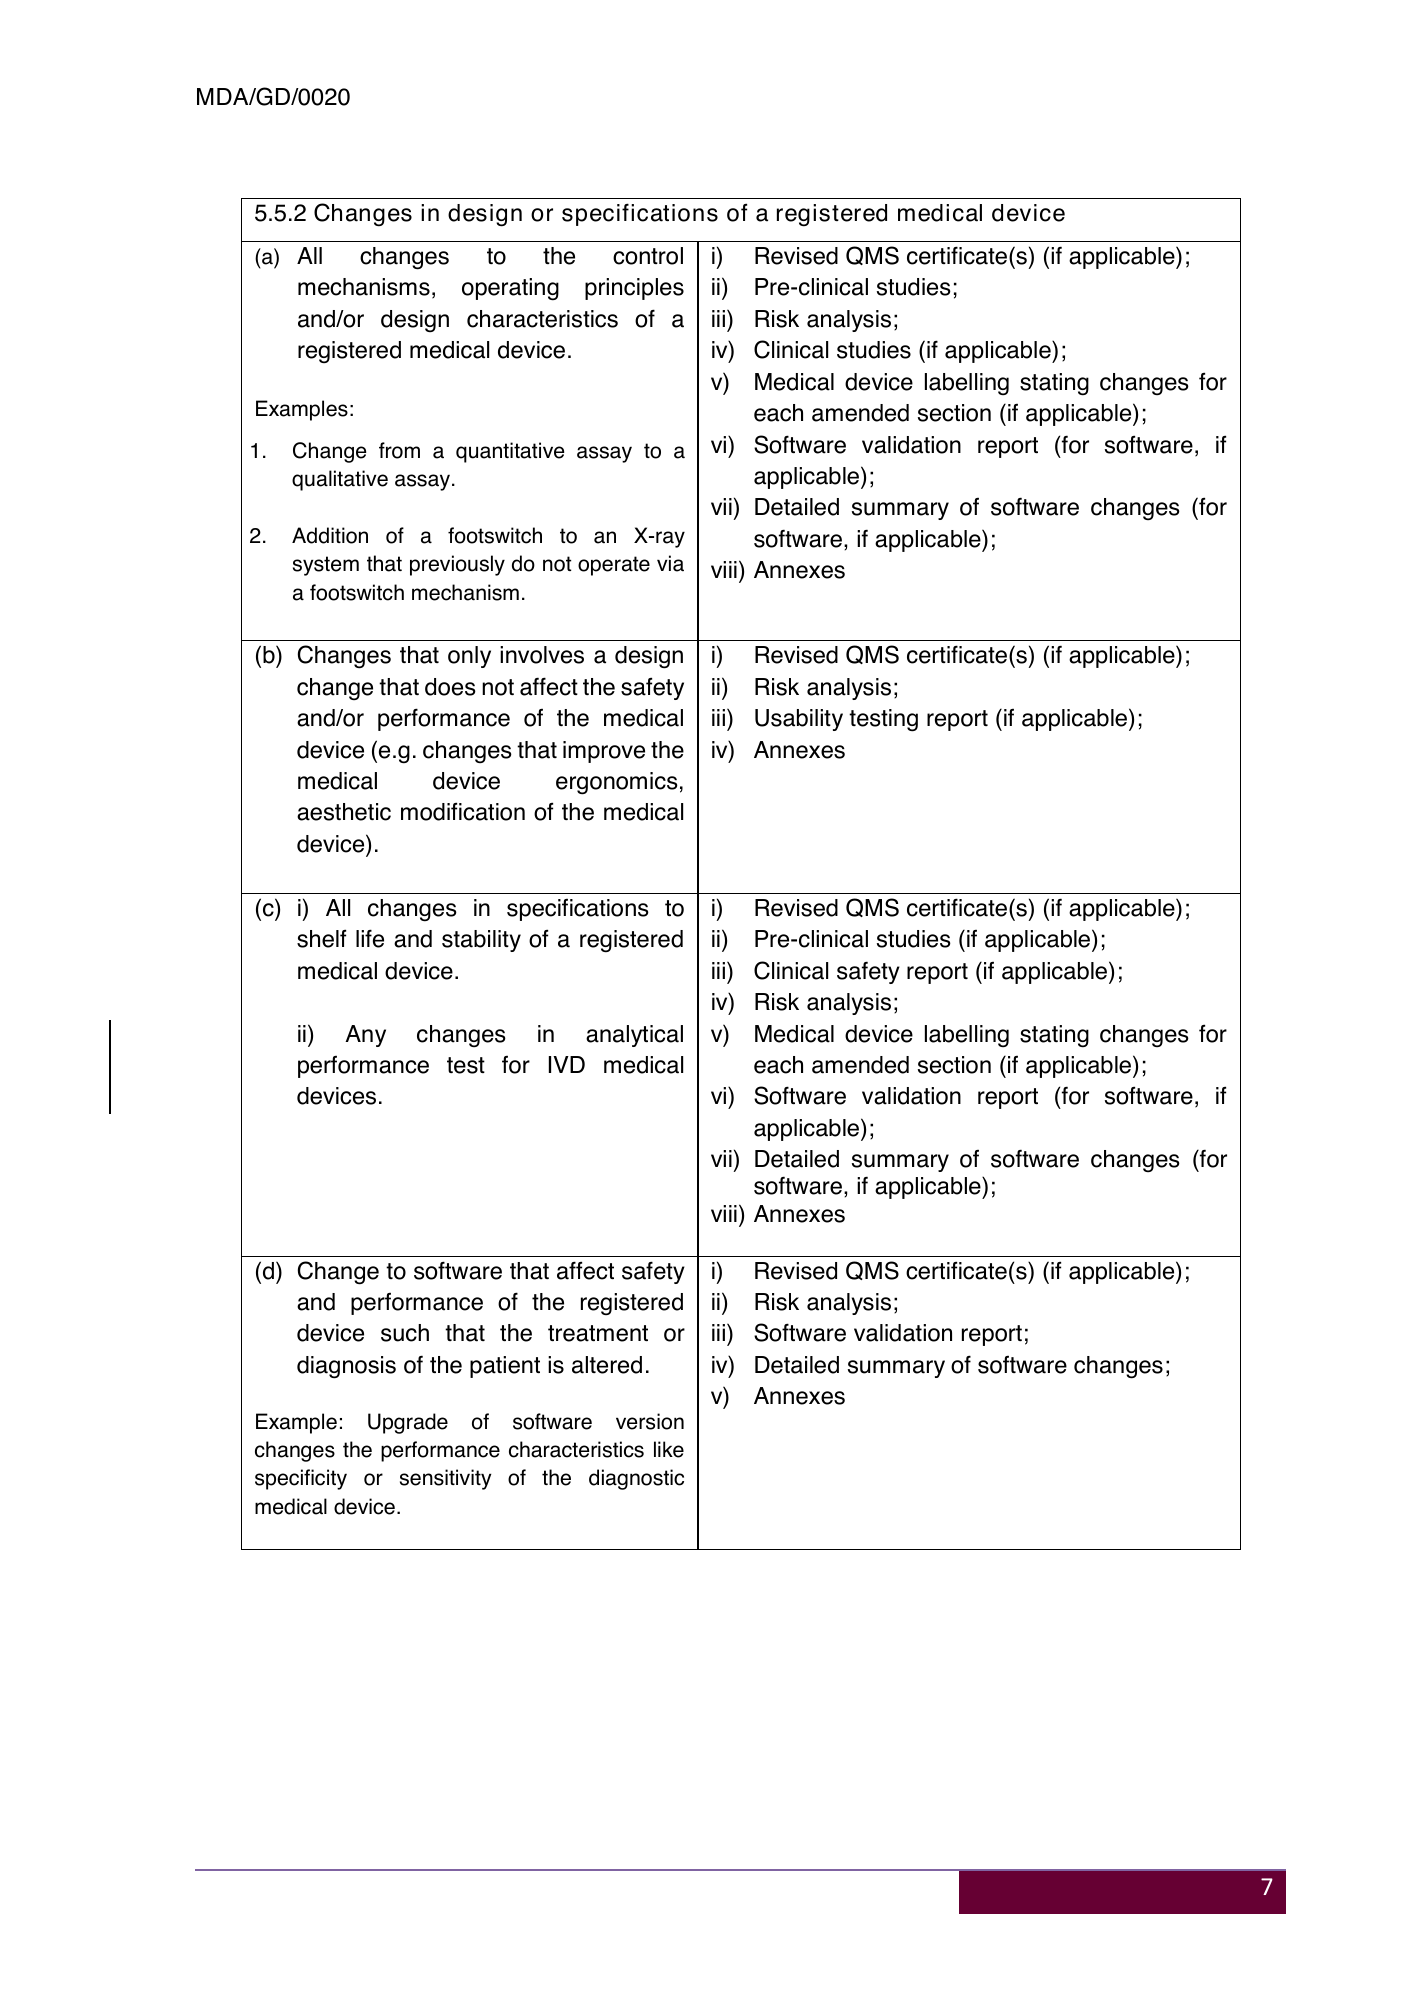
MDA/GD/0020 5.5.2 Changes in design or specifications of a registered medical device (a) All changes to the control i) Revised QMS certificate(s) (if applicable); mechanisms, operating principles ii) Pre-clinical studies; and/or design characteristics of a iii) Risk analysis; registered medical device. iv) Clinical studies (if applicable); v) Medical device labelling stating changes for Examples: each amended section (if applicable); 1. Change from a quantitative assay to a vi) Software validation report (for software, if qualitative assay. applicable); vii) Detailed summary of software changes (for 2. Addition of a footswitch to an X-ray software, if applicable); system that previously do not operate via viii) Annexes a footswitch mechanism. (b) Changes that only involves a design i) Revised QMS certificate(s) (if applicable); change that does not affect the safety ii) Risk analysis; and/or performance of the medical iii) Usability testing report (if applicable); device (e.g. changes that improve the iv) Annexes medical device ergonomics, aesthetic modification of the medical device). (c) i) All changes in specifications to i) Revised QMS certificate(s) (if applicable); shelf life and stability of a registered ii) Pre-clinical studies (if applicable); medical device. iii) Clinical safety report (if applicable); iv) Risk analysis; ii) Any changes in analytical v) Medical device labelling stating changes for performance test for IVD medical each amended section (if applicable); devices. vi) Software validation report (for software, if applicable); vii) Detailed summary of software changes (for software, if applicable); viii) Annexes (d) Change to software that affect safety i) Revised QMS certificate(s) (if applicable); and performance of the registered ii) Risk analysis; device such that the treatment or iii) Software validation report; diagnosis of the patient is altered. iv) Detailed summary of software changes; v) Annexes Example: Upgrade of software version changes the performance characteristics like specificity or sensitivity of the diagnostic medical device. 7
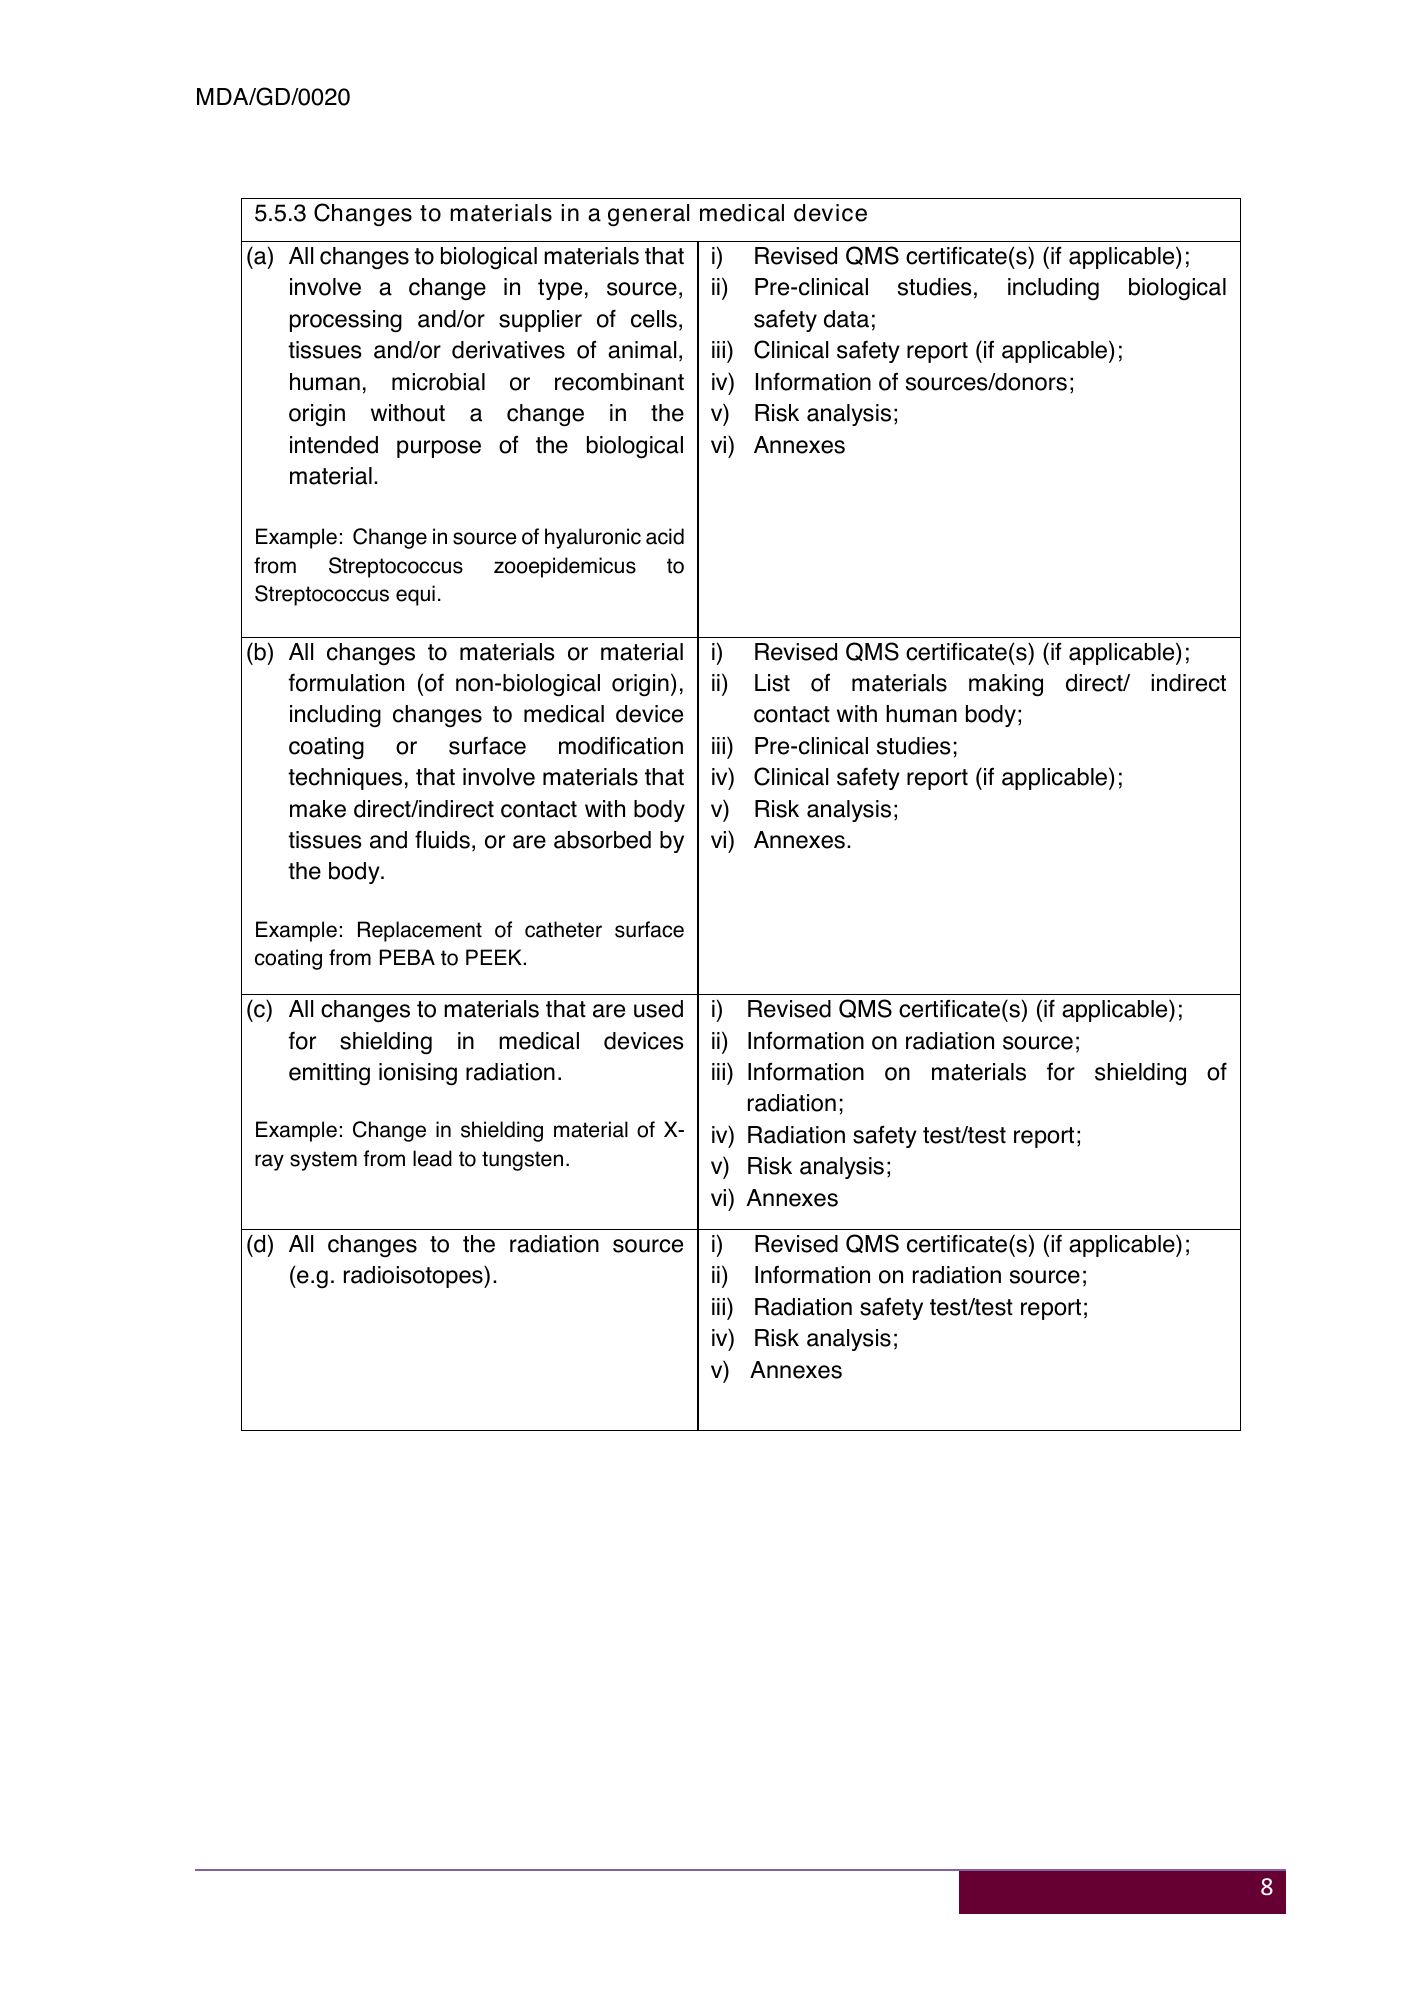
MDA/GD/0020 5.5.3 Changes to materials in a general medical device (a) All changes to biological materials that i) Revised QMS certificate(s) (if applicable); involve a change in type, source, ii) Pre-clinical studies, including biological processing and/or supplier of cells, safety data; tissues and/or derivatives of animal, iii) Clinical safety report (if applicable); human, microbial or recombinant iv) Information of sources/donors; origin without a change in the v) Risk analysis; intended purpose of the biological vi) Annexes material. Example: Change in source of hyaluronic acid from Streptococcus zooepidemicus to Streptococcus equi. (b) All changes to materials or material i) Revised QMS certificate(s) (if applicable); formulation (of non-biological origin), ii) List of materials making direct/ indirect including changes to medical device contact with human body; coating or surface modification iii) Pre-clinical studies; techniques, that involve materials that iv) Clinical safety report (if applicable); make direct/indirect contact with body v) Risk analysis; tissues and fluids, or are absorbed by vi) Annexes. the body. Example: Replacement of catheter surface coating from PEBA to PEEK. (c) All changes to materials that are used i) Revised QMS certificate(s) (if applicable); for shielding in medical devices ii) Information on radiation source; emitting ionising radiation. iii) Information on materials for shielding of radiation; Example: Change in shielding material of X- iv) Radiation safety test/test report; ray system from lead to tungsten. v) Risk analysis; vi) Annexes (d) All changes to the radiation source i) Revised QMS certificate(s) (if applicable); (e.g. radioisotopes). ii) Information on radiation source; iii) Radiation safety test/test report; iv) Risk analysis; v) Annexes 8
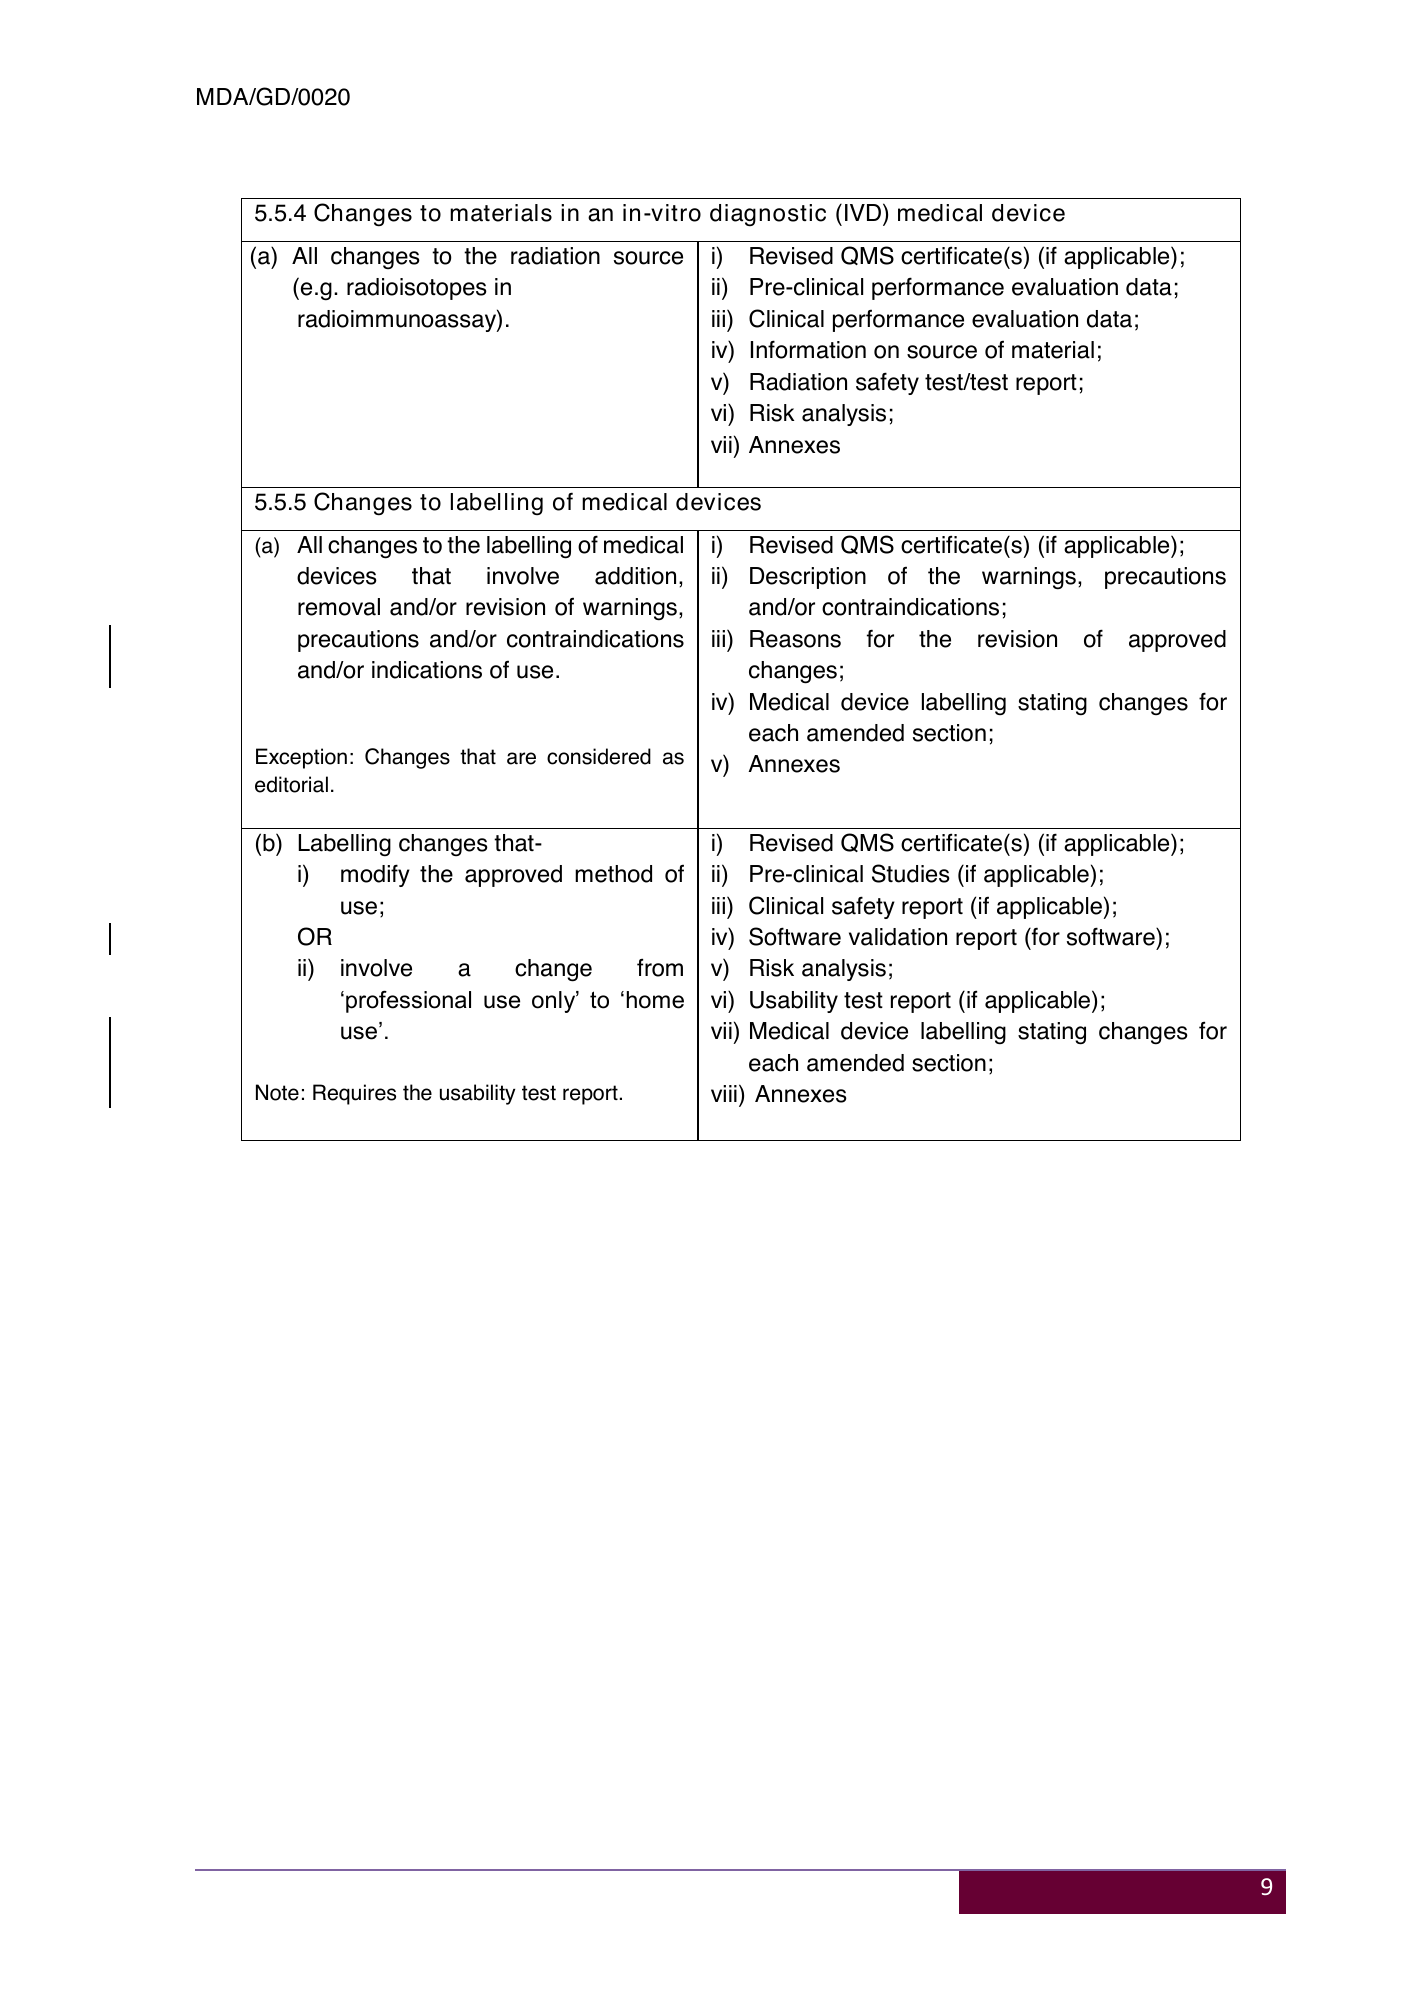
MDA/GD/0020 5.5.4 Changes to materials in an in-vitro diagnostic (IVD) medical device (a) All changes to the radiation source i) Revised QMS certificate(s) (if applicable); (e.g. radioisotopes in ii) Pre-clinical performance evaluation data; radioimmunoassay). iii) Clinical performance evaluation data; iv) Information on source of material; v) Radiation safety test/test report; vi) Risk analysis; vii) Annexes 5.5.5 Changes to labelling of medical devices (a) All changes to the labelling of medical i) Revised QMS certificate(s) (if applicable); devices that involve addition, ii) Description of the warnings, precautions removal and/or revision of warnings, and/or contraindications; precautions and/or contraindications iii) Reasons for the revision of approved and/or indications of use. changes; iv) Medical device labelling stating changes for each amended section; Exception: Changes that are considered as v) Annexes editorial. (b) Labelling changes that- i) Revised QMS certificate(s) (if applicable); i) modify the approved method of ii) Pre-clinical Studies (if applicable); use; iii) Clinical safety report (if applicable); OR iv) Software validation report (for software); ii) involve a change from v) Risk analysis; ‘professional use only’ to ‘home vi) Usability test report (if applicable); use’. vii) Medical device labelling stating changes for each amended section; Note: Requires the usability test report. viii) Annexes 9
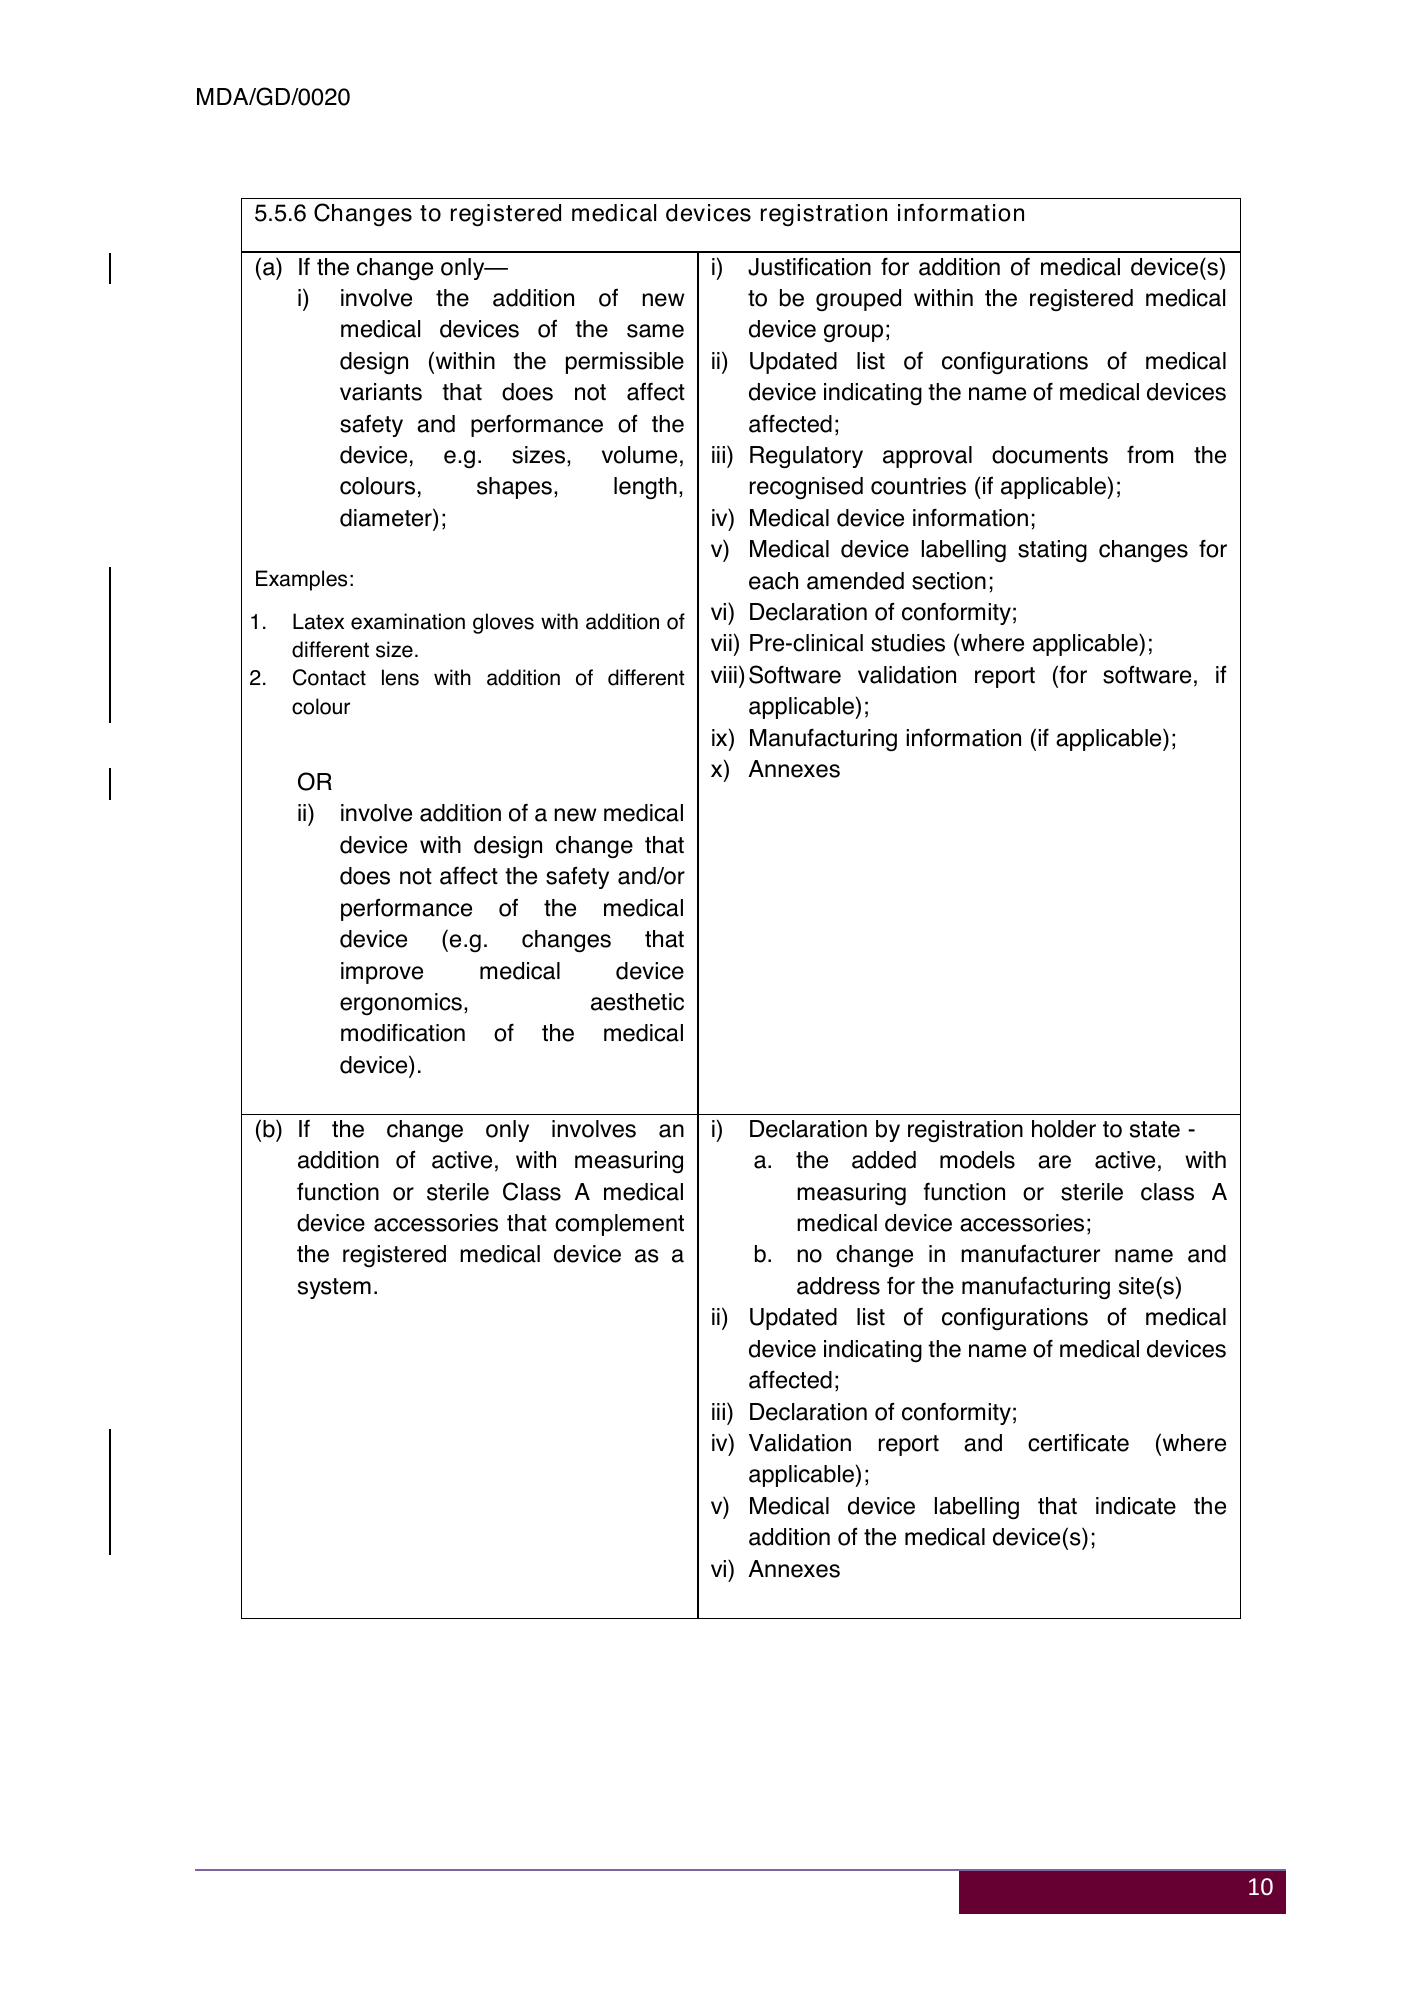
MDA/GD/0020 5.5.6 Changes to registered medical devices registration information (a) If the change only— i) Justification for addition of medical device(s) i) involve the addition of new to be grouped within the registered medical medical devices of the same device group; design (within the permissible ii) Updated list of configurations of medical variants that does not affect device indicating the name of medical devices safety and performance of the affected; device, e.g. sizes, volume, iii) Regulatory approval documents from the colours, shapes, length, recognised countries (if applicable); diameter); iv) Medical device information; v) Medical device labelling stating changes for Examples: each amended section; 1. Latex examination gloves with addition of vi) Declaration of conformity; different size. vii) Pre-clinical studies (where applicable); 2. Contact lens with addition of different viii) Software validation report (for software, if colour applicable); ix) Manufacturing information (if applicable); x) Annexes OR ii) involve addition of a new medical device with design change that does not affect the safety and/or performance of the medical device (e.g. changes that improve medical device ergonomics, aesthetic modification of the medical device). (b) If the change only involves an i) Declaration by registration holder to state - addition of active, with measuring a. the added models are active, with function or sterile Class A medical measuring function or sterile class A device accessories that complement medical device accessories; the registered medical device as a b. no change in manufacturer name and system. address for the manufacturing site(s) ii) Updated list of configurations of medical device indicating the name of medical devices affected; iii) Declaration of conformity; iv) Validation report and certificate (where applicable); v) Medical device labelling that indicate the addition of the medical device(s); vi) Annexes 10
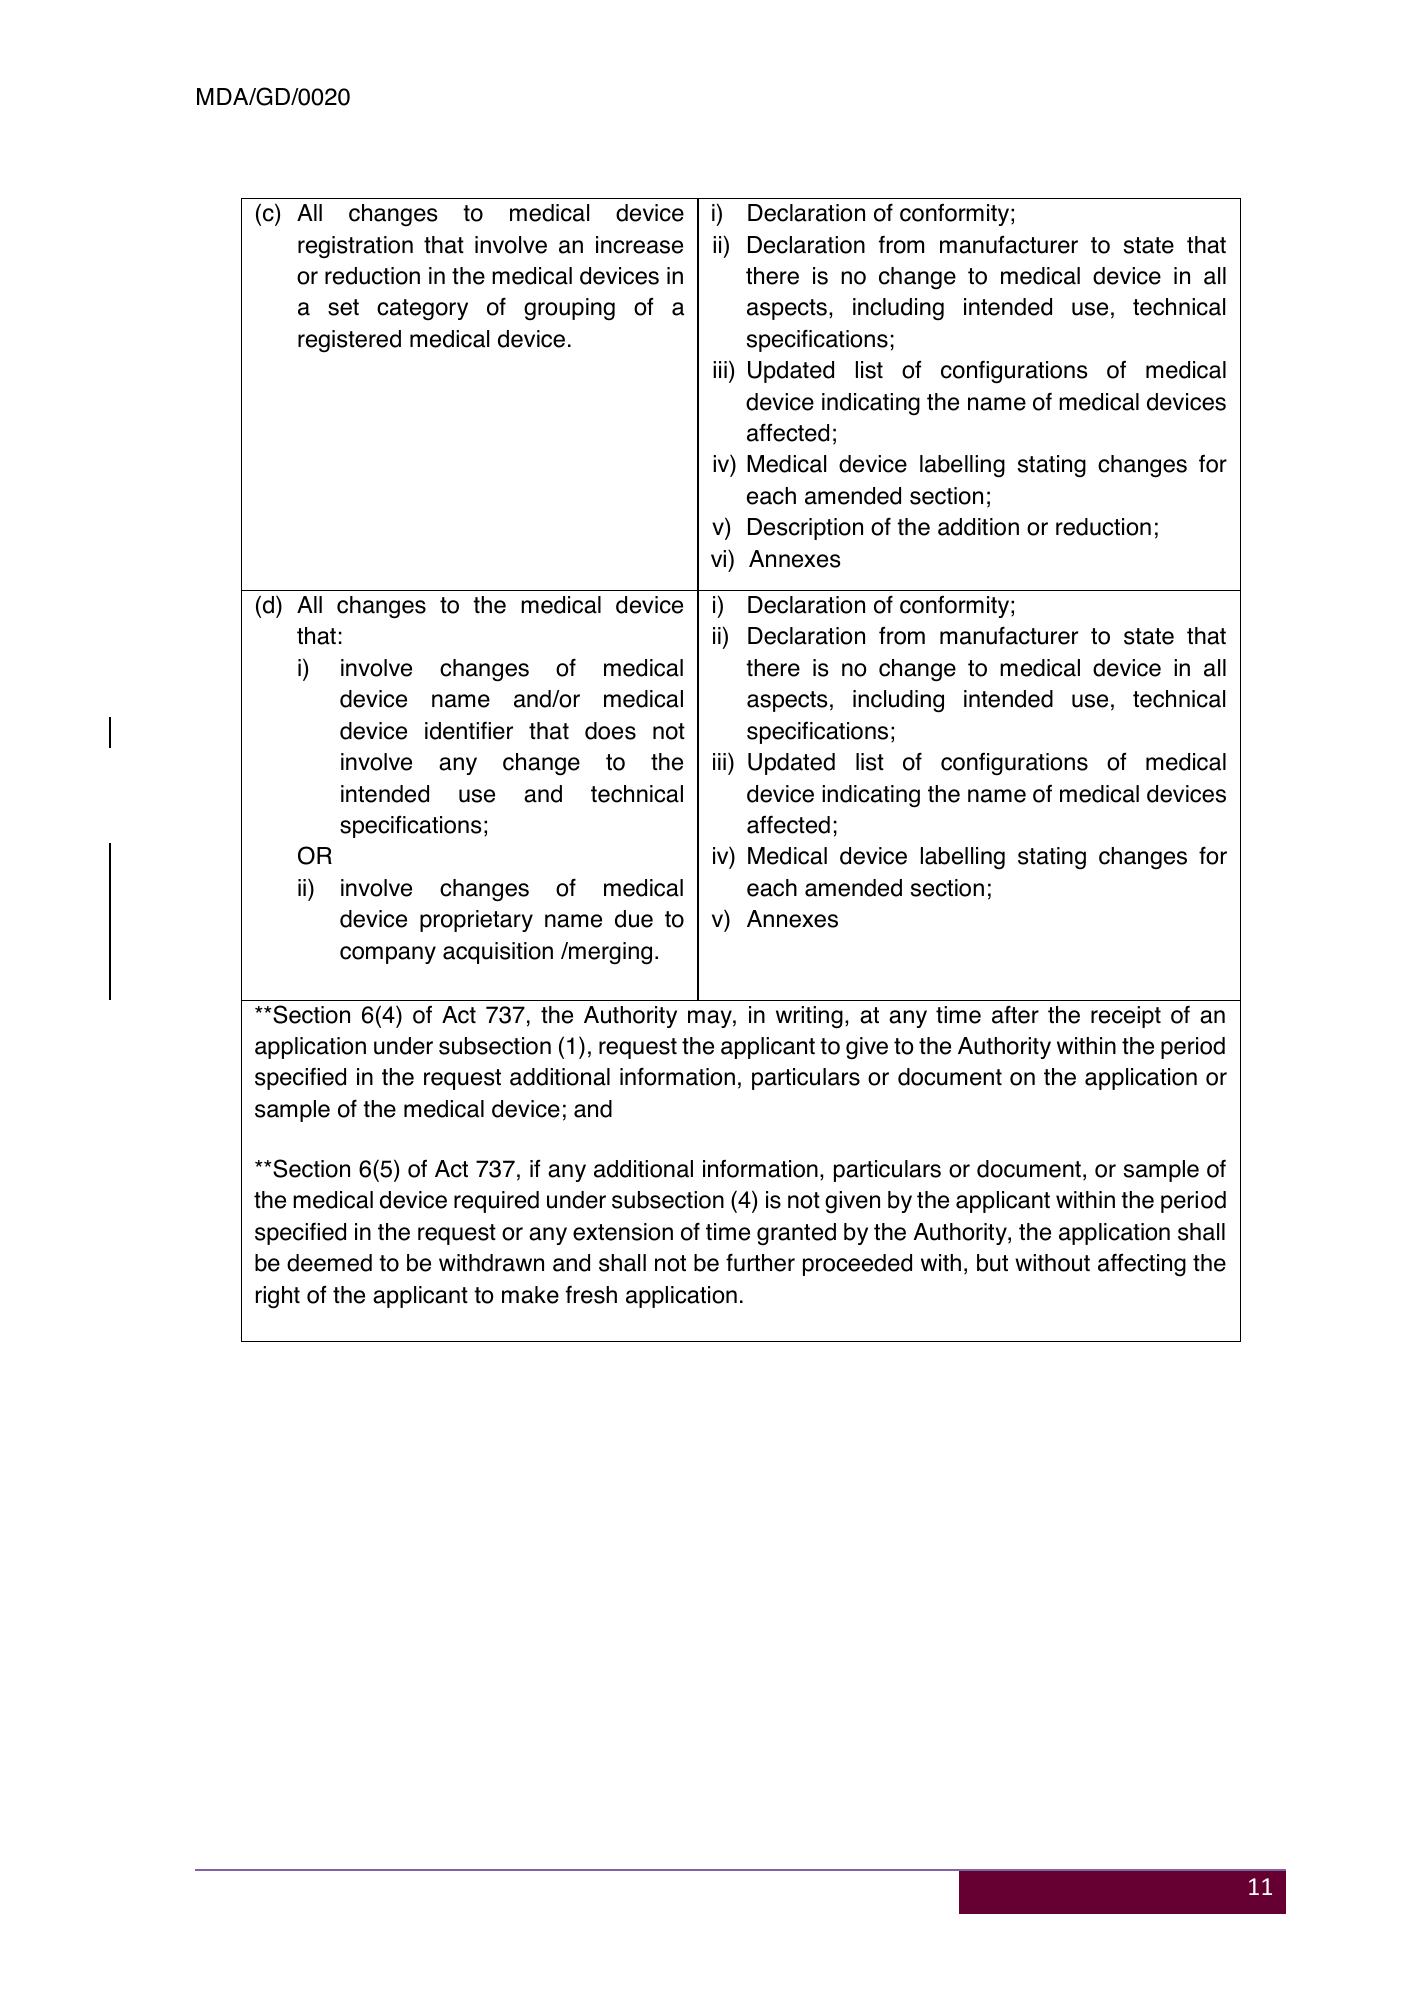
MDA/GD/0020 (c) All changes to medical device i) Declaration of conformity; registration that involve an increase ii) Declaration from manufacturer to state that or reduction in the medical devices in there is no change to medical device in all a set category of grouping of a aspects, including intended use, technical registered medical device. specifications; iii) Updated list of configurations of medical device indicating the name of medical devices affected; iv) Medical device labelling stating changes for each amended section; v) Description of the addition or reduction; vi) Annexes (d) All changes to the medical device i) Declaration of conformity; that: ii) Declaration from manufacturer to state that i) involve changes of medical there is no change to medical device in all device name and/or medical aspects, including intended use, technical device identifier that does not specifications; involve any change to the iii) Updated list of configurations of medical intended use and technical device indicating the name of medical devices specifications; affected; OR iv) Medical device labelling stating changes for ii) involve changes of medical each amended section; device proprietary name due to v) Annexes company acquisition /merging. **Section 6(4) of Act 737, the Authority may, in writing, at any time after the receipt of an application under subsection (1), request the applicant to give to the Authority within the period specified in the request additional information, particulars or document on the application or sample of the medical device; and **Section 6(5) of Act 737, if any additional information, particulars or document, or sample of the medical device required under subsection (4) is not given by the applicant within the period specified in the request or any extension of time granted by the Authority, the application shall be deemed to be withdrawn and shall not be further proceeded with, but without affecting the right of the applicant to make fresh application. 11
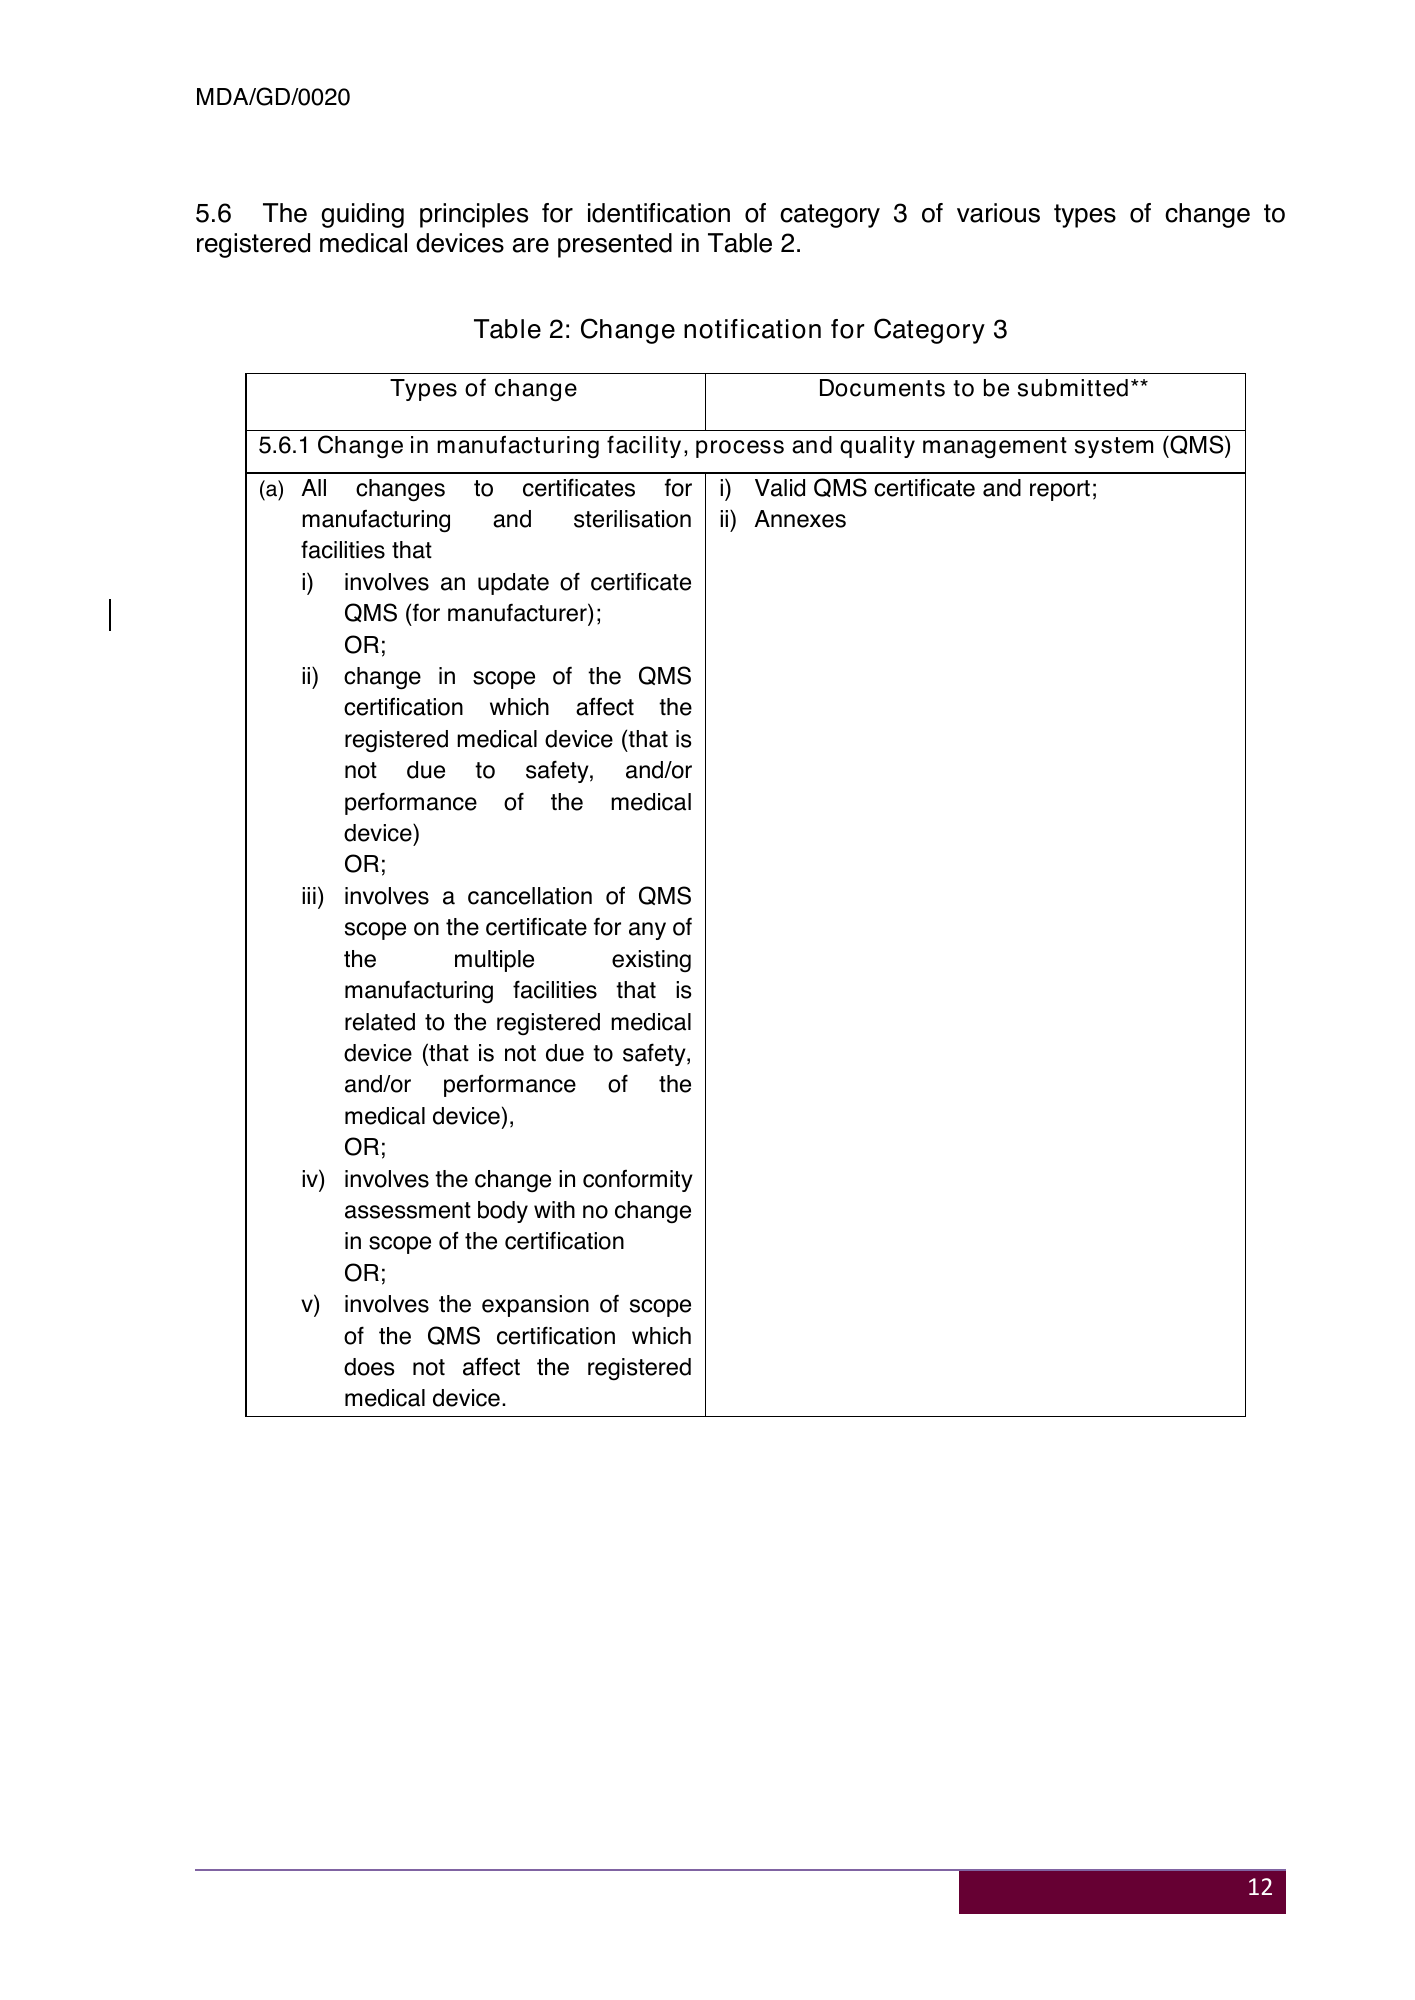
MDA/GD/0020 5.6 The guiding principles for identification of category 3 of various types of change to registered medical devices are presented in Table 2. Table 2: Change notification for Category 3 Types of change Documents to be submitted** 5.6.1 Change in manufacturing facility, process and quality management system (QMS) (a) All changes to certificates for i) Valid QMS certificate and report; manufacturing and sterilisation ii) Annexes facilities that i) involves an update of certificate QMS (for manufacturer); OR; ii) change in scope of the QMS certification which affect the registered medical device (that is not due to safety, and/or performance of the medical device) OR; iii) involves a cancellation of QMS scope on the certificate for any of the multiple existing manufacturing facilities that is related to the registered medical device (that is not due to safety, and/or performance of the medical device), OR; iv) involves the change in conformity assessment body with no change in scope of the certification OR; v) involves the expansion of scope of the QMS certification which does not affect the registered medical device. 12
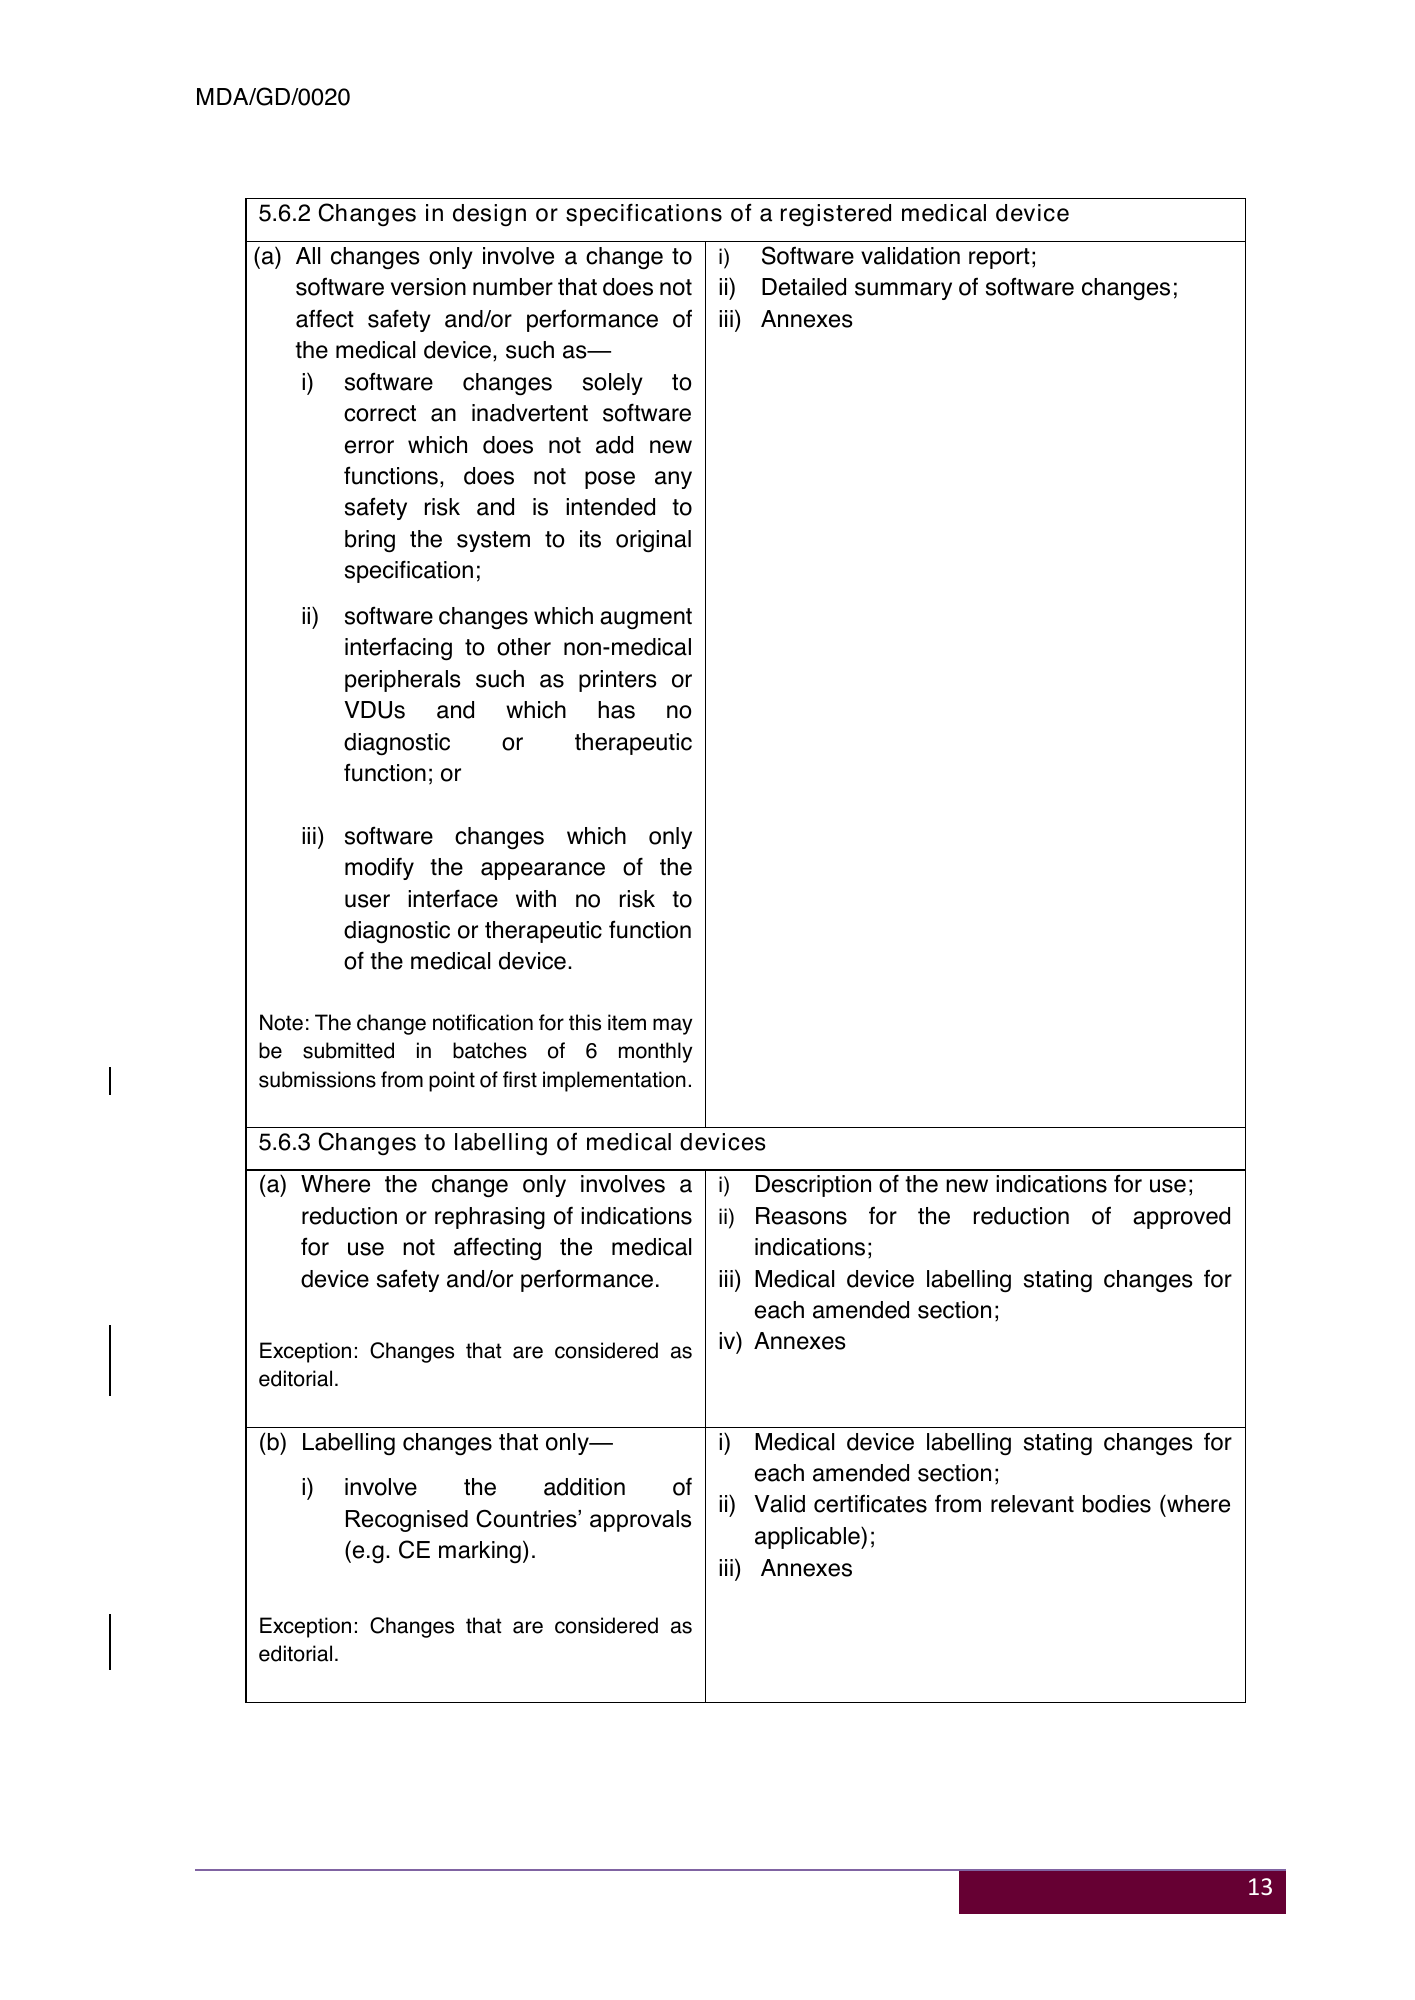
MDA/GD/0020 5.6.2 Changes in design or specifications of a registered medical device (a) All changes only involve a change to i) Software validation report; software version number that does not ii) Detailed summary of software changes; affect safety and/or performance of iii) Annexes the medical device, such as— i) software changes solely to correct an inadvertent software error which does not add new functions, does not pose any safety risk and is intended to bring the system to its original specification; ii) software changes which augment interfacing to other non-medical peripherals such as printers or VDUs and which has no diagnostic or therapeutic function; or iii) software changes which only modify the appearance of the user interface with no risk to diagnostic or therapeutic function of the medical device. Note: The change notification for this item may be submitted in batches of 6 monthly submissions from point of first implementation. 5.6.3 Changes to labelling of medical devices (a) Where the change only involves a i) Description of the new indications for use; reduction or rephrasing of indications ii) Reasons for the reduction of approved for use not affecting the medical indications; device safety and/or performance. iii) Medical device labelling stating changes for each amended section; Exception: Changes that are considered as iv) Annexes editorial. (b) Labelling changes that only— i) Medical device labelling stating changes for each amended section; i) involve the addition of ii) Valid certificates from relevant bodies (where Recognised Countries’ approvals applicable); (e.g. CE marking). iii) Annexes Exception: Changes that are considered as editorial. 13
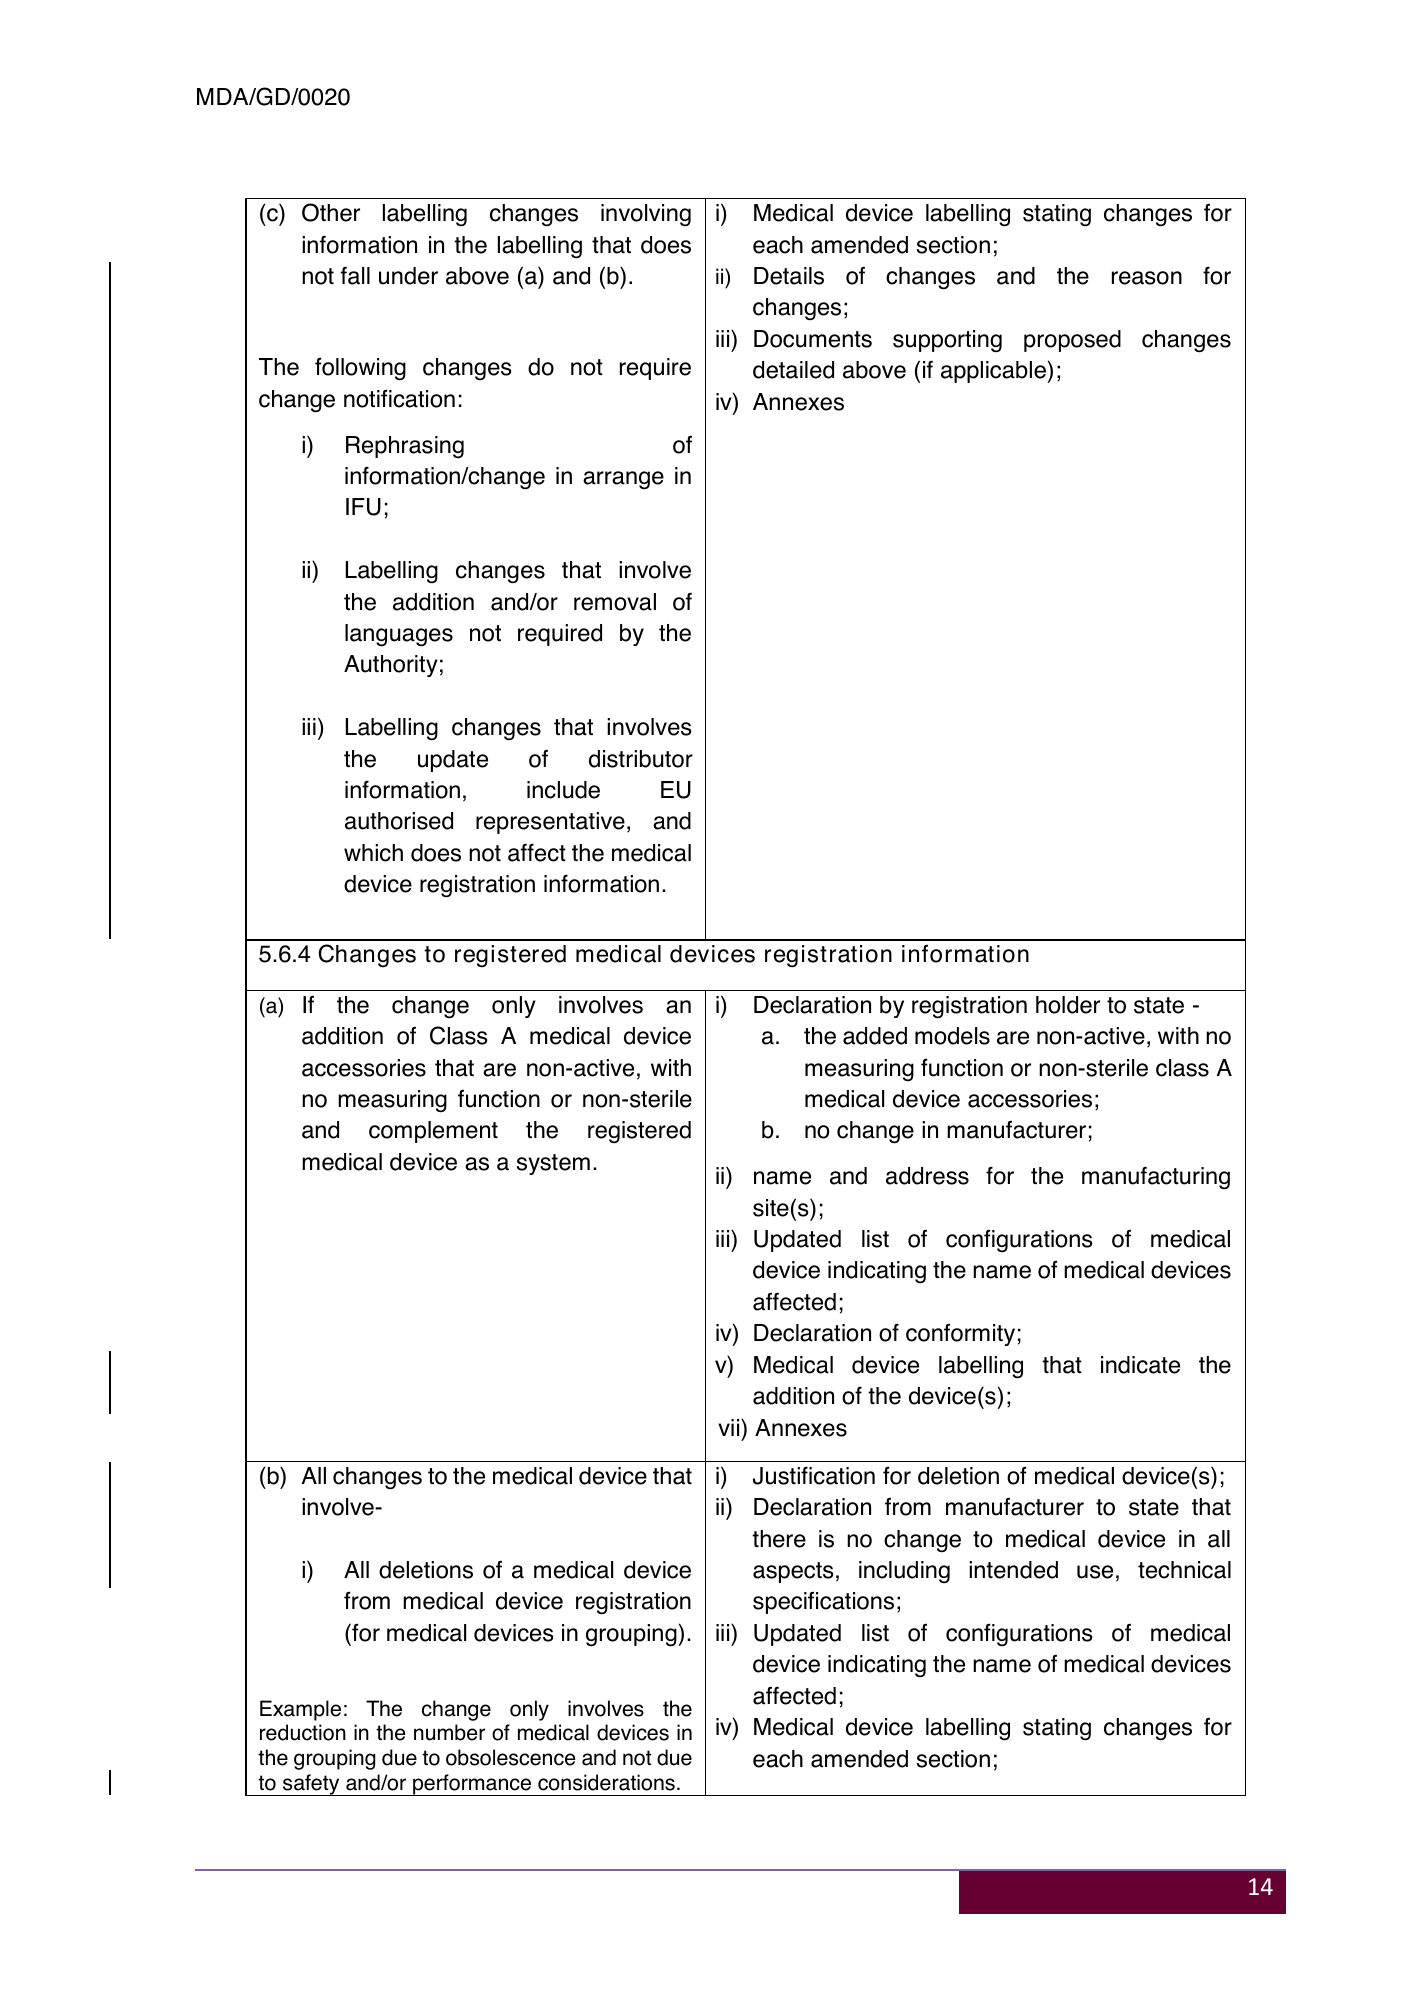
MDA/GD/0020 (c) Other labelling changes involving i) Medical device labelling stating changes for information in the labelling that does each amended section; not fall under above (a) and (b). ii) Details of changes and the reason for changes; iii) Documents supporting proposed changes The following changes do not require detailed above (if applicable); change notification: iv) Annexes i) Rephrasing of information/change in arrange in IFU; ii) Labelling changes that involve the addition and/or removal of languages not required by the Authority; iii) Labelling changes that involves the update of distributor information, include EU authorised representative, and which does not affect the medical device registration information. 5.6.4 Changes to registered medical devices registration information (a) If the change only involves an i) Declaration by registration holder to state - addition of Class A medical device a. the added models are non-active, with no accessories that are non-active, with measuring function or non-sterile class A no measuring function or non-sterile medical device accessories; and complement the registered b. no change in manufacturer; medical device as a system. ii) name and address for the manufacturing site(s); iii) Updated list of configurations of medical device indicating the name of medical devices affected; iv) Declaration of conformity; v) Medical device labelling that indicate the addition of the device(s); vii) Annexes (b) All changes to the medical device that i) Justification for deletion of medical device(s); involve- ii) Declaration from manufacturer to state that there is no change to medical device in all i) All deletions of a medical device aspects, including intended use, technical from medical device registration specifications; (for medical devices in grouping). iii) Updated list of configurations of medical device indicating the name of medical devices affected; Example: The change only involves the reduction in the number of medical devices in iv) Medical device labelling stating changes for the grouping due to obsolescence and not due each amended section; to safety and/or performance considerations. 14
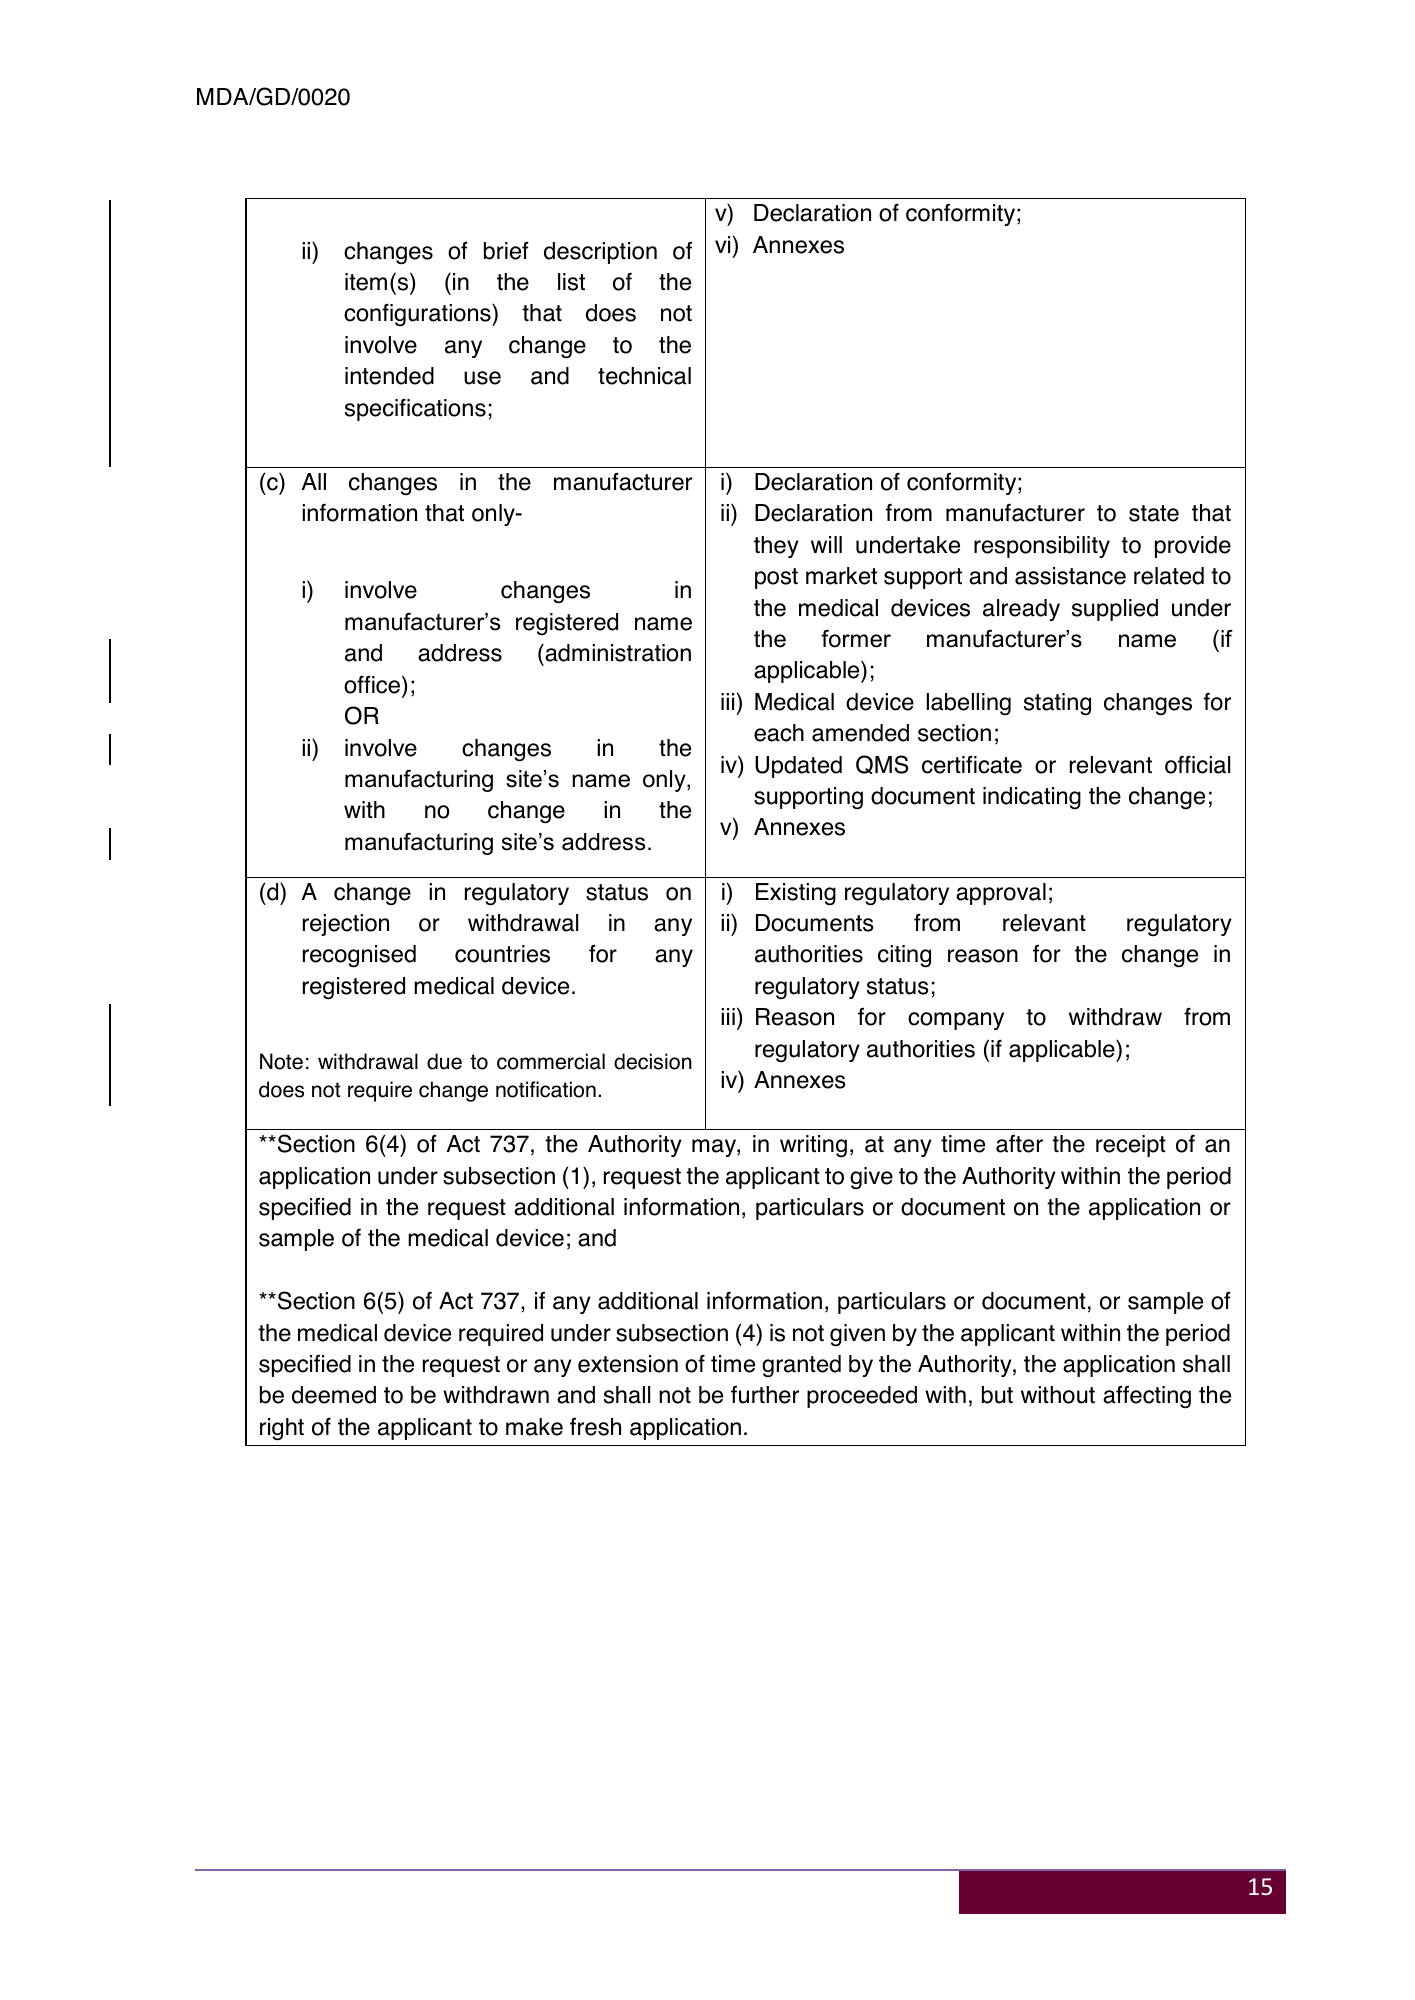
MDA/GD/0020 v) Declaration of conformity; ii) changes of brief description of vi) Annexes item(s) (in the list of the configurations) that does not involve any change to the intended use and technical specifications; (c) All changes in the manufacturer i) Declaration of conformity; information that only- ii) Declaration from manufacturer to state that they will undertake responsibility to provide post market support and assistance related to i) involve changes in the medical devices already supplied under manufacturer’s registered name the former manufacturer’s name (if and address (administration applicable); office); iii) Medical device labelling stating changes for OR each amended section; ii) involve changes in the iv) Updated QMS certificate or relevant official manufacturing site’s name only, supporting document indicating the change; with no change in the v) Annexes manufacturing site’s address. (d) A change in regulatory status on i) Existing regulatory approval; rejection or withdrawal in any ii) Documents from relevant regulatory recognised countries for any authorities citing reason for the change in registered medical device. regulatory status; iii) Reason for company to withdraw from regulatory authorities (if applicable); Note: withdrawal due to commercial decision does not require change notification. iv) Annexes **Section 6(4) of Act 737, the Authority may, in writing, at any time after the receipt of an application under subsection (1), request the applicant to give to the Authority within the period specified in the request additional information, particulars or document on the application or sample of the medical device; and **Section 6(5) of Act 737, if any additional information, particulars or document, or sample of the medical device required under subsection (4) is not given by the applicant within the period specified in the request or any extension of time granted by the Authority, the application shall be deemed to be withdrawn and shall not be further proceeded with, but without affecting the right of the applicant to make fresh application. 15
MDA/GD/0020 5.7 Changes to medical devices due to a mandatory reportable incident and/or field corrective action Changes to medical devices may arise from the occurrence of mandatory reportable incidents and/or field corrective actions (refer Section 40 and Section 41 of Act 737). The proposed changes to the medical device in these situations may have an impact on the safety and/or performance of the medical device. For any changes, the establishments shall submit a change notification based on Clause 5 for the purpose of FCA submission. Refer to MDA/GD/0013, Field Corrective Action (FCA) for further information. The determination of the category of change for notifications in the context of, or as a consequence of or arising from mandatory reportable incidents and/or field corrective actions shall be based on Clause 5. 16
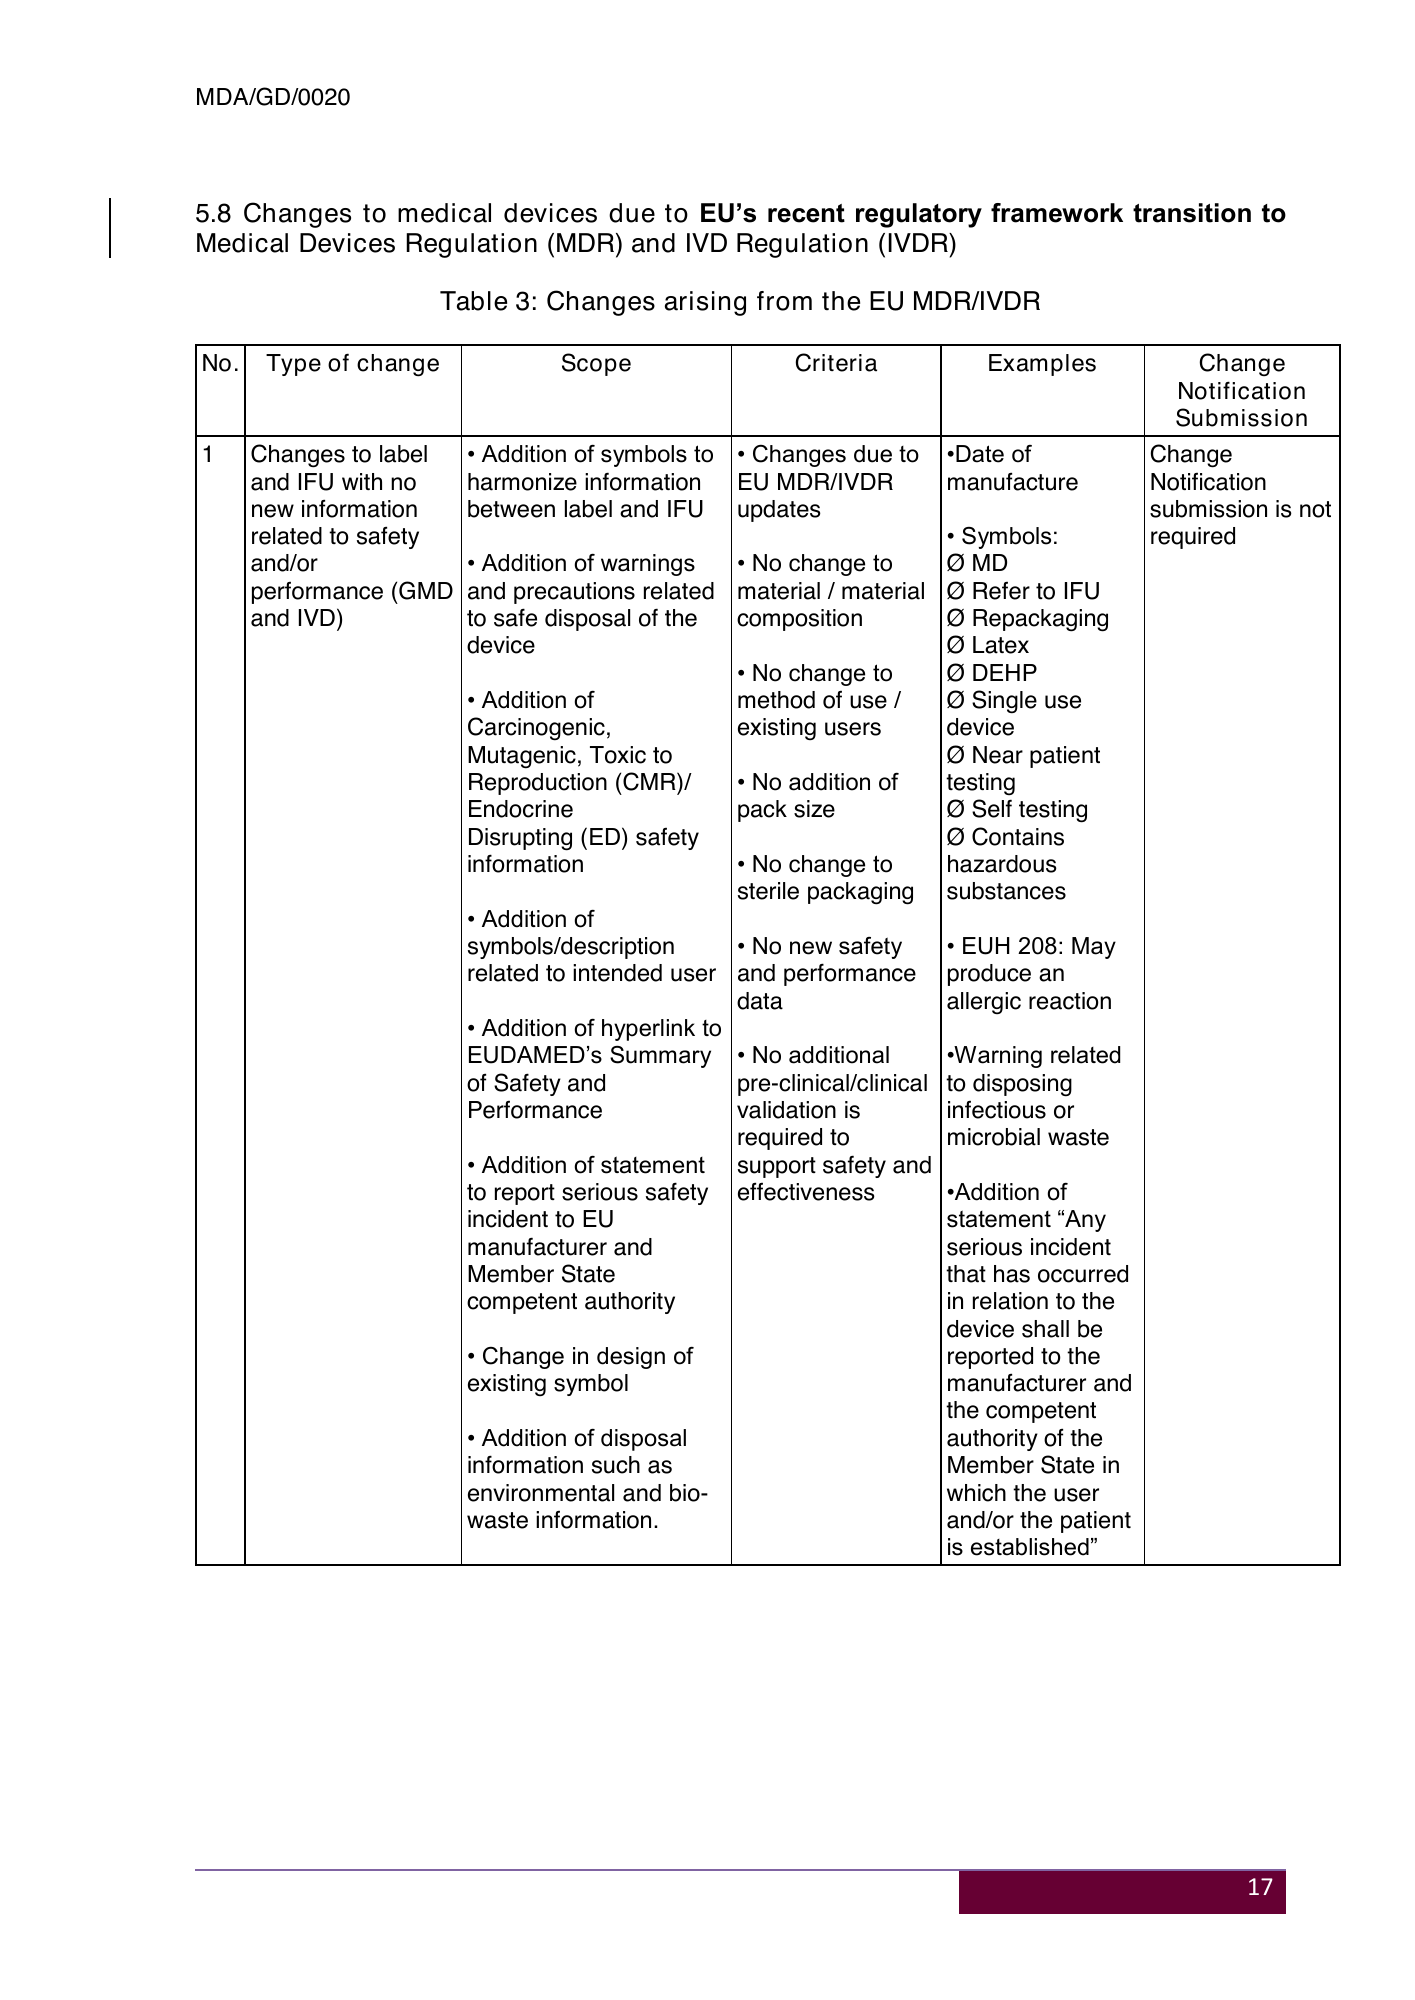
MDA/GD/0020 5.8 Changes to medical devices due to EU’s recent regulatory framework transition to Medical Devices Regulation (MDR) and IVD Regulation (IVDR) Table 3: Changes arising from the EU MDR/IVDR No. Type of change Scope Criteria Examples Change Notification Submission 1 Changes to label • Addition of symbols to • Changes due to •Date of Change and IFU with no harmonize information EU MDR/IVDR manufacture Notification new information between label and IFU updates submission is not related to safety • Symbols: required and/or • Addition of warnings • No change to Ø MD performance (GMD and precautions related material / material Ø Refer to IFU and IVD) to safe disposal of the composition Ø Repackaging device Ø Latex • No change to Ø DEHP • Addition of method of use / Ø Single use Carcinogenic, existing users device Mutagenic, Toxic to Ø Near patient Reproduction (CMR)/ • No addition of testing Endocrine pack size Ø Self testing Disrupting (ED) safety Ø Contains information • No change to hazardous sterile packaging substances • Addition of symbols/description • No new safety • EUH 208: May related to intended user and performance produce an data allergic reaction • Addition of hyperlink to EUDAMED’s Summary • No additional •Warning related of Safety and pre-clinical/clinical to disposing Performance validation is infectious or required to microbial waste • Addition of statement support safety and to report serious safety effectiveness •Addition of incident to EU statement “Any manufacturer and serious incident Member State that has occurred competent authority in relation to the device shall be • Change in design of reported to the existing symbol manufacturer and the competent • Addition of disposal authority of the information such as Member State in environmental and bio- which the user waste information. and/or the patient is established” 17
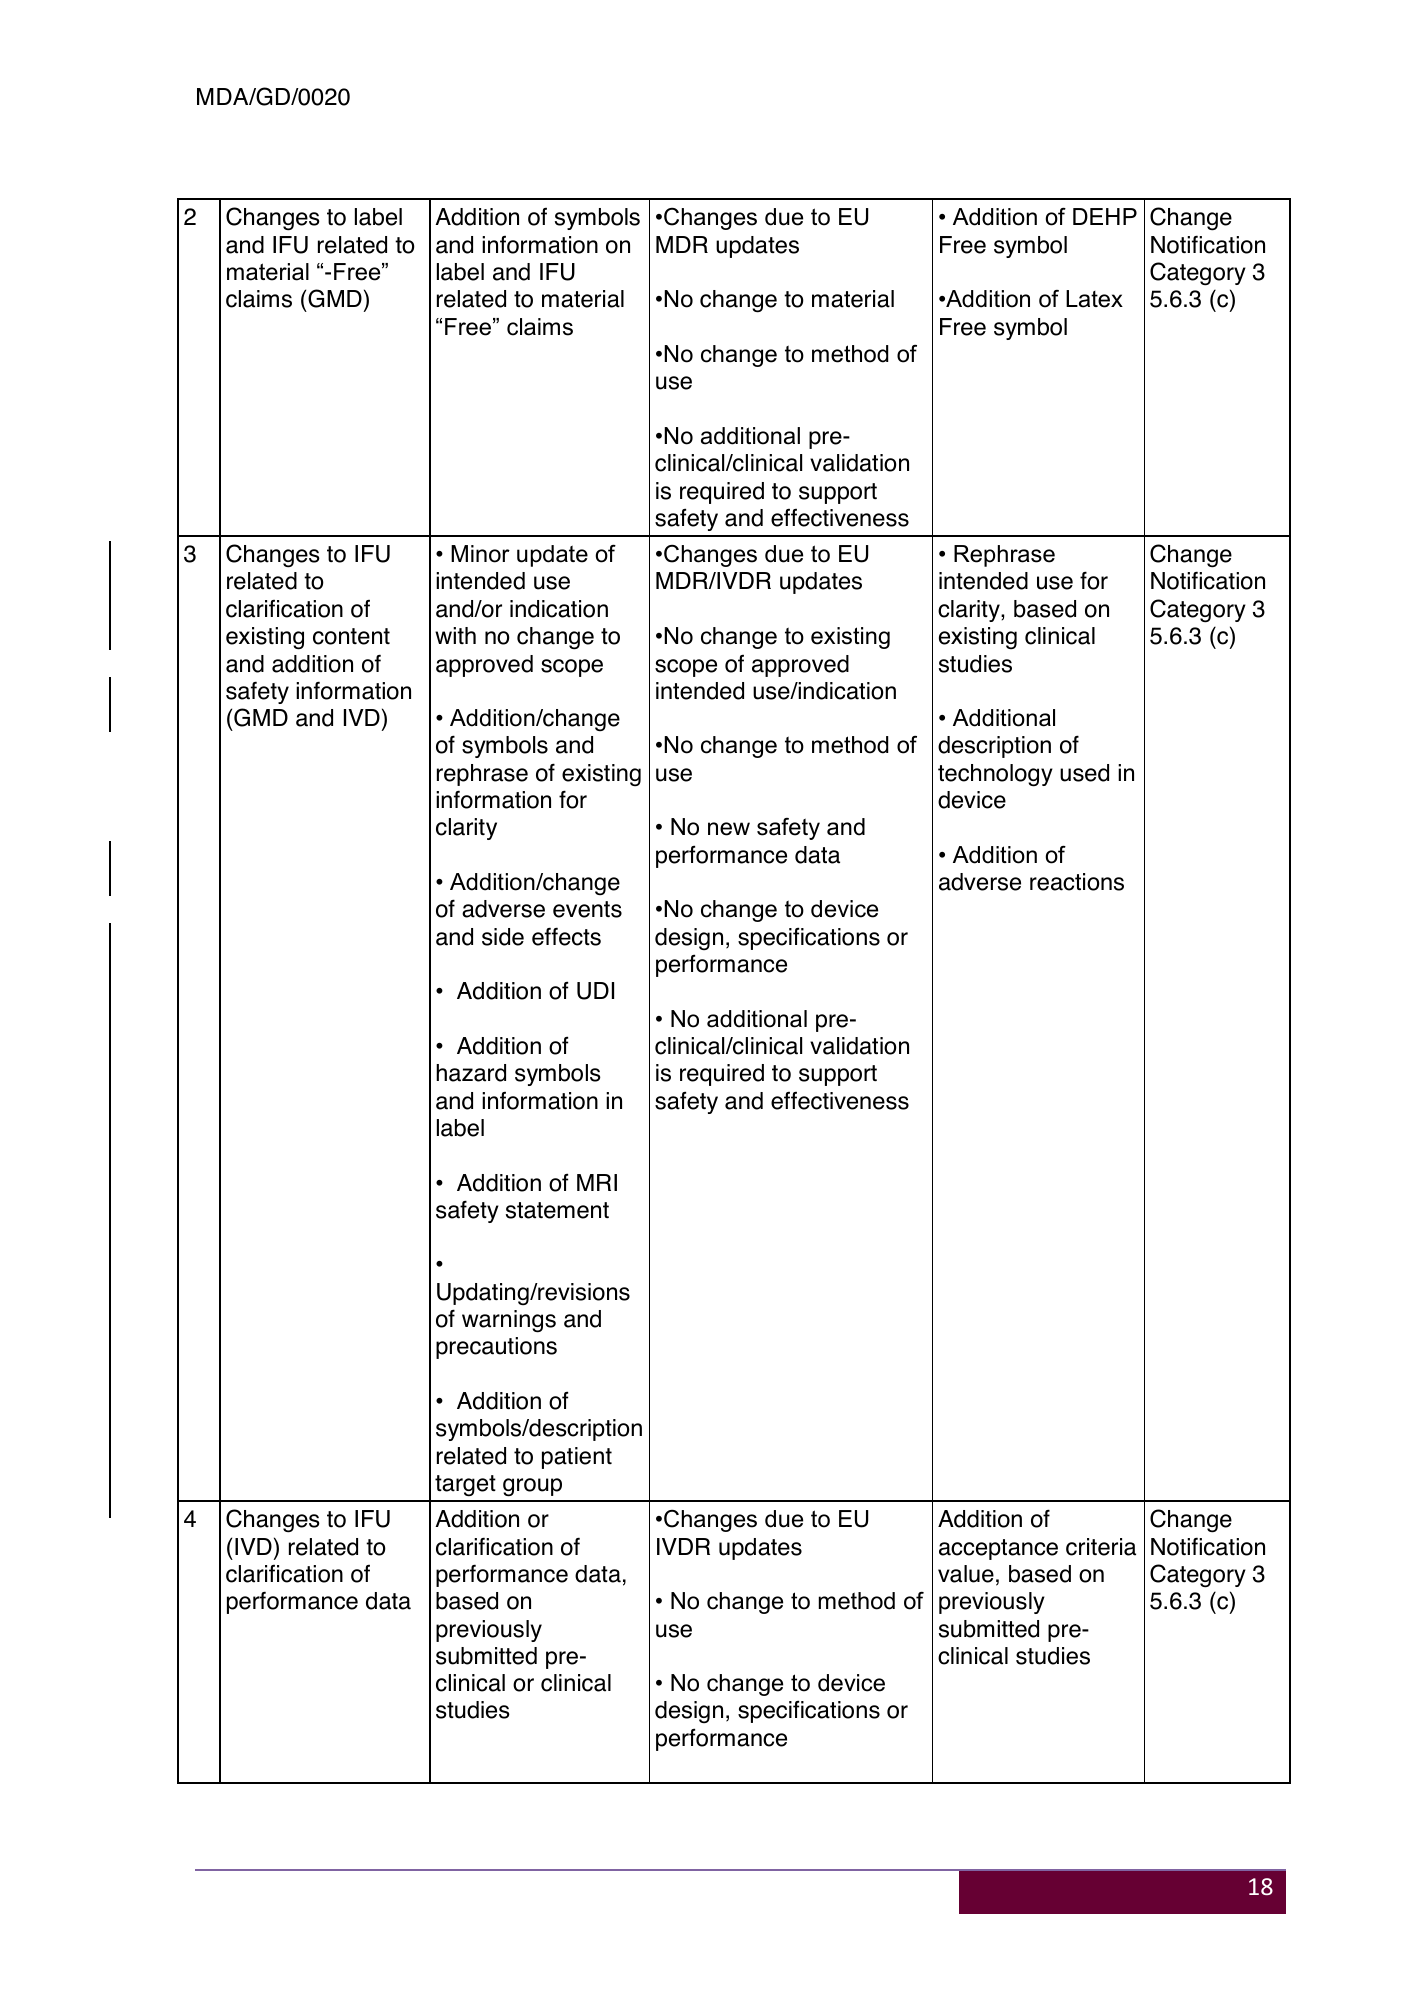
MDA/GD/0020 2 Changes to label Addition of symbols •Changes due to EU • Addition of DEHP Change and IFU related to and information on MDR updates Free symbol Notification material “-Free” label and IFU Category 3 claims (GMD) related to material •No change to material •Addition of Latex 5.6.3 (c) “Free” claims Free symbol •No change to method of use •No additional pre- clinical/clinical validation is required to support safety and effectiveness 3 Changes to IFU • Minor update of •Changes due to EU • Rephrase Change related to intended use MDR/IVDR updates intended use for Notification clarification of and/or indication clarity, based on Category 3 existing content with no change to •No change to existing existing clinical 5.6.3 (c) and addition of approved scope scope of approved studies safety information intended use/indication (GMD and IVD) • Addition/change • Additional of symbols and •No change to method of description of rephrase of existing use technology used in information for device clarity • No new safety and performance data • Addition of • Addition/change adverse reactions of adverse events •No change to device and side effects design, specifications or performance • Addition of UDI • No additional pre- • Addition of clinical/clinical validation hazard symbols is required to support and information in safety and effectiveness label • Addition of MRI safety statement • Updating/revisions of warnings and precautions • Addition of symbols/description related to patient target group 4 Changes to IFU Addition or •Changes due to EU Addition of Change (IVD) related to clarification of IVDR updates acceptance criteria Notification clarification of performance data, value, based on Category 3 performance data based on • No change to method of previously 5.6.3 (c) previously use submitted pre- submitted pre- clinical studies clinical or clinical • No change to device studies design, specifications or performance 18
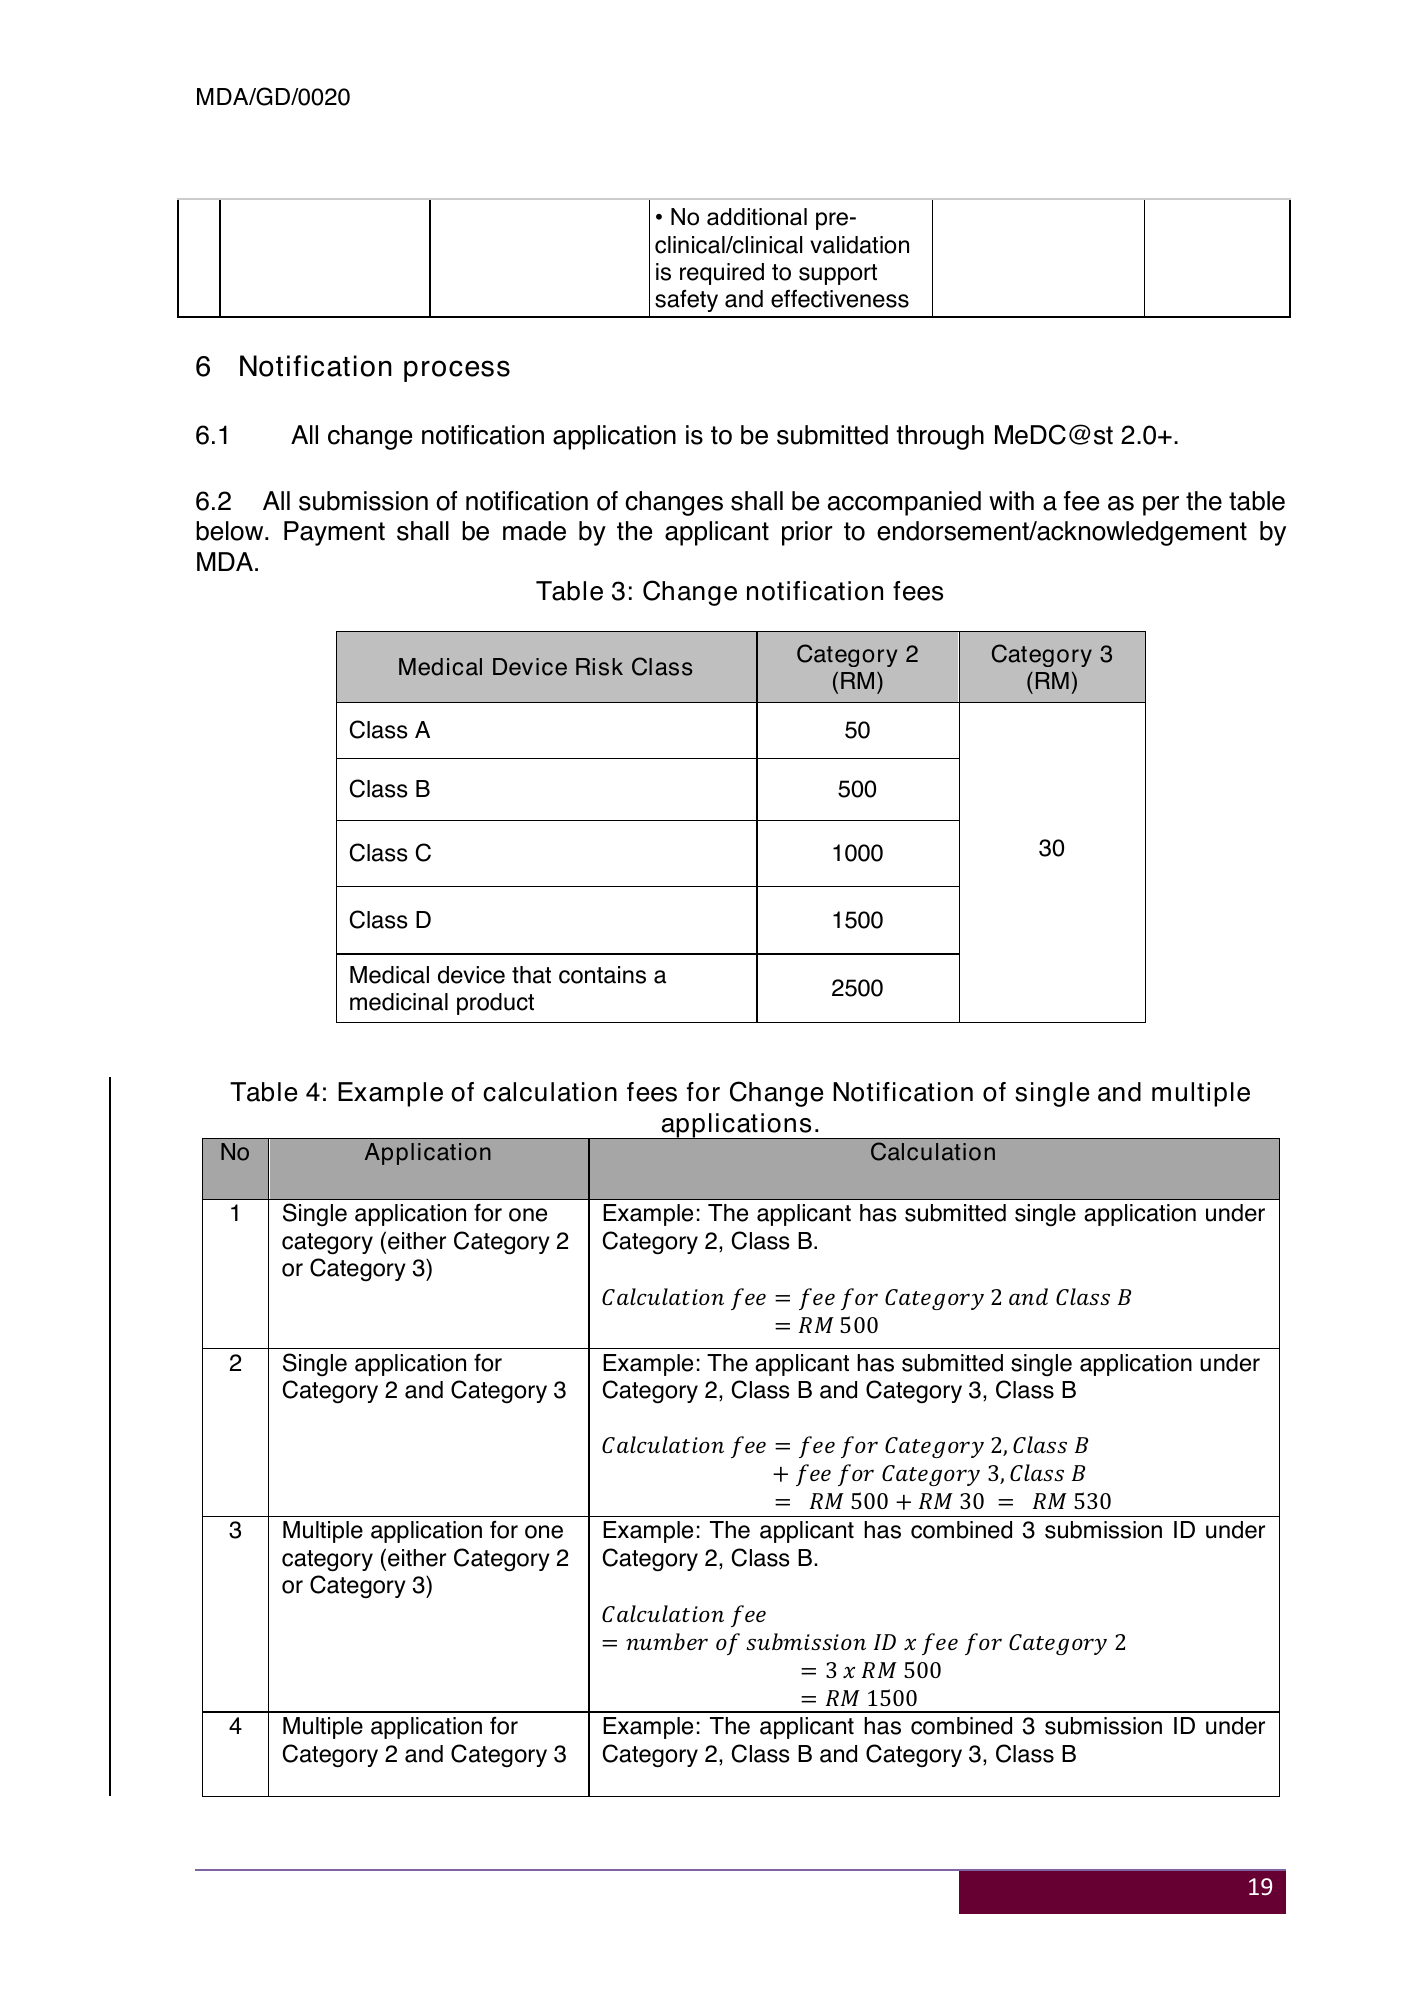
MDA/GD/0020 • No additional pre- clinical/clinical validation is required to support safety and effectiveness 6 Notification process 6.1 All change notification application is to be submitted through MeDC@st 2.0+. 6.2 All submission of notification of changes shall be accompanied with a fee as per the table below. Payment shall be made by the applicant prior to endorsement/acknowledgement by MDA. Table 3: Change notification fees Category 2 Category 3 Medical Device Risk Class (RM) (RM) Class A 50 Class B 500 Class C 1000 30 Class D 1500 Medical device that contains a 2500 medicinal product Table 4: Example of calculation fees for Change Notification of single and multiple applications. No Application Calculation 1 Single application for one Example: The applicant has submitted single application under category (either Category 2 Category 2, Class B. or Category 3) 𝐶𝑎𝑙𝑐𝑢𝑙𝑎𝑡𝑖𝑜𝑛 𝑓𝑒𝑒 = 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 2 𝑎𝑛𝑑 𝐶𝑙𝑎𝑠𝑠 𝐵 = 𝑅𝑀 500 2 Single application for Example: The applicant has submitted single application under Category 2 and Category 3 Category 2, Class B and Category 3, Class B 𝐶𝑎𝑙𝑐𝑢𝑙𝑎𝑡𝑖𝑜𝑛 𝑓𝑒𝑒 = 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 2, 𝐶𝑙𝑎𝑠𝑠 𝐵 + 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 3, 𝐶𝑙𝑎𝑠𝑠 𝐵 = 𝑅𝑀 500 + 𝑅𝑀 30 = 𝑅𝑀 530 3 Multiple application for one Example: The applicant has combined 3 submission ID under category (either Category 2 Category 2, Class B. or Category 3) 𝐶𝑎𝑙𝑐𝑢𝑙𝑎𝑡𝑖𝑜𝑛 𝑓𝑒𝑒 = 𝑛𝑢𝑚𝑏𝑒𝑟 𝑜𝑓 𝑠𝑢𝑏𝑚𝑖𝑠𝑠𝑖𝑜𝑛 𝐼𝐷 𝑥 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 2 = 3 𝑥 𝑅𝑀 500 = 𝑅𝑀 1500 4 Multiple application for Example: The applicant has combined 3 submission ID under Category 2 and Category 3 Category 2, Class B and Category 3, Class B 19
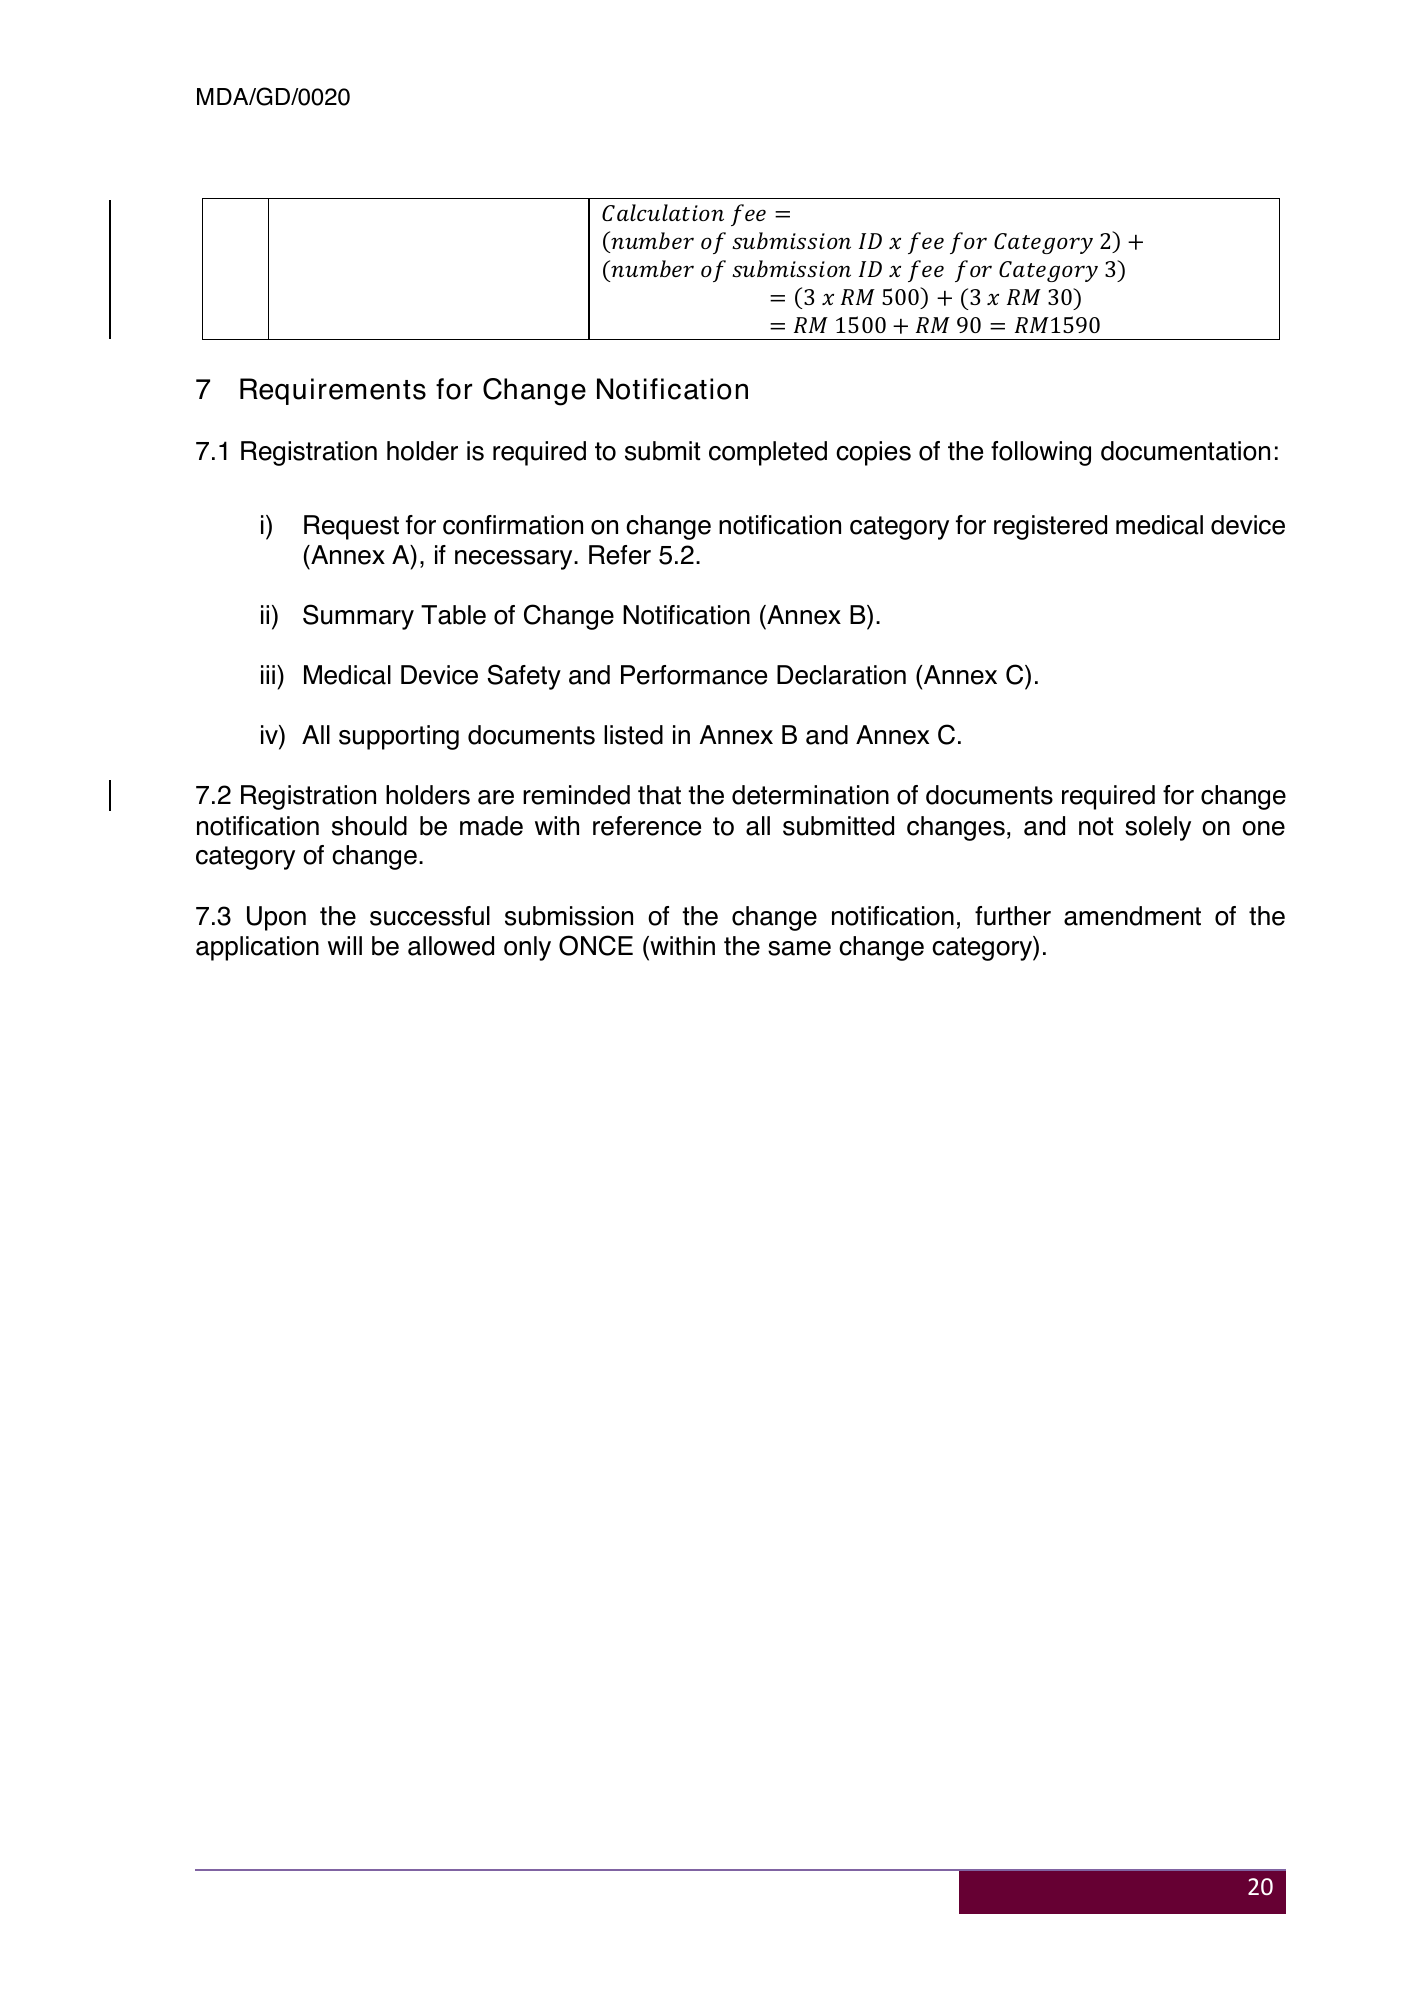
MDA/GD/0020 𝐶𝑎𝑙𝑐𝑢𝑙𝑎𝑡𝑖𝑜𝑛 𝑓𝑒𝑒 = (𝑛𝑢𝑚𝑏𝑒𝑟 𝑜𝑓 𝑠𝑢𝑏𝑚𝑖𝑠𝑠𝑖𝑜𝑛 𝐼𝐷 𝑥 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 2) + (𝑛𝑢𝑚𝑏𝑒𝑟 𝑜𝑓 𝑠𝑢𝑏𝑚𝑖𝑠𝑠𝑖𝑜𝑛 𝐼𝐷 𝑥 𝑓𝑒𝑒 𝑓𝑜𝑟 𝐶𝑎𝑡𝑒𝑔𝑜𝑟𝑦 3) = (3 𝑥 𝑅𝑀 500) + (3 𝑥 𝑅𝑀 30) = 𝑅𝑀 1500 + 𝑅𝑀 90 = 𝑅𝑀1590 7 Requirements for Change Notification 7.1 Registration holder is required to submit completed copies of the following documentation: i) Request for confirmation on change notification category for registered medical device (Annex A), if necessary. Refer 5.2. ii) Summary Table of Change Notification (Annex B). iii) Medical Device Safety and Performance Declaration (Annex C). iv) All supporting documents listed in Annex B and Annex C. 7.2 Registration holders are reminded that the determination of documents required for change notification should be made with reference to all submitted changes, and not solely on one category of change. 7.3 Upon the successful submission of the change notification, further amendment of the application will be allowed only ONCE (within the same change category). 20
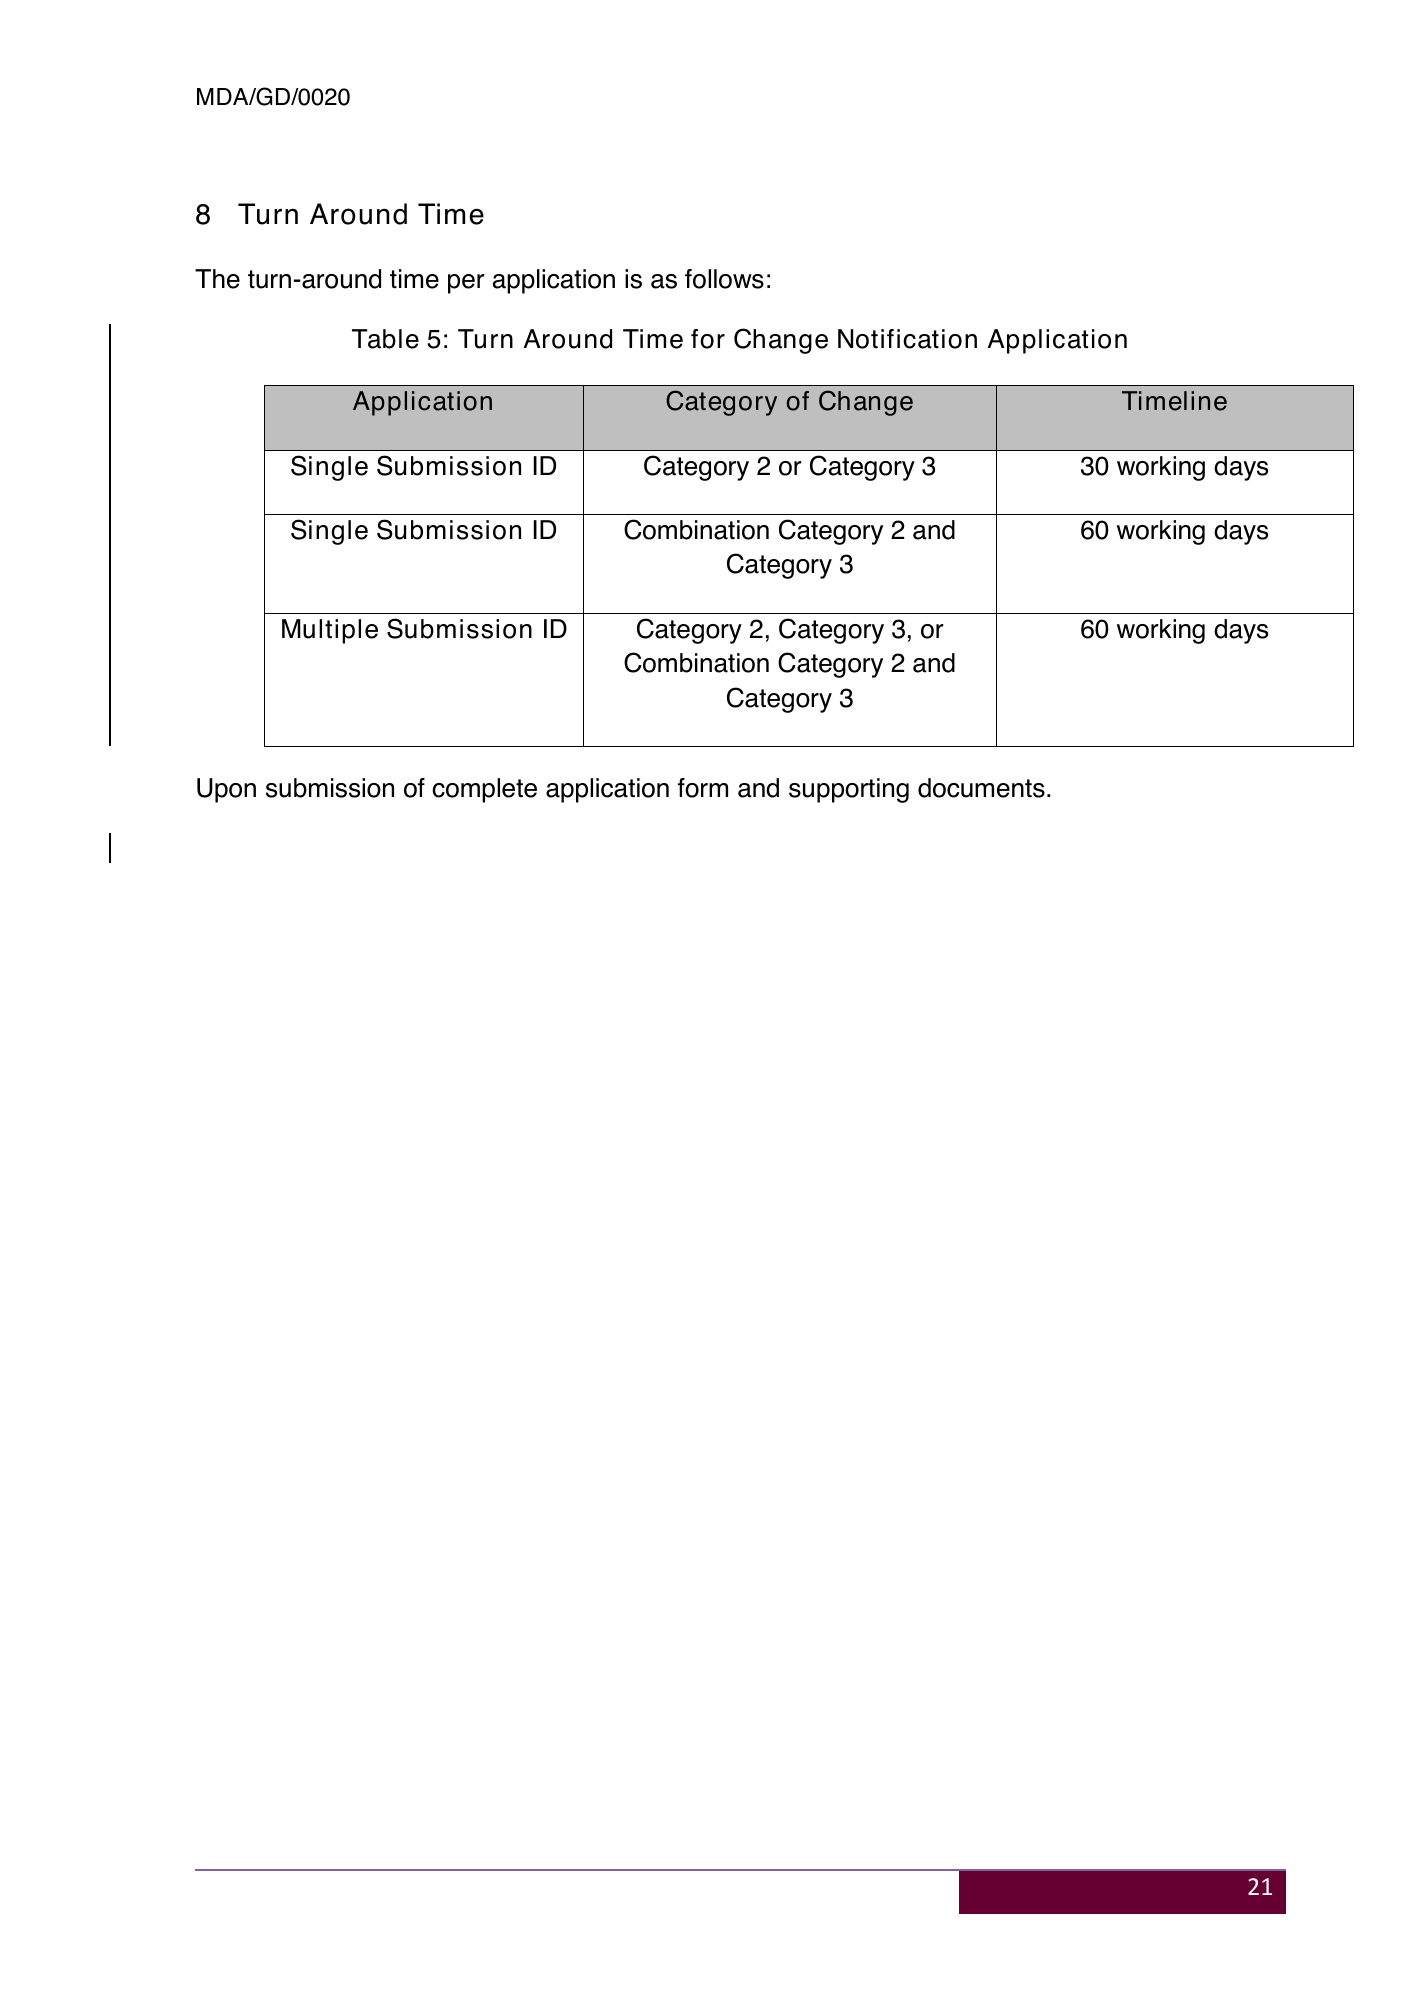
MDA/GD/0020 8 Turn Around Time The turn-around time per application is as follows: Table 5: Turn Around Time for Change Notification Application Application Category of Change Timeline Single Submission ID Category 2 or Category 3 30 working days Single Submission ID Combination Category 2 and 60 working days Category 3 Multiple Submission ID Category 2, Category 3, or 60 working days Combination Category 2 and Category 3 Upon submission of complete application form and supporting documents. 21
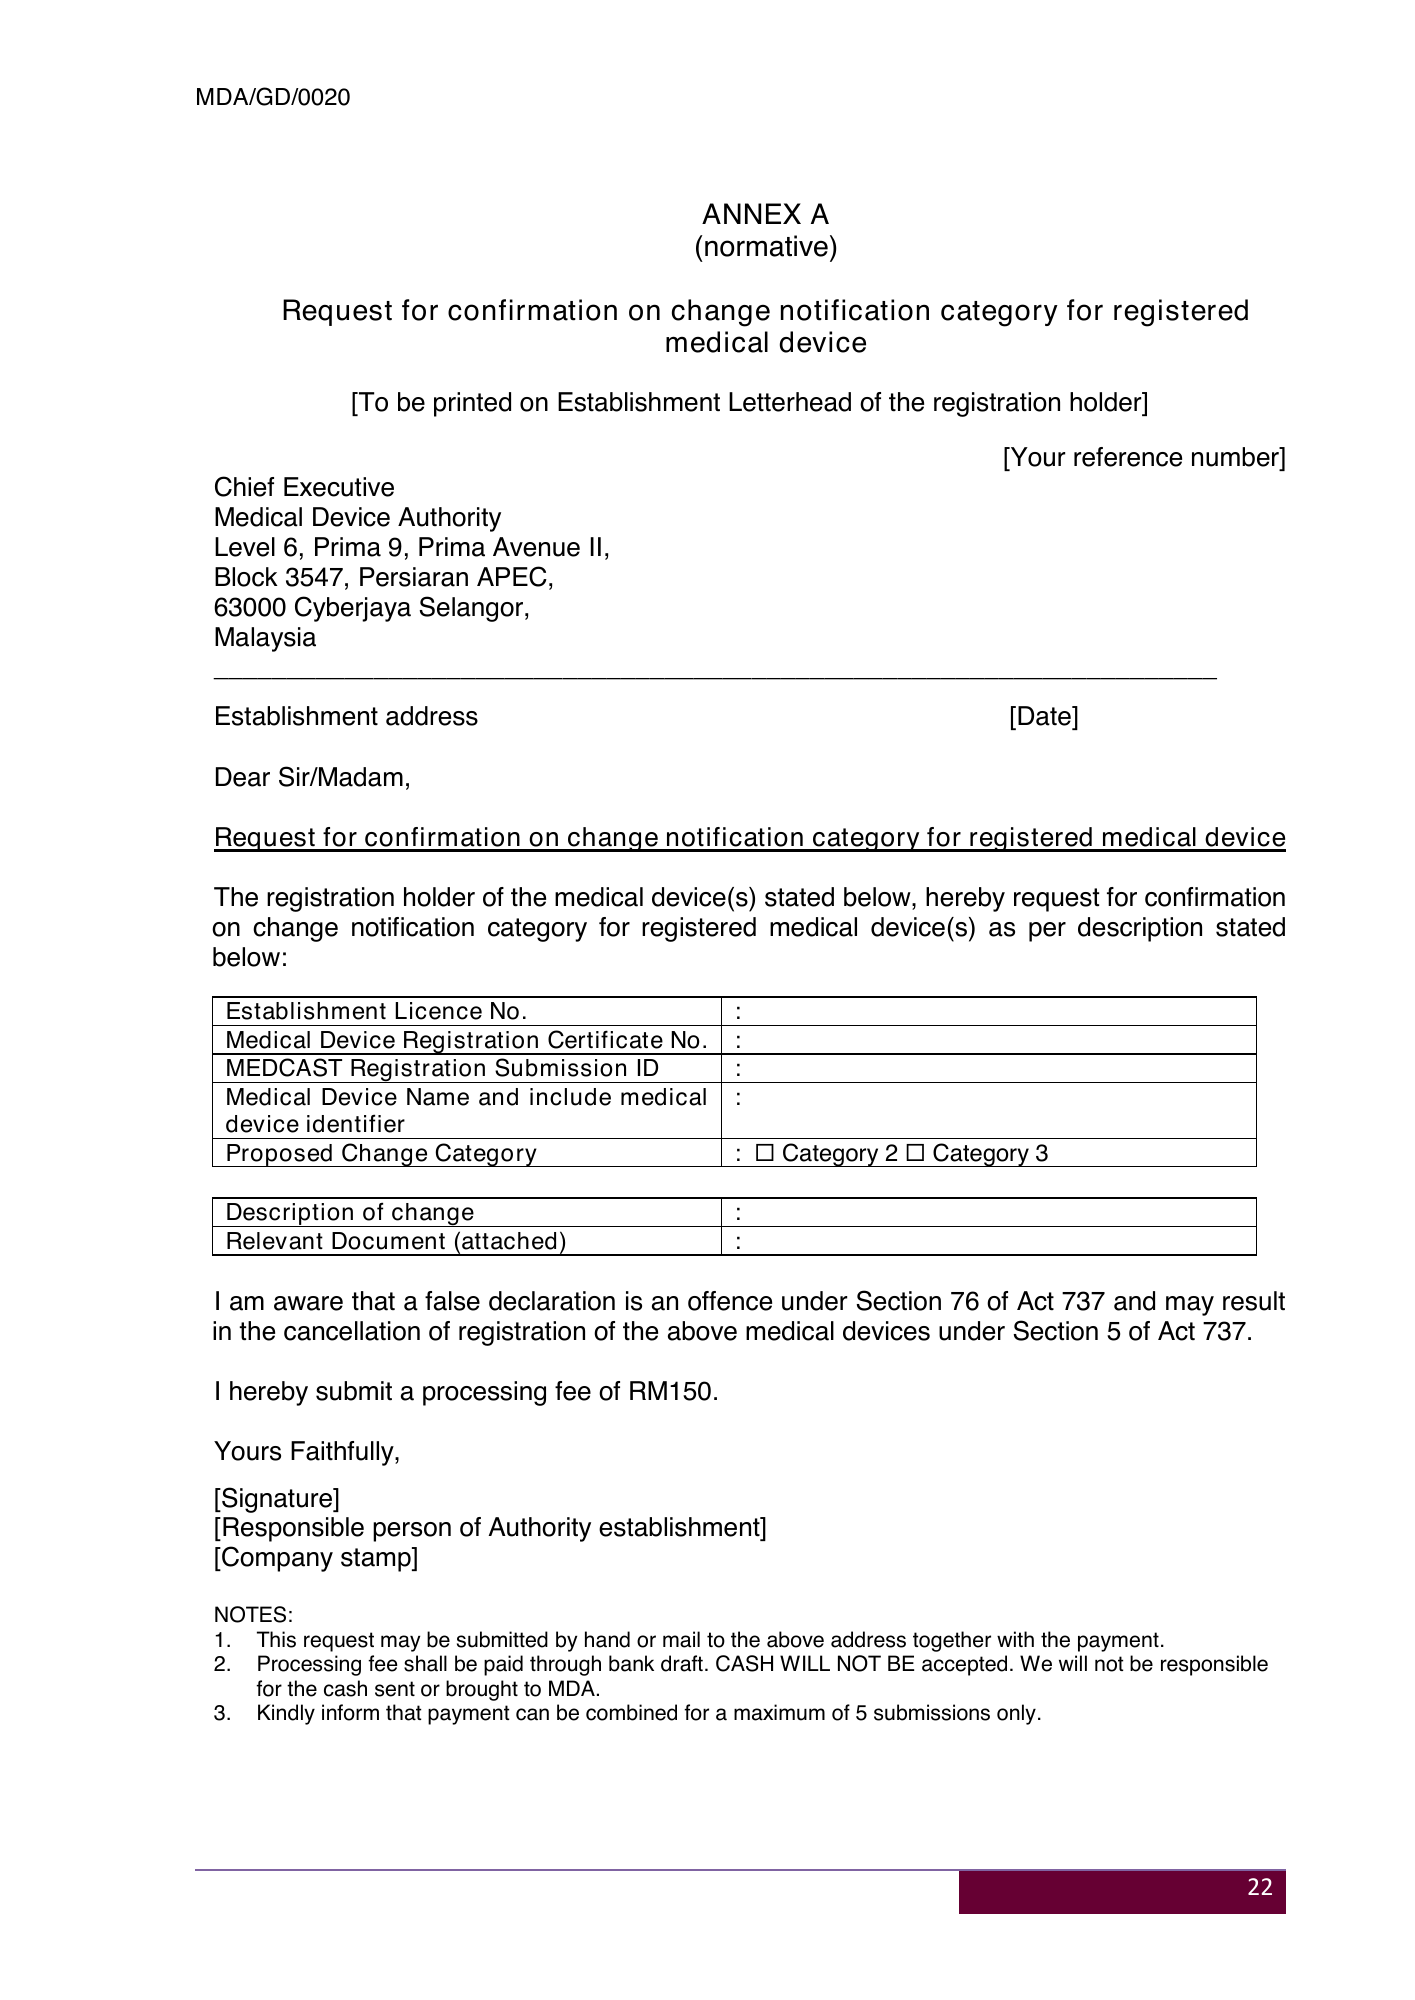
MDA/GD/0020 ANNEX A (normative) Request for confirmation on change notification category for registered medical device [To be printed on Establishment Letterhead of the registration holder] [Your reference number] Chief Executive Medical Device Authority Level 6, Prima 9, Prima Avenue II, Block 3547, Persiaran APEC, 63000 Cyberjaya Selangor, Malaysia _____________________________________________________________________ Establishment address [Date] Dear Sir/Madam, Request for confirmation on change notification category for registered medical device The registration holder of the medical device(s) stated below, hereby request for confirmation on change notification category for registered medical device(s) as per description stated below: Establishment Licence No. : Medical Device Registration Certificate No. : MEDCAST Registration Submission ID : Medical Device Name and include medical : device identifier Proposed Change Category : Category 2 Category 3 Description of change : Relevant Document (attached) : I am aware that a false declaration is an offence under Section 76 of Act 737 and may result in the cancellation of registration of the above medical devices under Section 5 of Act 737. I hereby submit a processing fee of RM150. Yours Faithfully, [Signature] [Responsible person of Authority establishment] [Company stamp] NOTES: 1. This request may be submitted by hand or mail to the above address together with the payment. 2. Processing fee shall be paid through bank draft. CASH WILL NOT BE accepted. We will not be responsible for the cash sent or brought to MDA. 3. Kindly inform that payment can be combined for a maximum of 5 submissions only. 22
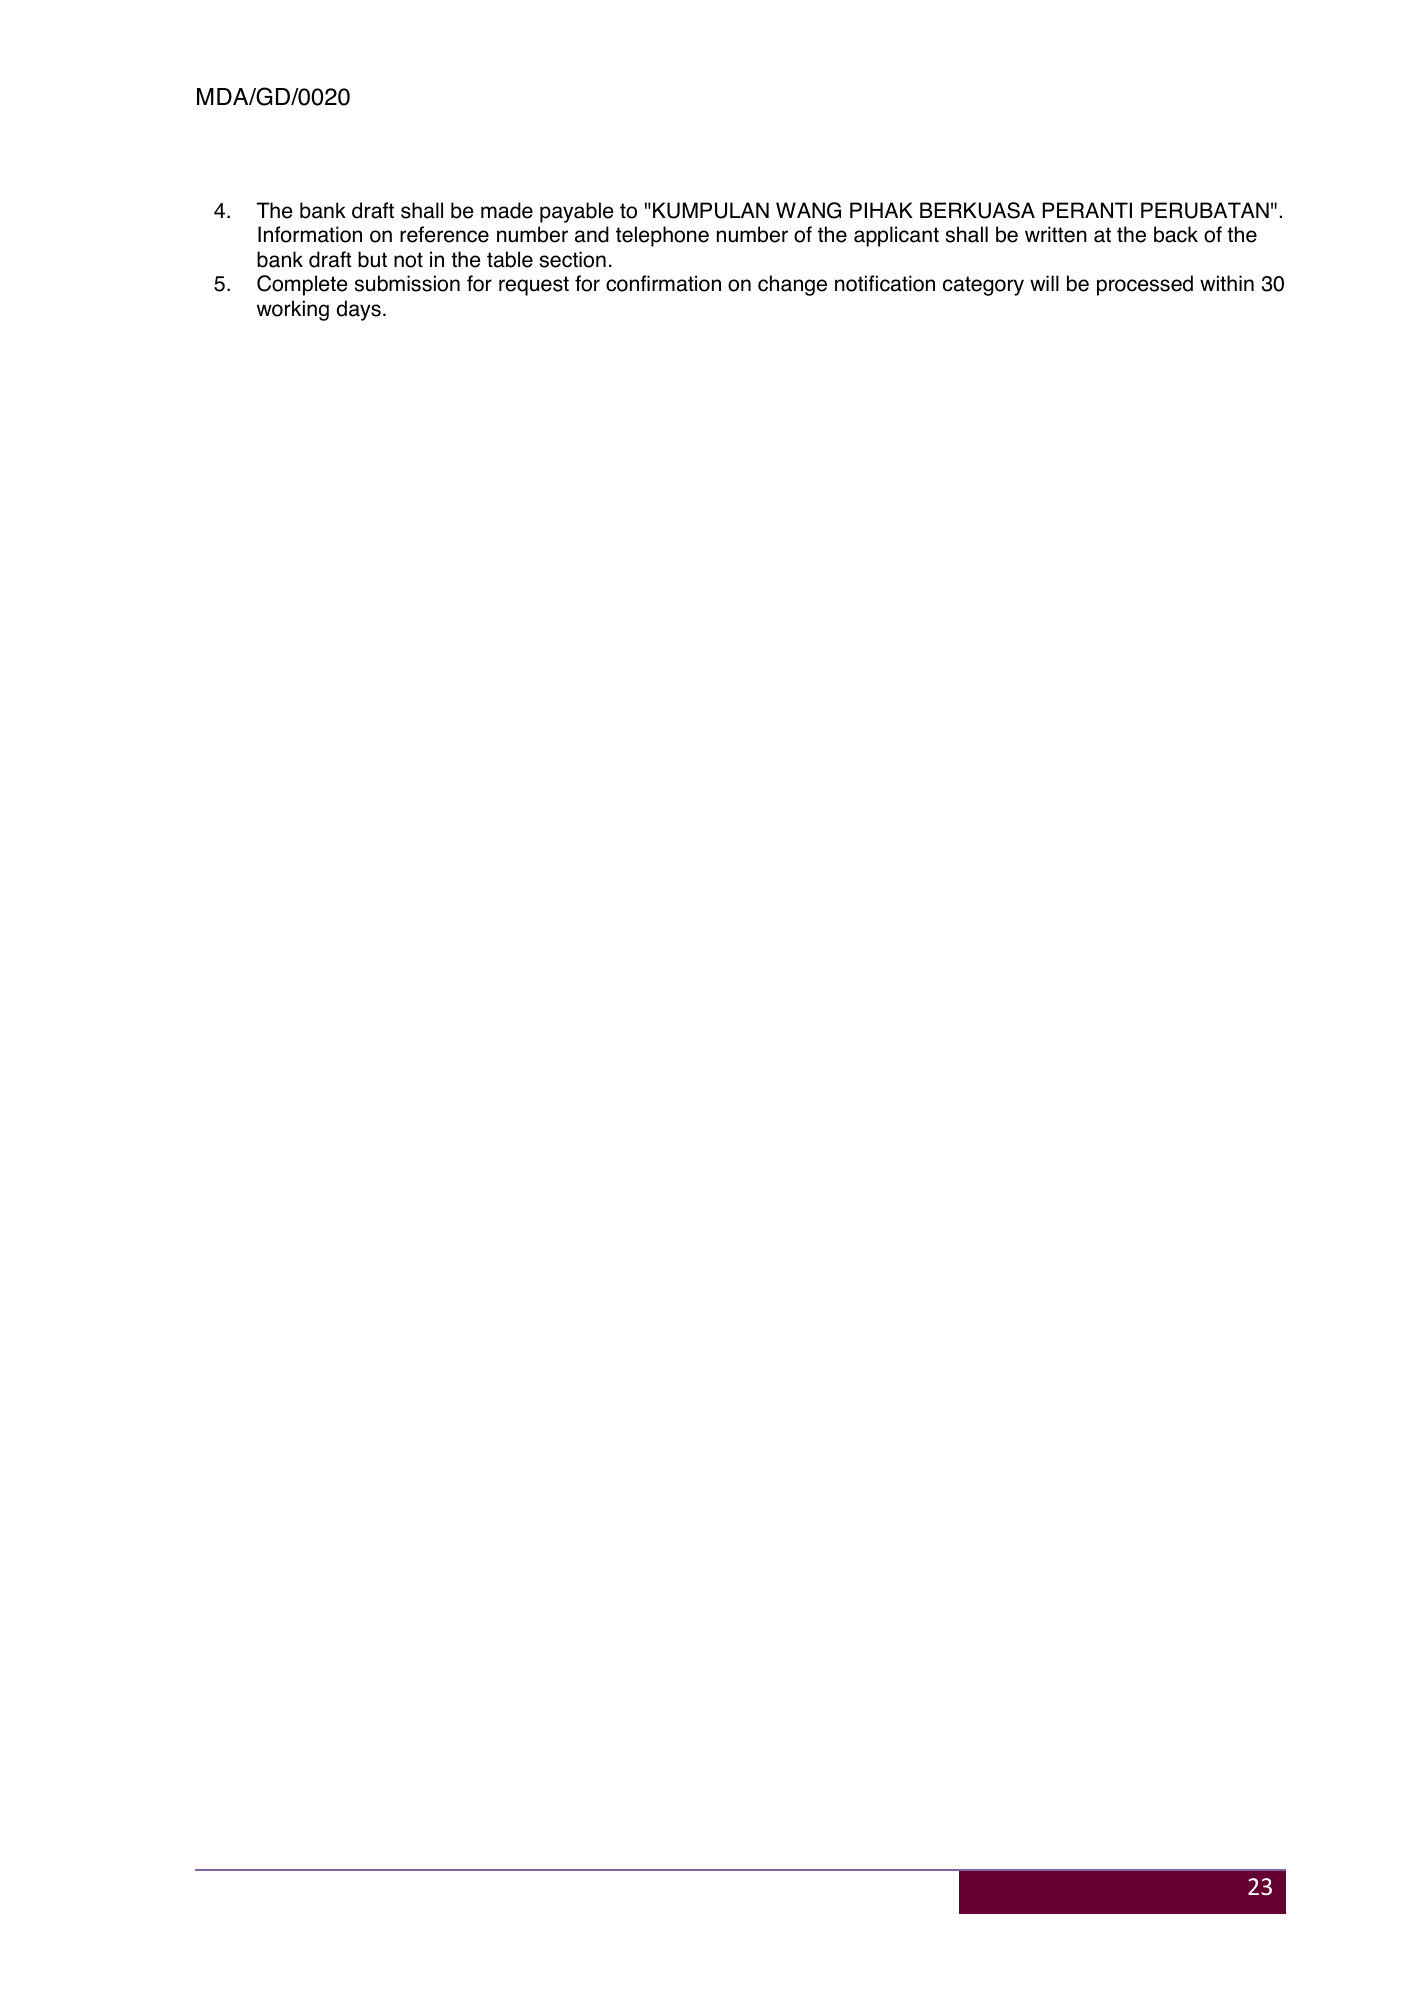
MDA/GD/0020 4. The bank draft shall be made payable to "KUMPULAN WANG PIHAK BERKUASA PERANTI PERUBATAN". Information on reference number and telephone number of the applicant shall be written at the back of the bank draft but not in the table section. 5. Complete submission for request for confirmation on change notification category will be processed within 30 working days. 23
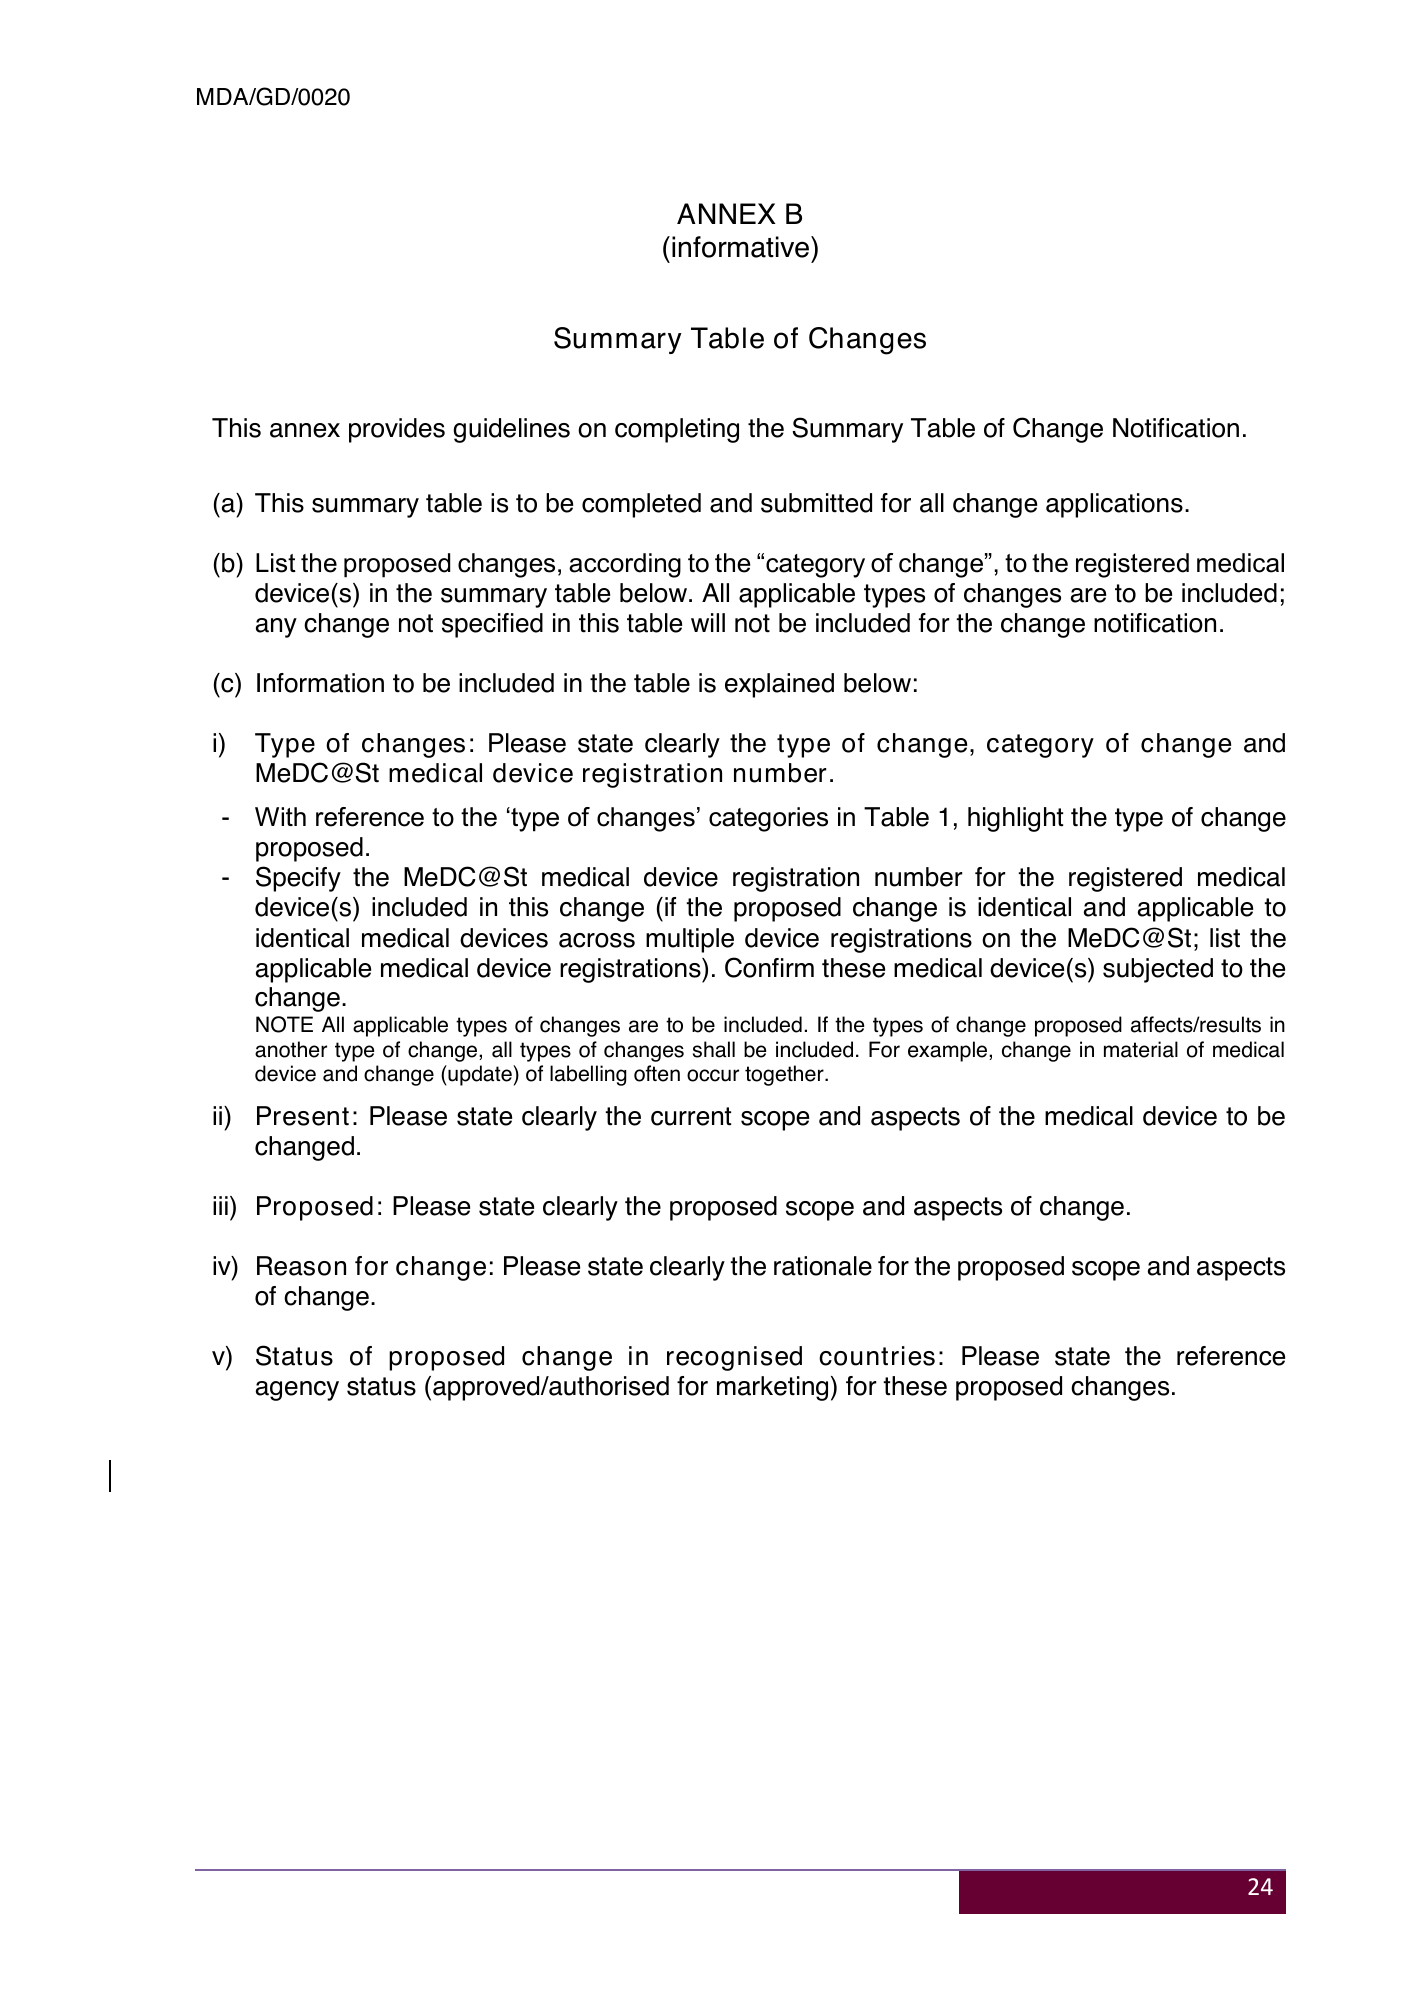
MDA/GD/0020 ANNEX B (informative) Summary Table of Changes This annex provides guidelines on completing the Summary Table of Change Notification. (a) This summary table is to be completed and submitted for all change applications. (b) List the proposed changes, according to the “category of change”, to the registered medical device(s) in the summary table below. All applicable types of changes are to be included; any change not specified in this table will not be included for the change notification. (c) Information to be included in the table is explained below: i) Type of changes: Please state clearly the type of change, category of change and MeDC@St medical device registration number. - With reference to the ‘type of changes’ categories in Table 1, highlight the type of change proposed. - Specify the MeDC@St medical device registration number for the registered medical device(s) included in this change (if the proposed change is identical and applicable to identical medical devices across multiple device registrations on the MeDC@St; list the applicable medical device registrations). Confirm these medical device(s) subjected to the change. NOTE All applicable types of changes are to be included. If the types of change proposed affects/results in another type of change, all types of changes shall be included. For example, change in material of medical device and change (update) of labelling often occur together. ii) Present: Please state clearly the current scope and aspects of the medical device to be changed. iii) Proposed: Please state clearly the proposed scope and aspects of change. iv) Reason for change: Please state clearly the rationale for the proposed scope and aspects of change. v) Status of proposed change in recognised countries: Please state the reference agency status (approved/authorised for marketing) for these proposed changes. 24
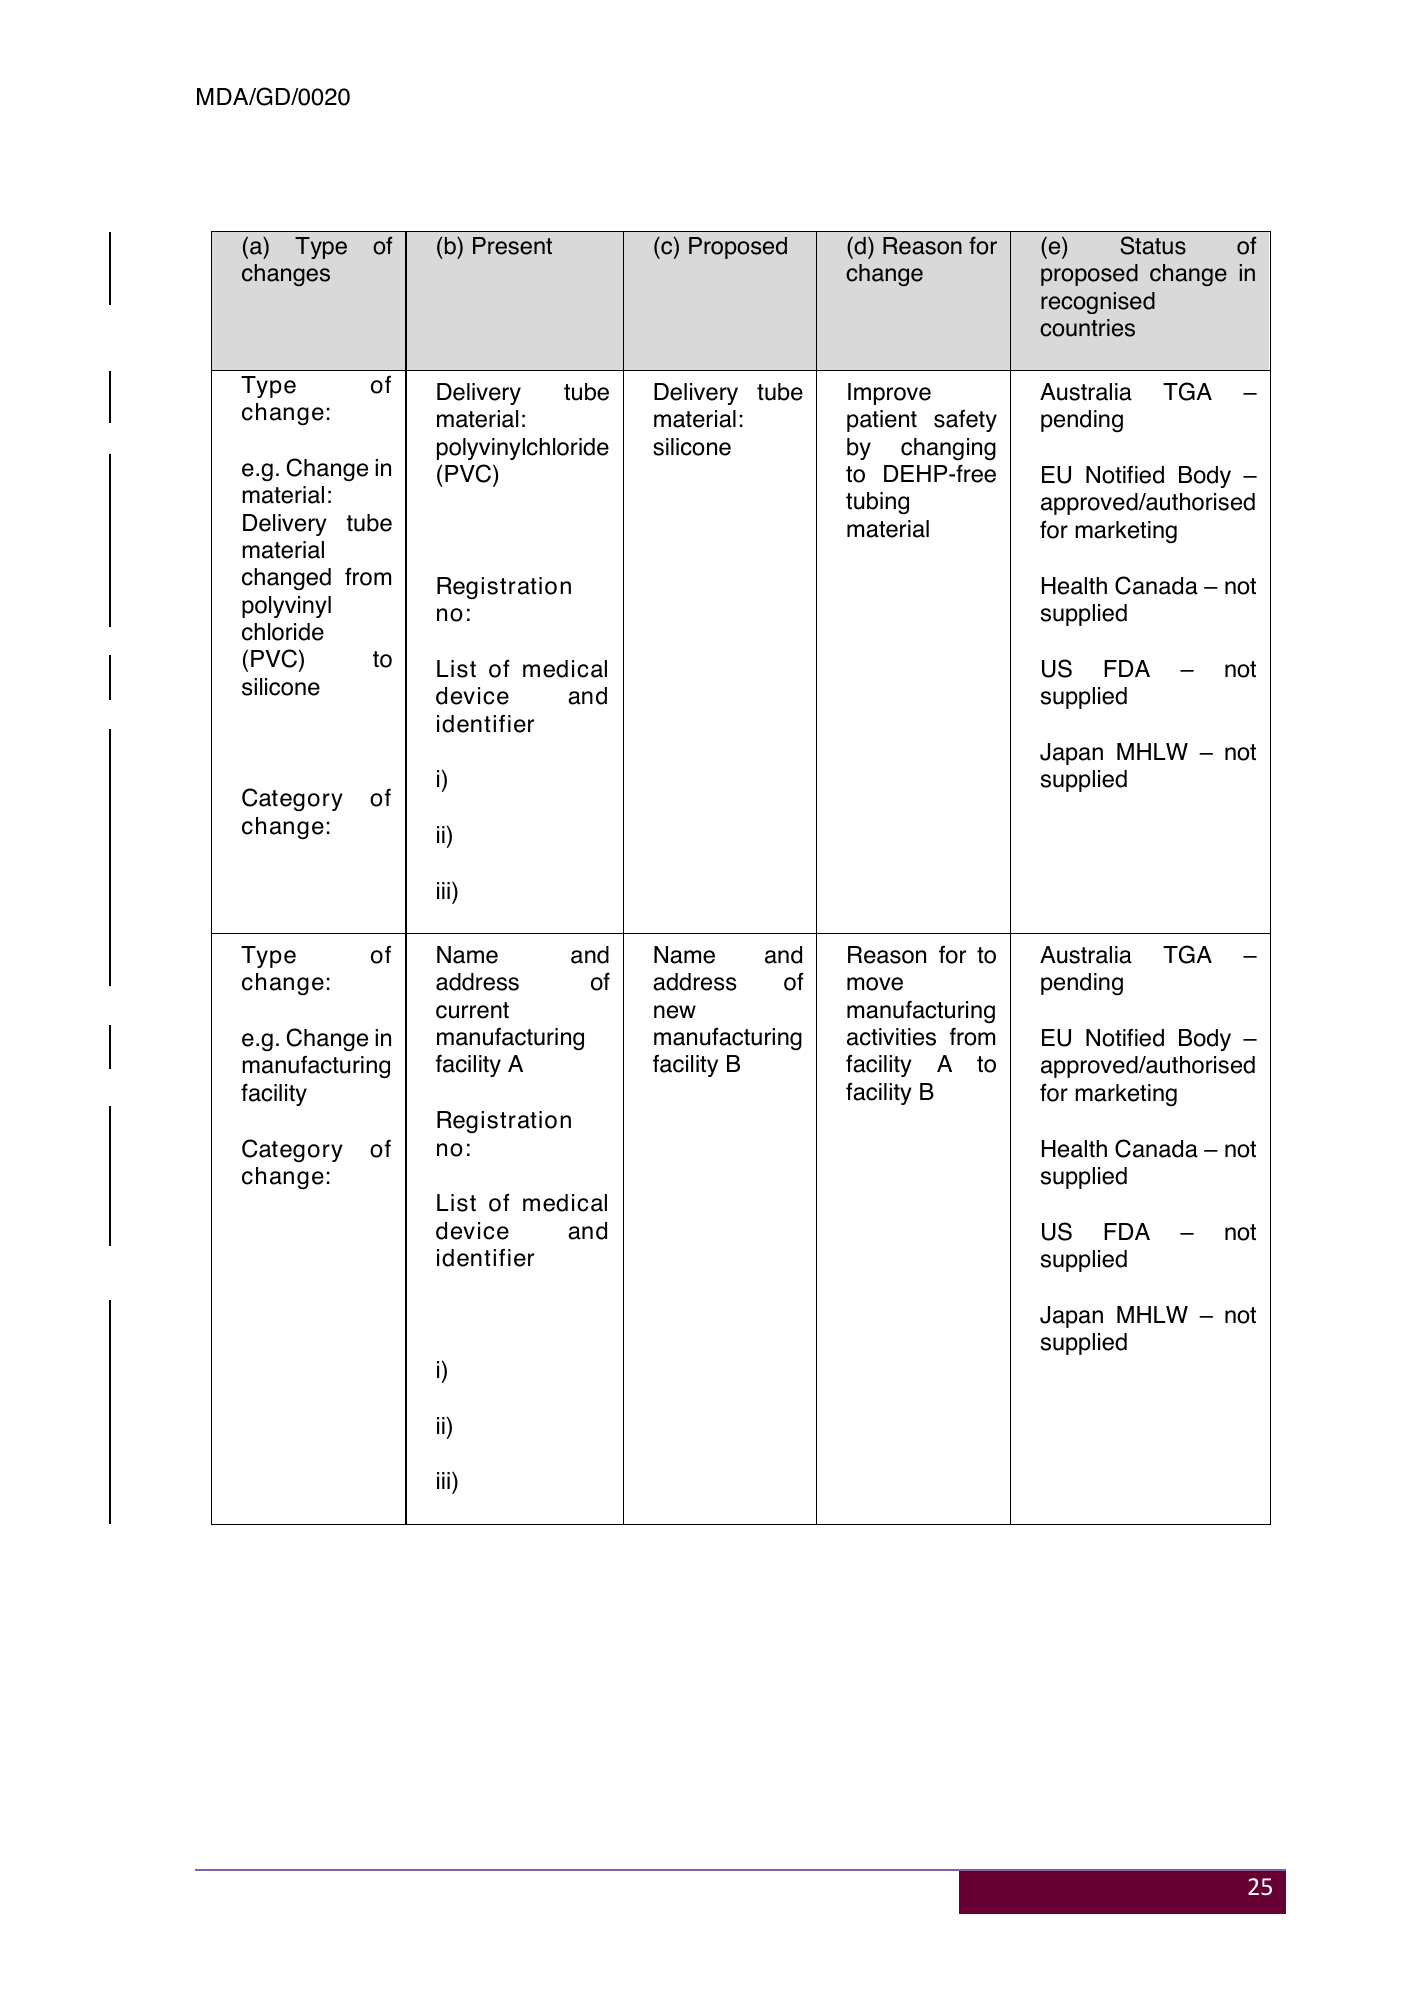
MDA/GD/0020 (a) Type of (b) Present (c) Proposed (d) Reason for (e) Status of changes change proposed change in recognised countries Type of Delivery tube Delivery tube Improve Australia TGA – change: material: material: patient safety pending polyvinylchloride silicone by changing e.g. Change in (PVC) to DEHP-free EU Notified Body – material: tubing approved/authorised Delivery tube material for marketing material changed from Registration Health Canada – not polyvinyl no: supplied chloride (PVC) to List of medical US FDA – not silicone device and supplied identifier Japan MHLW – not i) supplied Category of change: ii) iii) Type of Name and Name and Reason for to Australia TGA – change: address of address of move pending current new manufacturing e.g. Change in manufacturing manufacturing activities from EU Notified Body – manufacturing facility A facility B facility A to approved/authorised facility facility B for marketing Registration Category of no: Health Canada – not change: supplied List of medical device and US FDA – not identifier supplied Japan MHLW – not supplied i) ii) iii) 25
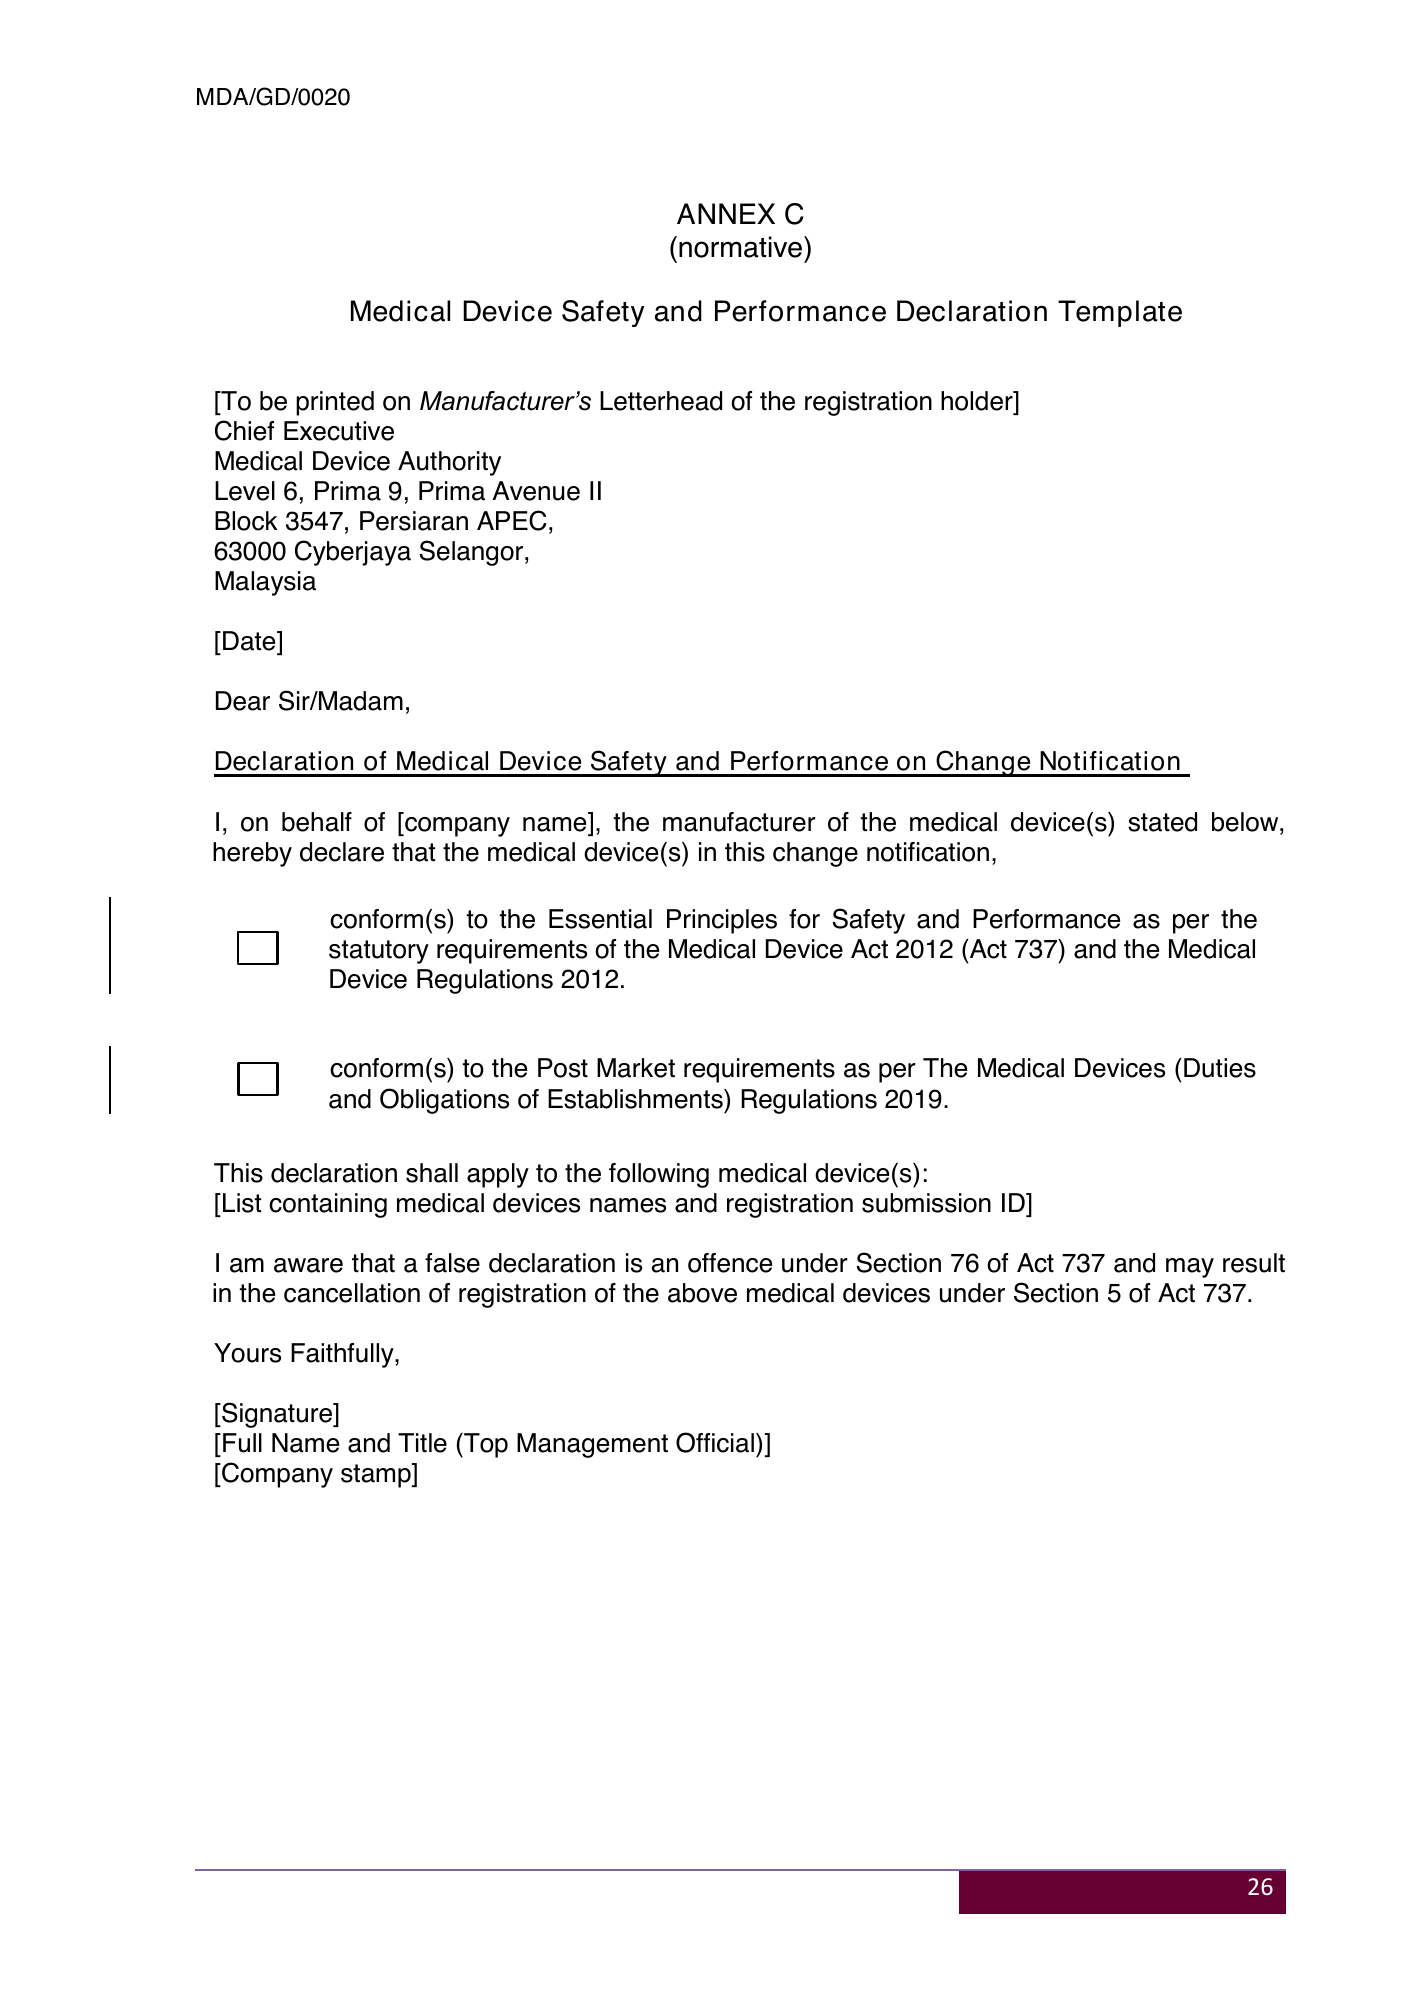
MDA/GD/0020 ANNEX C (normative) Medical Device Safety and Performance Declaration Template [To be printed on Manufacturer’s Letterhead of the registration holder] Chief Executive Medical Device Authority Level 6, Prima 9, Prima Avenue II Block 3547, Persiaran APEC, 63000 Cyberjaya Selangor, Malaysia [Date] Dear Sir/Madam, Declaration of Medical Device Safety and Performance on Change Notification I, on behalf of [company name], the manufacturer of the medical device(s) stated below, hereby declare that the medical device(s) in this change notification, conform(s) to the Essential Principles for Safety and Performance as per the statutory requirements of the Medical Device Act 2012 (Act 737) and the Medical Device Regulations 2012. conform(s) to the Post Market requirements as per The Medical Devices (Duties and Obligations of Establishments) Regulations 2019. This declaration shall apply to the following medical device(s): [List containing medical devices names and registration submission ID] I am aware that a false declaration is an offence under Section 76 of Act 737 and may result in the cancellation of registration of the above medical devices under Section 5 of Act 737. Yours Faithfully, [Signature] [Full Name and Title (Top Management Official)] [Company stamp] 26
MDA/GD/0020 MEDICAL DEVICE AUTHORITY MINISTRY OF HEALTH, MALAYSIA Contact Information: Medical Device Authority Ministry of Health Malaysia Level 6, Prima 9, Prima Avenue II Block 3547, Persiaran APEC 63000 Cyberjaya, Selangor MALAYSIA T: (03) 8230 0300 F: (03) 8230 0200 Website: http://www.mda.gov.my 27