Currentness under review. Whether this instrument remains applicable is under review. Consult the later instruments and source context below before relying on it.
Link directly to a passage with its anchor — for example #pdf-page-1 — to open it highlighted. This reference version is generated from the authority’s publication and does not replace it. Language: English.
Contents
Republic of Sudan
Federal Ministry of Health
National Medicines & Poisons Board
Secretariat General
Requirements for Variations
General Notes:
It is important to note that "National Medicines and Poisons Board NMPB" reserves the right to request any additional information and data not specifically described in this document, in order to assess adequately the safety, efficacy and quality of drug products.
Applicants should be aware that deficient documentation can lead to rejection of the application.
A. Variation required submission of application:
Variations that affect the pharmaceutical and non-pharmaceutical products or their manufacturers are classified as follows:
1. Change in the name and /or address of the marketing authorization holder (MAH).
2. Change in the manufacturing Site of the finished product.
3. Changes in the composition formula of the finished Product (excipients).
4. Change in trade name of medicinal product.
5. Change in any part of the secondary/outer pack of the finished product.
6. Change in any part of the primary/inner pack of the finished product.
7. Change in the shelf life of finished product.
8. Change in the finished product specifications.
9. Change/ addition in pack size of the finished product.
10. Change in the shape or dimensions of the pharmaceutical form (for solid dosage form)
11. Change of the name / Modification of the address of a foreign manufacturer
12. Changing or adding a new agency.
B. Variation not required submission of application:
• For any change (s) other than the previously mentioned variation in (A), a notification letter only from the manufacturer to submitted to NMPB.
The following documents should take into consideration when submitting any variation application:
Code. No RCh/R/01/01 Issue. NO (3) Issue Date 01/08/2012 Change. NO (1) Change Date28/07/2020 Expire date01/08/2022 Page 1of5
Original PDF page 1 — retained source fields, tables and layout.
1. Change in the name and /or address of the marketing authorization holder (MAH):
1.1 Justification for changing the name and /or address of the marketing authorization holder (MAH).
1.2 A formal document from a relevant official body in which the new name or new address is mention.
1.3 Replacement of the relevant pages of the Dossier that affected by the variation (inner, outer pack).
2. Change in the manufacturing Site of the finished product:
2.1 Justification for changing the manufacturing site.
2.2 Replacement of the relevant pages of the dossier that are affected by the variation (Inner, outer pack).
2.3 Certificate of a Pharmaceutical Product (CPP) (for foreign products) from the relevant competent health authority for human & veterinary Pharmaceutical Products (The composition should be attached with CPP), and free sale certificate for cosmetic, dietary supplement & health products, and European Certificate (EC) or FDA for medical devices.
2.4 Real time stability studies from the new manufacturing site for at least two pilot-scale batches and commitment letter to complete the stability studies to cover the shelf life must be submitted (must cover at least three months).
2.5 A declaration by the company that the manufacturing process will remain the same. In addition, the API(s), excipient(s) and their source(s), dosage form, concentration, the primary and secondary packaging, labeling, and all specifications for the product must remain the same as previously approved in the old site. A clarification of any proposed change(s) to the manufacturing of the product at the new manufacturing site should be provided and justified.
2.6 For solid dosage forms, data from comparative dissolution tests with demonstration of similarity of dissolution profile, performed on the last three batches from the previous site and the first three batches from the new site should be submitted.
2.7 Validation of the analytical methods needed for batch release (according to the release specifications) from the proposed secondary packaging site and/or validation for transportation process from manufacturing site to secondary packaging site along with release certificate from secondary packaging site covering all processes from receiving the semi-finished product to final pack.
N.B. If the change is related to Manufacturer responsible for quality control or the finished product release the stability study is not required.
3. Changes in the composition formula of the finished Product (excipients):
Code. No RCh/R/01/01 Issue. NO (3) Issue Date 01/08/2019 Change. NO (1) Change Date28/07/2020 Expire date01/08/2022 Page 2of5
Original PDF page 2 — retained source fields, tables and layout.
3.1 Justification for the change/choice of excipients etc. must be given by appropriate development pharmaceutics.
3.2 Copy of the approval document (for the change) from health authority in country of origin.
3.3 Certificate of a Pharmaceutical Product (CPP) (for foreign products) from the relevant competent health authority for human & veterinary Pharmaceutical Products (The composition should be attached with CPP) and free sale certificate for cosmetic, dietary supplement & health products.
3.4 Real time stability studies from the new manufacturing site for at least two pilot-scale batches and commitment letter to complete the stability studies to cover the shelf life must be submitted (must cover at least three months).
3.5 A certificate stating that the product is free from (TSE/BSE) and a Halal certificate if source of the active, Inactive and/ or any other substance within the Product is of animal source.
3.6 Comparative table that clearly outline the change (current and proposed).
3.7 For solid dosage forms, comparative dissolution Profile data of at least two pilot scale batches of the finished product in the new and old composition. For herbal medicinal products, comparative disintegration data may be acceptable. (Unless the change is related to the coloring of flavoring agent)
3.8 For veterinary medicines intended for use in food producing, justification that the excipient does not have pharmacological activity at the dose at which it is administered to the target animal.
4. Change in trade name of medicinal product:
4.1 Justification for changing the trade name of medicinal product.
4.2 Replacement of the relevant pages of the dossier that are affected by the variation (Inner, outer pack).
4.3 Certificate of a Pharmaceutical Product (CPP) (for foreign products) from the relevant competent health authority for human & veterinary Pharmaceutical Products (The composition should be attached with CPP), and free sale certificate for cosmetic, dietary supplement & health products, and European Certificate (EC) or FDA for medical devices.
5. Change in any part of the secondary/outer pack of the finished product:
5.1 New outer pack artwork (Colored Mock- up)
6. Change in any part of the primary/inner pack of the finished product:
6.1 New inner pack artwork (Colored Mock- up).
Code. No RCh/R/01/01 Issue. NO (3) Issue Date 01/08/2019 Change. NO (1) Change Date28/07/2020 Expire date01/08/2022 Page 3of5
Original PDF page 3 — retained source fields, tables and layout.
6.2 Real time stability studies from the new manufacturing site for at least two pilot-scale batches and commitment letter to complete the stability studies to cover the shelf life must be submitted (must cover at least three months).
7. Change in the shelf life of finished product:
A. Reduction of shelf life of finished:
1. Justification for the reduction in the shelf-life from the manufacturer.
2. Copy of the approval document (for the change) from health authority in country of origin.
B. Extension of shelf life of finished:
1. Copy of the approval document (for the change) from health authority in country of origin.
2. Recent real time stability studies covering the proposed shelf life, on at least three production scale batches of the finished product and/or after first opening or reconstitution (in-use stability).
8. Change in the finished product specifications:
8.1 Justification of the new specification parameter and limits.
8.2 Comparative table of current and proposed specifications.
8.3 Details of any new analytical method and validation data.
8.4 Comparative study between pharmacopeia method and in-house method with statistical data (in case of in-house method)
9. Change/ addition in pack size of the finished product:
9.1 Justification for the new/remaining pack-size, showing that the new/remaining size is/are consistent with the dosage regimen and duration of use as approved in the summary of product characteristics.
9.2 Real time stability studies for at least two pilot-scale batches and commitment letter to complete the stability studies to cover the shelf life must submitted (must cover at least three months).
9.3 A declaration that material of the primary / inner package has not been changed from the previously approved one.
10. Change in the shape or dimensions of the pharmaceutical form (for solid dosage form):
Copy of the approval document (for the change) from health authority in country of Origin.
Comparative dissolution data on at least one pilot batch of the current and
Code. No RCh/R/01/01 Issue. NO (3) Issue Date 01/08/2019 Change. NO (1) Change Date28/07/2020 Expire date01/08/2022 Page 4of5
Original PDF page 4 — retained source fields, tables and layout.
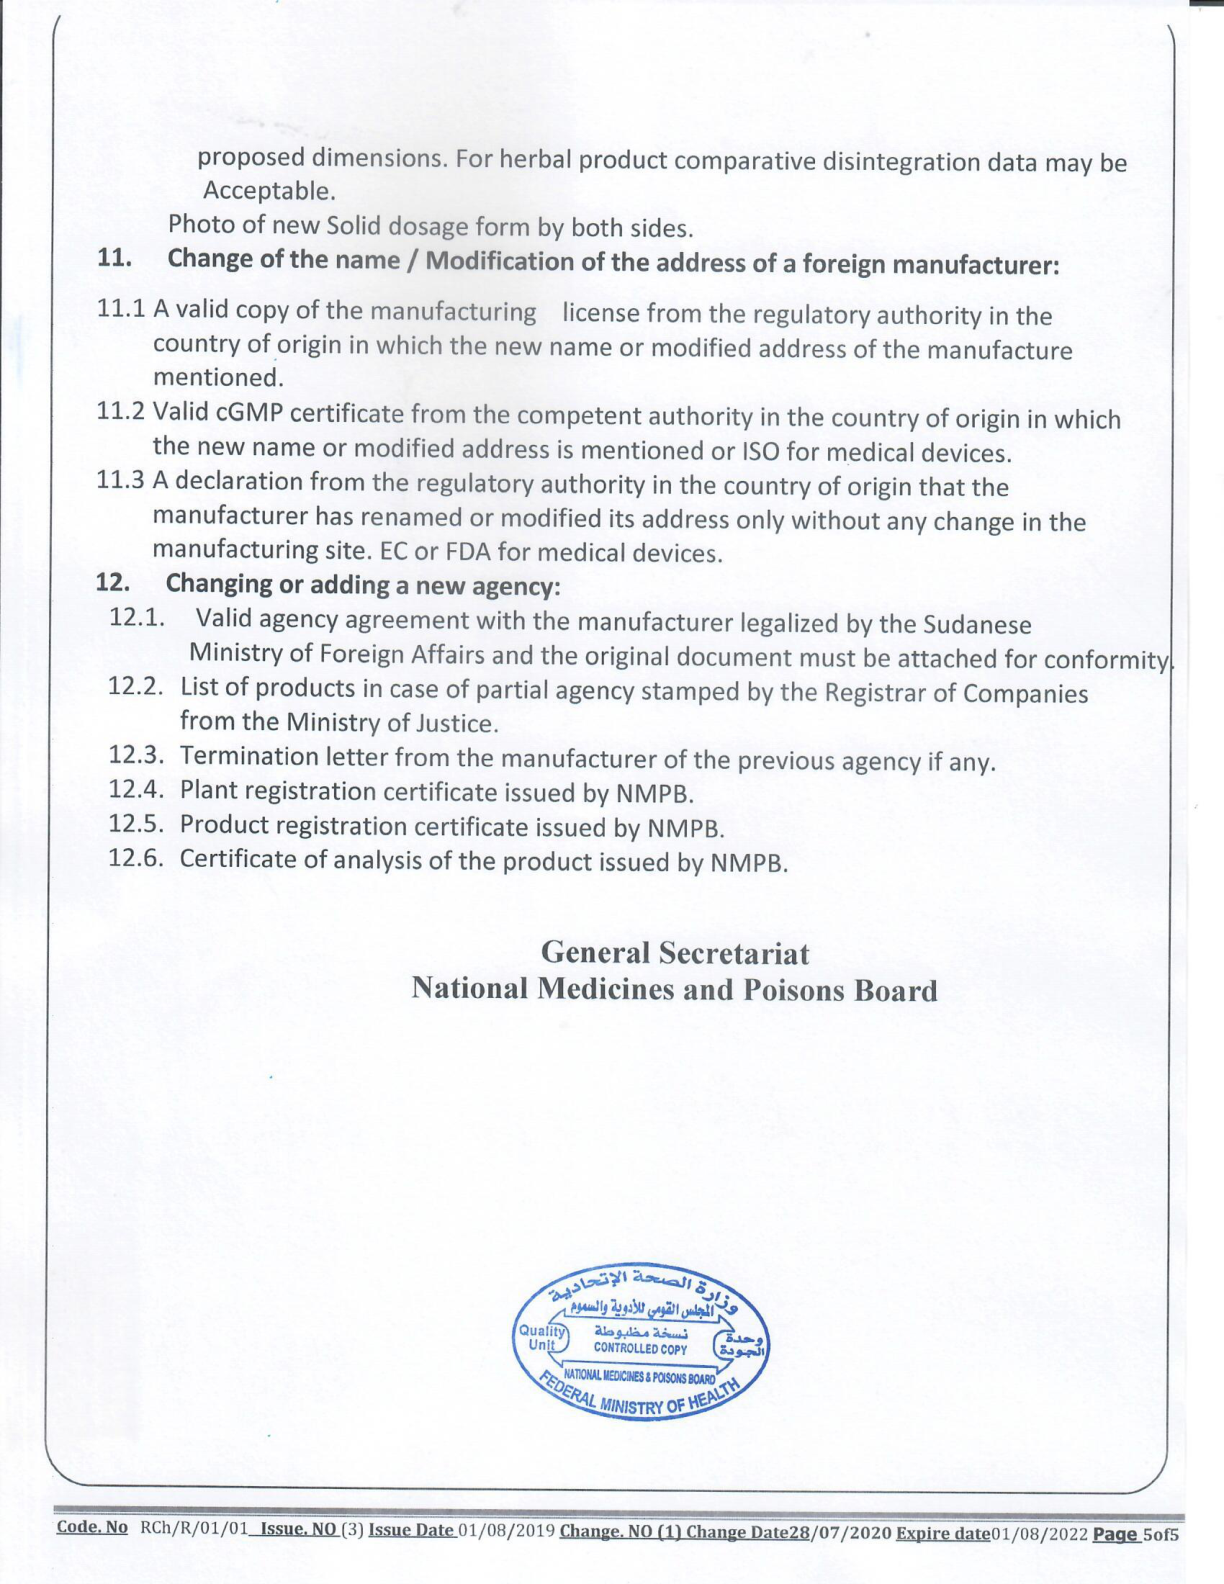
proposed dimensions. For herbal product comparative disintegration data may be Acceptable.
Photo of new Solid dosage form by both sides.
11. Change of the name / Modification of the address of a foreign manufacturer:
11.1 A valid copy of the manufacturing license from the regulatory authority in the country of origin in which the new name or modified address of the manufacture mentioned.
11.2 Valid cGMP certificate from the competent authority in the country of origin in which the new name or modified address is mentioned or ISO for medical devices.
11.3 A declaration from the regulatory authority in the country of origin that the manufacturer has renamed or modified its address only without any change in the manufacturing site. EC or FDA for medical devices.
12. Changing or adding a new agency:
12.1. Valid agency agreement with the manufacturer legalized by the Sudanese Ministry of Foreign Affairs and the original document must be attached for conformity.
12.2. List of products in case of partial agency stamped by the Registrar of Companies from the Ministry of Justice.
12.3. Termination letter from the manufacturer of the previous agency if any.
12.4. Plant registration certificate issued by NMPB.
12.5. Product registration certificate issued by NMPB.
12.6. Certificate of analysis of the product issued by NMPB.
General Secretariat
National Medicines and Poisons Board
Code. No RCh/R/01/01 Issue. NO (3) Issue Date 01/08/2019 Change. NO (1) Change Date28/07/2020 Expire date01/08/2022 Page 5of5
Original PDF page 5 — retained source fields, tables and layout.